return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHFCH3 (2-Fluoropropane)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-217.872967
Energy at 298.15K-217.880757
HF Energy-217.147017
Nuclear repulsion energy131.161168
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3160 3087        
2 A' 3153 3080        
3 A' 3067 2997        
4 A' 3059 2988        
5 A' 1554 1518        
6 A' 1537 1501        
7 A' 1461 1427        
8 A' 1410 1377        
9 A' 1234 1205        
10 A' 1183 1155        
11 A' 981 958        
12 A' 843 824        
13 A' 480 469        
14 A' 354 346        
15 A' 272 265        
16 A" 3157 3084        
17 A" 3146 3074        
18 A" 3064 2993        
19 A" 1532 1496        
20 A" 1526 1491        
21 A" 1460 1426        
22 A" 1414 1382        
23 A" 1196 1168        
24 A" 970 947        
25 A" 949 927        
26 A" 405 396        
27 A" 230 225        

Unscaled Zero Point Vibrational Energy (zpe) 21397.3 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 20903.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.28305 0.26604 0.15677

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.289 0.243 0.000
F2 -0.899 1.045 0.000
H3 1.129 0.964 0.000
C4 0.289 -0.587 1.279
C5 0.289 -0.587 -1.279
H6 1.218 -1.185 1.339
H7 1.218 -1.185 -1.339
H8 0.234 0.068 2.165
H9 0.234 0.068 -2.165
H10 -0.576 -1.273 1.287
H11 -0.576 -1.273 -1.287

Atom - Atom Distances (Å)
  C1 F2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.43321.10621.52481.52482.16642.16642.17272.17272.16892.1689
F21.43322.02892.38952.38953.35283.35282.63122.63122.67072.6707
H31.10622.02892.17842.17842.53322.53322.50802.50803.09293.0929
C41.52482.38952.17842.55831.10562.84171.10313.50631.10472.7934
C51.52482.38952.17842.55832.84171.10563.50631.10312.79341.1047
H62.16643.35282.53321.10562.84172.67871.79403.84931.79673.1813
H72.16643.35282.53322.84171.10562.67873.84931.79403.18131.7967
H82.17272.63122.50801.10313.50631.79403.84934.32991.79653.7906
H92.17272.63122.50803.50631.10313.84931.79404.32993.79061.7965
H102.16892.67073.09291.10472.79341.79673.18131.79653.79062.5730
H112.16892.67073.09292.79341.10473.18131.79673.79061.79652.5730

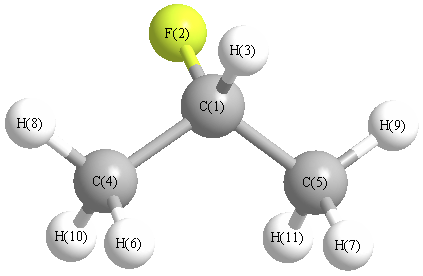
picture of 2-Fluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H8 110.512 C1 C4 H10 110.111
C1 C5 H7 109.869 C1 C5 H9 110.512
C1 C5 H11 110.111 F2 C1 H3 105.340
F2 C1 C4 107.723 F2 C1 C5 107.723
H3 C1 C4 110.771 H3 C1 C5 110.771
C4 C1 C5 114.044 H7 C5 H9 108.634
H7 C5 H11 108.753 H8 C4 H10 108.920
H9 C5 H11 108.920
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability