Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -377.497831 |
| Energy at 298.15K | |
| HF Energy | -377.497831 |
| Nuclear repulsion energy | 190.284893 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3168 |
3035 |
15.37 |
33.53 |
0.70 |
0.83 |
| 2 |
A |
3127 |
2997 |
19.88 |
93.07 |
0.25 |
0.41 |
| 3 |
A |
3095 |
2966 |
16.01 |
123.58 |
0.05 |
0.10 |
| 4 |
A |
1471 |
1410 |
8.32 |
3.70 |
0.74 |
0.85 |
| 5 |
A |
1446 |
1385 |
12.71 |
1.48 |
0.55 |
0.71 |
| 6 |
A |
1378 |
1320 |
15.18 |
2.26 |
0.74 |
0.85 |
| 7 |
A |
1322 |
1267 |
9.21 |
1.34 |
0.75 |
0.86 |
| 8 |
A |
1256 |
1204 |
14.01 |
3.48 |
0.67 |
0.80 |
| 9 |
A |
1164 |
1115 |
80.21 |
4.09 |
0.64 |
0.78 |
| 10 |
A |
1130 |
1083 |
14.93 |
1.89 |
0.26 |
0.42 |
| 11 |
A |
1110 |
1063 |
278.01 |
0.11 |
0.55 |
0.71 |
| 12 |
A |
1089 |
1043 |
17.69 |
7.01 |
0.39 |
0.56 |
| 13 |
A |
906 |
868 |
34.66 |
4.28 |
0.24 |
0.39 |
| 14 |
A |
572 |
548 |
3.42 |
1.90 |
0.25 |
0.40 |
| 15 |
A |
470 |
451 |
18.44 |
0.55 |
0.73 |
0.84 |
| 16 |
A |
422 |
405 |
5.54 |
1.28 |
0.43 |
0.60 |
| 17 |
A |
241 |
231 |
8.10 |
0.07 |
0.54 |
0.70 |
| 18 |
A |
112 |
108 |
8.21 |
0.05 |
0.72 |
0.83 |
Unscaled Zero Point Vibrational Energy (zpe) 11739.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 11250.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.776 |
-0.585 |
-0.290 |
| C2 |
0.463 |
0.019 |
0.327 |
| F3 |
-1.885 |
0.111 |
0.155 |
| F4 |
1.531 |
-0.768 |
-0.004 |
| F5 |
0.693 |
1.261 |
-0.182 |
| H6 |
-0.723 |
-0.512 |
-1.383 |
| H7 |
-0.868 |
-1.633 |
0.021 |
| H8 |
0.414 |
0.104 |
1.420 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5107 | 1.3837 | 2.3318 | 2.3618 | 1.0969 | 1.0969 | 2.1945 |
C2 | 1.5107 | | 2.3571 | 1.3675 | 1.3617 | 2.1483 | 2.1439 | 1.0976 | F3 | 1.3837 | 2.3571 | | 3.5316 | 2.8437 | 2.0268 | 2.0233 | 2.6241 | F4 | 2.3318 | 1.3675 | 3.5316 | | 2.2027 | 2.6548 | 2.5506 | 2.0101 | F5 | 2.3618 | 1.3617 | 2.8437 | 2.2027 | | 2.5675 | 3.2946 | 1.9958 | H6 | 1.0969 | 2.1483 | 2.0268 | 2.6548 | 2.5675 | | 1.8028 | 3.0874 | H7 | 1.0969 | 2.1439 | 2.0233 | 2.5506 | 3.2946 | 1.8028 | | 2.5727 | H8 | 2.1945 | 1.0976 | 2.6241 | 2.0101 | 1.9958 | 3.0874 | 2.5727 | |
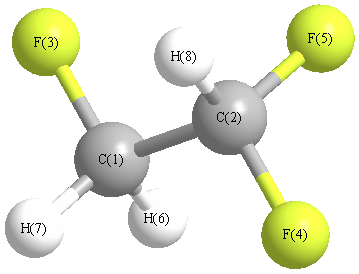 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.126 |
|
C1 |
C2 |
F5 |
110.514 |
| C1 |
C2 |
H8 |
113.625 |
|
C2 |
C1 |
F3 |
108.970 |
| C2 |
C1 |
H6 |
109.936 |
|
C2 |
C1 |
H7 |
109.586 |
| F3 |
C1 |
H6 |
109.039 |
|
F3 |
C1 |
H7 |
108.752 |
| F4 |
C2 |
F5 |
107.626 |
|
F4 |
C2 |
H8 |
108.765 |
| F5 |
C2 |
H8 |
108.011 |
|
H6 |
C1 |
H7 |
110.525 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.708 |
|
|
|
| 2 |
C |
1.167 |
|
|
|
| 3 |
F |
-0.434 |
|
|
|
| 4 |
F |
-0.449 |
|
|
|
| 5 |
F |
-0.440 |
|
|
|
| 6 |
H |
-0.148 |
|
|
|
| 7 |
H |
-0.134 |
|
|
|
| 8 |
H |
-0.269 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.313 |
-1.497 |
0.365 |
1.573 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.717 |
1.716 |
1.581 |
| y |
1.716 |
-28.115 |
0.530 |
| z |
1.581 |
0.530 |
-24.886 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.217 |
1.716 |
1.581 |
| y |
1.716 |
0.687 |
0.530 |
| z |
1.581 |
0.530 |
5.529 |
|
| Polar |
|---|
| 3z2-r2 | 11.059 |
|---|
| x2-y2 | -4.603 |
|---|
| xy | 1.716 |
|---|
| xz | 1.581 |
|---|
| yz | 0.530 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.614 |
-0.056 |
-0.019 |
| y |
-0.056 |
4.443 |
0.060 |
| z |
-0.019 |
0.060 |
4.166 |
<r2> (average value of r
2) Å
2
| <r2> |
111.942 |
| (<r2>)1/2 |
10.580 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -377.495882 |
| Energy at 298.15K | |
| HF Energy | -377.495882 |
| Nuclear repulsion energy | 193.013331 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3099 |
2970 |
48.91 |
136.33 |
0.06 |
0.11 |
| 2 |
A' |
3082 |
2953 |
4.51 |
71.40 |
0.22 |
0.37 |
| 3 |
A' |
1465 |
1404 |
9.73 |
3.29 |
0.74 |
0.85 |
| 4 |
A' |
1428 |
1368 |
21.84 |
2.28 |
0.39 |
0.56 |
| 5 |
A' |
1391 |
1333 |
13.96 |
1.24 |
0.70 |
0.82 |
| 6 |
A' |
1193 |
1143 |
118.83 |
4.17 |
0.18 |
0.31 |
| 7 |
A' |
1107 |
1061 |
31.12 |
3.59 |
0.75 |
0.85 |
| 8 |
A' |
879 |
842 |
30.43 |
7.39 |
0.10 |
0.18 |
| 9 |
A' |
758 |
727 |
51.14 |
2.23 |
0.46 |
0.63 |
| 10 |
A' |
510 |
489 |
11.82 |
1.05 |
0.74 |
0.85 |
| 11 |
A' |
230 |
220 |
1.50 |
0.19 |
0.38 |
0.55 |
| 12 |
A" |
3149 |
3018 |
12.56 |
48.08 |
0.75 |
0.86 |
| 13 |
A" |
1368 |
1311 |
20.87 |
1.02 |
0.75 |
0.86 |
| 14 |
A" |
1275 |
1221 |
20.88 |
6.40 |
0.75 |
0.86 |
| 15 |
A" |
1132 |
1085 |
125.08 |
1.69 |
0.75 |
0.86 |
| 16 |
A" |
946 |
907 |
60.57 |
2.12 |
0.75 |
0.86 |
| 17 |
A" |
366 |
351 |
0.47 |
0.23 |
0.75 |
0.86 |
| 18 |
A" |
113 |
108 |
1.99 |
0.09 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 11745.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 11255.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.737 |
-0.835 |
0.000 |
| C2 |
0.358 |
0.626 |
0.000 |
| F3 |
-0.388 |
-1.630 |
0.000 |
| F4 |
-0.388 |
0.931 |
1.099 |
| F5 |
-0.388 |
0.931 |
-1.099 |
| H6 |
1.327 |
-1.055 |
-0.900 |
| H7 |
1.327 |
-1.055 |
0.900 |
| H8 |
1.250 |
1.270 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
F5 |
H6 |
H7 |
H8 |
| C1 | | 1.5090 | 1.3778 | 2.3644 | 2.3644 | 1.0981 | 1.0981 | 2.1667 |
C2 | 1.5090 | | 2.3760 | 1.3627 | 1.3627 | 2.1387 | 2.1387 | 1.1003 | F3 | 1.3778 | 2.3760 | | 2.7863 | 2.7863 | 2.0200 | 2.0200 | 3.3308 | F4 | 2.3644 | 1.3627 | 2.7863 | | 2.1970 | 3.2983 | 2.6316 | 2.0015 | F5 | 2.3644 | 1.3627 | 2.7863 | 2.1970 | | 2.6316 | 3.2983 | 2.0015 | H6 | 1.0981 | 2.1387 | 2.0200 | 3.2983 | 2.6316 | | 1.7994 | 2.4947 | H7 | 1.0981 | 2.1387 | 2.0200 | 2.6316 | 3.2983 | 1.7994 | | 2.4947 | H8 | 2.1667 | 1.1003 | 3.3308 | 2.0015 | 2.0015 | 2.4947 | 2.4947 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.739 |
|
C1 |
C2 |
F5 |
110.739 |
| C1 |
C2 |
H8 |
111.311 |
|
C2 |
C1 |
F3 |
110.705 |
| C2 |
C1 |
H6 |
109.221 |
|
C2 |
C1 |
H7 |
109.221 |
| F3 |
C1 |
H6 |
108.825 |
|
F3 |
C1 |
H7 |
108.825 |
| F4 |
C2 |
F5 |
107.439 |
|
F4 |
C2 |
H8 |
108.239 |
| F5 |
C2 |
H8 |
108.239 |
|
H6 |
C1 |
H7 |
110.034 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.721 |
|
|
|
| 2 |
C |
1.196 |
|
|
|
| 3 |
F |
-0.437 |
|
|
|
| 4 |
F |
-0.450 |
|
|
|
| 5 |
F |
-0.450 |
|
|
|
| 6 |
H |
-0.161 |
|
|
|
| 7 |
H |
-0.161 |
|
|
|
| 8 |
H |
-0.258 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.238 |
0.049 |
0.000 |
3.239 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.404 |
-0.359 |
0.000 |
| y |
-0.359 |
-29.043 |
0.000 |
| z |
0.000 |
0.000 |
-28.786 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.510 |
-0.359 |
0.000 |
| y |
-0.359 |
-1.948 |
0.000 |
| z |
0.000 |
0.000 |
-1.562 |
|
| Polar |
|---|
| 3z2-r2 | -3.124 |
|---|
| x2-y2 | 3.639 |
|---|
| xy | -0.359 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.412 |
-0.023 |
0.000 |
| y |
-0.023 |
4.487 |
0.000 |
| z |
0.000 |
0.000 |
4.354 |
<r2> (average value of r
2) Å
2
| <r2> |
103.725 |
| (<r2>)1/2 |
10.185 |
