Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -154.351872 |
| Energy at 298.15K | |
| HF Energy | -154.351872 |
| Nuclear repulsion energy | 74.661511 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3862 |
3701 |
19.24 |
|
|
|
| 2 |
A' |
3177 |
3044 |
8.46 |
|
|
|
| 3 |
A' |
3020 |
2894 |
58.69 |
|
|
|
| 4 |
A' |
1499 |
1437 |
0.04 |
|
|
|
| 5 |
A' |
1469 |
1408 |
5.37 |
|
|
|
| 6 |
A' |
1411 |
1352 |
7.83 |
|
|
|
| 7 |
A' |
1231 |
1180 |
40.30 |
|
|
|
| 8 |
A' |
1073 |
1028 |
83.24 |
|
|
|
| 9 |
A' |
1000 |
958 |
47.11 |
|
|
|
| 10 |
A' |
620 |
594 |
18.06 |
|
|
|
| 11 |
A' |
390 |
374 |
29.08 |
|
|
|
| 12 |
A" |
3293 |
3156 |
9.36 |
|
|
|
| 13 |
A" |
3064 |
2936 |
41.21 |
|
|
|
| 14 |
A" |
1276 |
1223 |
0.31 |
|
|
|
| 15 |
A" |
1160 |
1111 |
0.32 |
|
|
|
| 16 |
A" |
808 |
775 |
0.08 |
|
|
|
| 17 |
A" |
259 |
248 |
113.67 |
|
|
|
| 18 |
A" |
138i |
133i |
0.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14237.2 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 13643.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
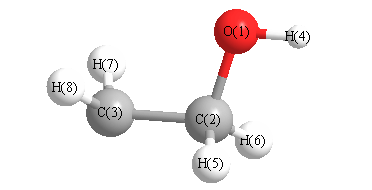
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.113 |
-0.371 |
0.000 |
| C2 |
0.000 |
0.535 |
0.000 |
| C3 |
1.257 |
-0.254 |
0.000 |
| H4 |
-1.922 |
0.150 |
0.000 |
| H5 |
-0.042 |
1.180 |
0.893 |
| H6 |
-0.042 |
1.180 |
-0.893 |
| H7 |
1.683 |
-0.616 |
-0.933 |
| H8 |
1.683 |
-0.616 |
0.933 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4350 | 2.3731 | 0.9619 | 2.0858 | 2.0858 | 2.9584 | 2.9584 |
C2 | 1.4350 | | 1.4843 | 1.9600 | 1.1023 | 1.1023 | 2.2428 | 2.2428 | C3 | 2.3731 | 1.4843 | | 3.2046 | 2.1310 | 2.1310 | 1.0881 | 1.0881 | H4 | 0.9619 | 1.9600 | 3.2046 | | 2.3224 | 2.3224 | 3.8021 | 3.8021 | H5 | 2.0858 | 1.1023 | 2.1310 | 2.3224 | | 1.7853 | 3.0883 | 2.4910 | H6 | 2.0858 | 1.1023 | 2.1310 | 2.3224 | 1.7853 | | 2.4910 | 3.0883 | H7 | 2.9584 | 2.2428 | 1.0881 | 3.8021 | 3.0883 | 2.4910 | | 1.8666 | H8 | 2.9584 | 2.2428 | 1.0881 | 3.8021 | 2.4910 | 3.0883 | 1.8666 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.748 |
|
O1 |
C2 |
H5 |
109.891 |
| O1 |
C2 |
H6 |
109.891 |
|
C2 |
O1 |
H4 |
108.089 |
| C2 |
C3 |
H7 |
120.581 |
|
C2 |
C3 |
H8 |
120.581 |
| C3 |
C2 |
H5 |
110.075 |
|
C3 |
C2 |
H6 |
110.075 |
| H5 |
C2 |
H6 |
108.155 |
|
H7 |
C3 |
H8 |
118.133 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.479 |
|
|
|
| 2 |
C |
0.816 |
|
|
|
| 3 |
C |
0.505 |
|
|
|
| 4 |
H |
0.101 |
|
|
|
| 5 |
H |
-0.207 |
|
|
|
| 6 |
H |
-0.207 |
|
|
|
| 7 |
H |
-0.265 |
|
|
|
| 8 |
H |
-0.265 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.111 |
1.520 |
0.000 |
1.524 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.224 |
-2.890 |
0.000 |
| y |
-2.890 |
-20.479 |
0.000 |
| z |
0.000 |
0.000 |
-19.032 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.531 |
-2.890 |
0.000 |
| y |
-2.890 |
-2.351 |
0.000 |
| z |
0.000 |
0.000 |
-0.181 |
|
| Polar |
|---|
| 3z2-r2 | -0.361 |
|---|
| x2-y2 | 3.255 |
|---|
| xy | -2.890 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.622 |
-0.192 |
0.000 |
| y |
-0.192 |
4.636 |
0.000 |
| z |
0.000 |
0.000 |
4.390 |
<r2> (average value of r
2) Å
2
| <r2> |
50.679 |
| (<r2>)1/2 |
7.119 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -154.353642 |
| Energy at 298.15K | -154.358178 |
| HF Energy | -154.353642 |
| Nuclear repulsion energy | 75.054133 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3896 |
3734 |
34.69 |
|
|
|
| 2 |
A |
3314 |
3175 |
3.81 |
|
|
|
| 3 |
A |
3190 |
3057 |
5.02 |
|
|
|
| 4 |
A |
2977 |
2853 |
49.96 |
|
|
|
| 5 |
A |
2941 |
2818 |
60.10 |
|
|
|
| 6 |
A |
1473 |
1411 |
11.34 |
|
|
|
| 7 |
A |
1457 |
1396 |
2.41 |
|
|
|
| 8 |
A |
1404 |
1346 |
1.47 |
|
|
|
| 9 |
A |
1269 |
1216 |
64.19 |
|
|
|
| 10 |
A |
1218 |
1167 |
1.40 |
|
|
|
| 11 |
A |
1140 |
1093 |
63.56 |
|
|
|
| 12 |
A |
1065 |
1021 |
9.62 |
|
|
|
| 13 |
A |
951 |
911 |
8.08 |
|
|
|
| 14 |
A |
883 |
846 |
18.34 |
|
|
|
| 15 |
A |
451 |
432 |
43.16 |
|
|
|
| 16 |
A |
422 |
404 |
25.02 |
|
|
|
| 17 |
A |
282 |
270 |
86.35 |
|
|
|
| 18 |
A |
66 |
63 |
22.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14198.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 13606.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.107 |
-0.378 |
-0.026 |
| C2 |
-0.009 |
0.520 |
0.018 |
| C3 |
1.250 |
-0.253 |
-0.012 |
| H4 |
-1.921 |
0.127 |
0.036 |
| H5 |
-0.056 |
1.142 |
0.936 |
| H6 |
-0.043 |
1.226 |
-0.834 |
| H7 |
2.203 |
0.261 |
-0.101 |
| H8 |
1.229 |
-1.330 |
0.134 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4189 | 2.3602 | 0.9598 | 2.0836 | 2.0878 | 3.3724 | 2.5271 |
C2 | 1.4189 | | 1.4780 | 1.9512 | 1.1099 | 1.1074 | 2.2310 | 2.2289 | C3 | 2.3602 | 1.4780 | | 3.1936 | 2.1330 | 2.1301 | 1.0872 | 1.0867 | H4 | 0.9598 | 1.9512 | 3.1936 | | 2.3061 | 2.3429 | 4.1285 | 3.4714 | H5 | 2.0836 | 1.1099 | 2.1330 | 2.3061 | | 1.7723 | 2.6373 | 2.8989 | H6 | 2.0878 | 1.1074 | 2.1301 | 2.3429 | 1.7723 | | 2.5526 | 3.0147 | H7 | 3.3724 | 2.2310 | 1.0872 | 4.1285 | 2.6373 | 2.5526 | | 1.8809 | H8 | 2.5271 | 2.2289 | 1.0867 | 3.4714 | 2.8989 | 3.0147 | 1.8809 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
109.106 |
|
O1 |
C2 |
H5 |
110.371 |
| O1 |
C2 |
H6 |
110.868 |
|
C2 |
O1 |
H4 |
108.693 |
| C2 |
C3 |
H7 |
120.089 |
|
C2 |
C3 |
H8 |
119.928 |
| C3 |
C2 |
H5 |
110.213 |
|
C3 |
C2 |
H6 |
110.127 |
| H5 |
C2 |
H6 |
106.130 |
|
H7 |
C3 |
H8 |
119.810 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.474 |
|
|
|
| 2 |
C |
0.904 |
|
|
|
| 3 |
C |
0.427 |
|
|
|
| 4 |
H |
0.112 |
|
|
|
| 5 |
H |
-0.185 |
|
|
|
| 6 |
H |
-0.194 |
|
|
|
| 7 |
H |
-0.310 |
|
|
|
| 8 |
H |
-0.279 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.265 |
1.686 |
0.201 |
1.719 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.454 |
-1.841 |
-0.251 |
| y |
-1.841 |
-19.435 |
-0.245 |
| z |
-0.251 |
-0.245 |
-20.752 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.639 |
-1.841 |
-0.251 |
| y |
-1.841 |
-0.831 |
-0.245 |
| z |
-0.251 |
-0.245 |
-2.808 |
|
| Polar |
|---|
| 3z2-r2 | -5.615 |
|---|
| x2-y2 | 2.980 |
|---|
| xy | -1.841 |
|---|
| xz | -0.251 |
|---|
| yz | -0.245 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.492 |
-0.203 |
-0.041 |
| y |
-0.203 |
4.830 |
0.024 |
| z |
-0.041 |
0.024 |
4.301 |
<r2> (average value of r
2) Å
2
| <r2> |
50.449 |
| (<r2>)1/2 |
7.103 |
