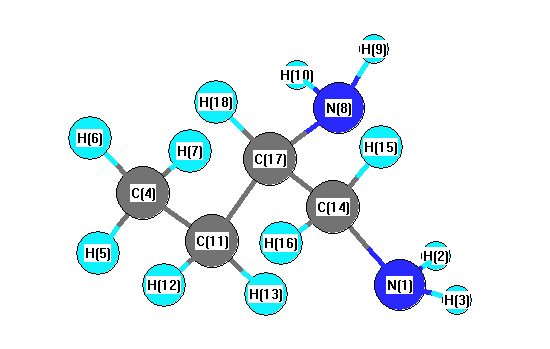
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3623 |
3472 |
5.59 |
|
|
|
| 2 |
A |
3620 |
3469 |
1.94 |
|
|
|
| 3 |
A |
3534 |
3386 |
0.62 |
|
|
|
| 4 |
A |
3530 |
3383 |
4.00 |
|
|
|
| 5 |
A |
3138 |
3007 |
41.94 |
|
|
|
| 6 |
A |
3133 |
3002 |
37.79 |
|
|
|
| 7 |
A |
3111 |
2981 |
9.09 |
|
|
|
| 8 |
A |
3082 |
2953 |
42.50 |
|
|
|
| 9 |
A |
3069 |
2941 |
12.37 |
|
|
|
| 10 |
A |
3046 |
2919 |
32.00 |
|
|
|
| 11 |
A |
2967 |
2844 |
130.57 |
|
|
|
| 12 |
A |
2951 |
2828 |
45.28 |
|
|
|
| 13 |
A |
1641 |
1573 |
40.83 |
|
|
|
| 14 |
A |
1625 |
1558 |
43.86 |
|
|
|
| 15 |
A |
1495 |
1433 |
6.36 |
|
|
|
| 16 |
A |
1485 |
1423 |
8.52 |
|
|
|
| 17 |
A |
1478 |
1416 |
5.35 |
|
|
|
| 18 |
A |
1462 |
1401 |
1.82 |
|
|
|
| 19 |
A |
1432 |
1372 |
0.46 |
|
|
|
| 20 |
A |
1417 |
1358 |
8.98 |
|
|
|
| 21 |
A |
1395 |
1337 |
3.95 |
|
|
|
| 22 |
A |
1378 |
1321 |
13.49 |
|
|
|
| 23 |
A |
1334 |
1278 |
1.62 |
|
|
|
| 24 |
A |
1296 |
1242 |
1.01 |
|
|
|
| 25 |
A |
1281 |
1227 |
2.35 |
|
|
|
| 26 |
A |
1257 |
1205 |
1.65 |
|
|
|
| 27 |
A |
1191 |
1141 |
3.51 |
|
|
|
| 28 |
A |
1160 |
1112 |
5.46 |
|
|
|
| 29 |
A |
1113 |
1067 |
6.62 |
|
|
|
| 30 |
A |
1079 |
1034 |
2.33 |
|
|
|
| 31 |
A |
1068 |
1024 |
4.05 |
|
|
|
| 32 |
A |
1013 |
971 |
3.36 |
|
|
|
| 33 |
A |
982 |
941 |
17.76 |
|
|
|
| 34 |
A |
933 |
894 |
14.65 |
|
|
|
| 35 |
A |
896 |
859 |
50.97 |
|
|
|
| 36 |
A |
845 |
810 |
115.24 |
|
|
|
| 37 |
A |
817 |
783 |
38.18 |
|
|
|
| 38 |
A |
763 |
731 |
1.54 |
|
|
|
| 39 |
A |
623 |
597 |
18.16 |
|
|
|
| 40 |
A |
443 |
424 |
4.69 |
|
|
|
| 41 |
A |
378 |
362 |
2.51 |
|
|
|
| 42 |
A |
332 |
319 |
23.09 |
|
|
|
| 43 |
A |
264 |
253 |
47.76 |
|
|
|
| 44 |
A |
261 |
250 |
14.94 |
|
|
|
| 45 |
A |
234 |
224 |
5.71 |
|
|
|
| 46 |
A |
214 |
205 |
3.57 |
|
|
|
| 47 |
A |
140 |
134 |
3.96 |
|
|
|
| 48 |
A |
84 |
80 |
0.67 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36806.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 35271.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.160 |
|
|
|
| 2 |
H |
0.046 |
|
|
|
| 3 |
H |
-0.104 |
|
|
|
| 4 |
C |
0.475 |
|
|
|
| 5 |
H |
-0.135 |
|
|
|
| 6 |
H |
-0.131 |
|
|
|
| 7 |
H |
-0.161 |
|
|
|
| 8 |
N |
-0.128 |
|
|
|
| 9 |
H |
-0.032 |
|
|
|
| 10 |
H |
-0.070 |
|
|
|
| 11 |
C |
0.755 |
|
|
|
| 12 |
H |
-0.266 |
|
|
|
| 13 |
H |
-0.260 |
|
|
|
| 14 |
C |
1.034 |
|
|
|
| 15 |
H |
-0.233 |
|
|
|
| 16 |
H |
-0.301 |
|
|
|
| 17 |
C |
0.024 |
|
|
|
| 18 |
H |
-0.353 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.015 |
1.181 |
1.267 |
1.732 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.510 |
0.373 |
1.055 |
| y |
0.373 |
-39.287 |
-0.433 |
| z |
1.055 |
-0.433 |
-41.391 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.829 |
0.373 |
1.055 |
| y |
0.373 |
-0.337 |
-0.433 |
| z |
1.055 |
-0.433 |
-3.492 |
|
| Polar |
|---|
| 3z2-r2 | -6.984 |
|---|
| x2-y2 | 2.777 |
|---|
| xy | 0.373 |
|---|
| xz | 1.055 |
|---|
| yz | -0.433 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.610 |
0.031 |
0.004 |
| y |
0.031 |
10.130 |
0.069 |
| z |
0.004 |
0.069 |
9.762 |
<r2> (average value of r
2) Å
2
| <r2> |
206.187 |
| (<r2>)1/2 |
14.359 |