Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -218.349443 |
| Energy at 298.15K | -218.357256 |
| HF Energy | -218.349443 |
| Nuclear repulsion energy | 127.851679 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3148 |
3017 |
26.38 |
|
|
|
| 2 |
A' |
3073 |
2945 |
41.69 |
|
|
|
| 3 |
A' |
3062 |
2934 |
32.20 |
|
|
|
| 4 |
A' |
3055 |
2928 |
11.85 |
|
|
|
| 5 |
A' |
1496 |
1434 |
6.46 |
|
|
|
| 6 |
A' |
1488 |
1426 |
2.72 |
|
|
|
| 7 |
A' |
1473 |
1412 |
1.10 |
|
|
|
| 8 |
A' |
1414 |
1355 |
10.91 |
|
|
|
| 9 |
A' |
1398 |
1339 |
2.83 |
|
|
|
| 10 |
A' |
1316 |
1262 |
0.81 |
|
|
|
| 11 |
A' |
1142 |
1095 |
1.20 |
|
|
|
| 12 |
A' |
1074 |
1029 |
53.23 |
|
|
|
| 13 |
A' |
1049 |
1006 |
74.20 |
|
|
|
| 14 |
A' |
906 |
868 |
8.01 |
|
|
|
| 15 |
A' |
451 |
433 |
6.75 |
|
|
|
| 16 |
A' |
268 |
257 |
3.38 |
|
|
|
| 17 |
A" |
3141 |
3010 |
73.45 |
|
|
|
| 18 |
A" |
3122 |
2992 |
2.22 |
|
|
|
| 19 |
A" |
3104 |
2975 |
4.10 |
|
|
|
| 20 |
A" |
1479 |
1417 |
7.25 |
|
|
|
| 21 |
A" |
1307 |
1253 |
0.03 |
|
|
|
| 22 |
A" |
1259 |
1207 |
0.56 |
|
|
|
| 23 |
A" |
1180 |
1130 |
0.97 |
|
|
|
| 24 |
A" |
888 |
851 |
1.10 |
|
|
|
| 25 |
A" |
759 |
727 |
2.28 |
|
|
|
| 26 |
A" |
227 |
218 |
0.00 |
|
|
|
| 27 |
A" |
122 |
117 |
3.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21200.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 20316.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.120 |
-0.778 |
0.000 |
| C2 |
0.000 |
0.727 |
0.000 |
| C3 |
-1.457 |
1.176 |
0.000 |
| F4 |
1.464 |
-1.160 |
0.000 |
| H5 |
-0.342 |
-1.222 |
0.894 |
| H6 |
-0.342 |
-1.222 |
-0.894 |
| H7 |
0.520 |
1.123 |
-0.883 |
| H8 |
0.520 |
1.123 |
0.883 |
| H9 |
-1.529 |
2.270 |
0.000 |
| H10 |
-1.991 |
0.809 |
-0.887 |
| H11 |
-1.991 |
0.809 |
0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5107 | 2.5112 | 1.3977 | 1.0994 | 1.0994 | 2.1341 | 2.1341 | 3.4654 | 2.7856 | 2.7856 |
C2 | 1.5107 | | 1.5246 | 2.3888 | 2.1714 | 2.1714 | 1.0984 | 1.0984 | 2.1715 | 2.1809 | 2.1809 | C3 | 2.5112 | 1.5246 | | 3.7404 | 2.7914 | 2.7914 | 2.1662 | 2.1662 | 1.0962 | 1.0983 | 1.0983 | F4 | 1.3977 | 2.3888 | 3.7404 | | 2.0162 | 2.0162 | 2.6233 | 2.6233 | 4.5520 | 4.0742 | 4.0742 | H5 | 1.0994 | 2.1714 | 2.7914 | 2.0162 | | 1.7882 | 3.0654 | 2.4978 | 3.7945 | 3.1645 | 2.6156 | H6 | 1.0994 | 2.1714 | 2.7914 | 2.0162 | 1.7882 | | 2.4978 | 3.0654 | 3.7945 | 2.6156 | 3.1645 | H7 | 2.1341 | 1.0984 | 2.1662 | 2.6233 | 3.0654 | 2.4978 | | 1.7660 | 2.5086 | 2.5304 | 3.0880 | H8 | 2.1341 | 1.0984 | 2.1662 | 2.6233 | 2.4978 | 3.0654 | 1.7660 | | 2.5086 | 3.0880 | 2.5304 | H9 | 3.4654 | 2.1715 | 1.0962 | 4.5520 | 3.7945 | 3.7945 | 2.5086 | 2.5086 | | 1.7706 | 1.7706 | H10 | 2.7856 | 2.1809 | 1.0983 | 4.0742 | 3.1645 | 2.6156 | 2.5304 | 3.0880 | 1.7706 | | 1.7740 | H11 | 2.7856 | 2.1809 | 1.0983 | 4.0742 | 2.6156 | 3.1645 | 3.0880 | 2.5304 | 1.7706 | 1.7740 | |
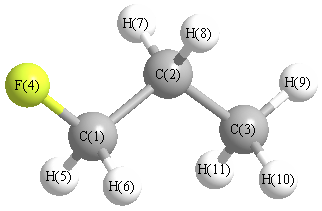 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.651 |
|
C1 |
C2 |
H7 |
108.735 |
| C1 |
C2 |
H8 |
108.735 |
|
C2 |
C1 |
F4 |
110.381 |
| C2 |
C1 |
H5 |
111.626 |
|
C2 |
C1 |
H6 |
111.626 |
| C2 |
C3 |
H9 |
110.845 |
|
C2 |
C3 |
H10 |
111.467 |
| C2 |
C3 |
H11 |
111.467 |
|
C3 |
C2 |
H7 |
110.290 |
| C3 |
C2 |
H8 |
110.290 |
|
F4 |
C1 |
H5 |
107.084 |
| F4 |
C1 |
H6 |
107.084 |
|
H5 |
C1 |
H6 |
108.831 |
| H7 |
C2 |
H8 |
107.008 |
|
H9 |
C3 |
H10 |
107.572 |
| H9 |
C3 |
H11 |
107.572 |
|
H10 |
C3 |
H11 |
107.729 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.915 |
|
|
|
| 2 |
C |
0.307 |
|
|
|
| 3 |
C |
0.428 |
|
|
|
| 4 |
F |
-0.467 |
|
|
|
| 5 |
H |
-0.247 |
|
|
|
| 6 |
H |
-0.247 |
|
|
|
| 7 |
H |
-0.159 |
|
|
|
| 8 |
H |
-0.159 |
|
|
|
| 9 |
H |
-0.143 |
|
|
|
| 10 |
H |
-0.113 |
|
|
|
| 11 |
H |
-0.113 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.851 |
0.877 |
0.000 |
2.048 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.770 |
2.331 |
0.000 |
| y |
2.331 |
-26.039 |
0.000 |
| z |
0.000 |
0.000 |
-24.673 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.414 |
2.331 |
0.000 |
| y |
2.331 |
0.682 |
0.000 |
| z |
0.000 |
0.000 |
2.732 |
|
| Polar |
|---|
| 3z2-r2 | 5.465 |
|---|
| x2-y2 | -2.731 |
|---|
| xy | 2.331 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.432 |
-0.422 |
0.000 |
| y |
-0.422 |
6.250 |
0.000 |
| z |
0.000 |
0.000 |
5.536 |
<r2> (average value of r
2) Å
2
| <r2> |
103.622 |
| (<r2>)1/2 |
10.179 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -218.349659 |
| Energy at 298.15K | |
| HF Energy | -218.349659 |
| Nuclear repulsion energy | 130.036413 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3155 |
3023 |
22.04 |
|
|
|
| 2 |
A |
3140 |
3009 |
46.74 |
|
|
|
| 3 |
A |
3127 |
2997 |
34.66 |
|
|
|
| 4 |
A |
3105 |
2975 |
14.00 |
|
|
|
| 5 |
A |
3066 |
2938 |
34.59 |
|
|
|
| 6 |
A |
3060 |
2932 |
27.96 |
|
|
|
| 7 |
A |
3058 |
2931 |
15.91 |
|
|
|
| 8 |
A |
1492 |
1430 |
2.62 |
|
|
|
| 9 |
A |
1484 |
1422 |
9.61 |
|
|
|
| 10 |
A |
1473 |
1411 |
6.33 |
|
|
|
| 11 |
A |
1455 |
1394 |
3.11 |
|
|
|
| 12 |
A |
1407 |
1349 |
8.98 |
|
|
|
| 13 |
A |
1401 |
1343 |
4.02 |
|
|
|
| 14 |
A |
1363 |
1306 |
0.40 |
|
|
|
| 15 |
A |
1296 |
1242 |
1.63 |
|
|
|
| 16 |
A |
1264 |
1211 |
1.32 |
|
|
|
| 17 |
A |
1168 |
1119 |
1.24 |
|
|
|
| 18 |
A |
1123 |
1077 |
4.31 |
|
|
|
| 19 |
A |
1099 |
1053 |
56.06 |
|
|
|
| 20 |
A |
980 |
940 |
56.48 |
|
|
|
| 21 |
A |
914 |
876 |
3.04 |
|
|
|
| 22 |
A |
889 |
852 |
4.91 |
|
|
|
| 23 |
A |
761 |
730 |
0.81 |
|
|
|
| 24 |
A |
476 |
456 |
4.16 |
|
|
|
| 25 |
A |
314 |
301 |
1.26 |
|
|
|
| 26 |
A |
210 |
201 |
1.54 |
|
|
|
| 27 |
A |
137 |
132 |
2.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21208.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 20323.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.791 |
0.512 |
0.301 |
| C2 |
-0.594 |
0.644 |
-0.288 |
| C3 |
-1.539 |
-0.479 |
0.118 |
| F4 |
1.413 |
-0.650 |
-0.171 |
| H5 |
1.434 |
1.356 |
0.018 |
| H6 |
0.761 |
0.433 |
1.398 |
| H7 |
-0.506 |
0.695 |
-1.382 |
| H8 |
-0.993 |
1.616 |
0.041 |
| H9 |
-2.529 |
-0.341 |
-0.334 |
| H10 |
-1.153 |
-1.453 |
-0.203 |
| H11 |
-1.670 |
-0.511 |
1.208 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5106 | 2.5387 | 1.4001 | 1.0981 | 1.0995 | 2.1328 | 2.1139 | 3.4859 | 2.8096 | 2.8154 |
C2 | 1.5106 | | 1.5230 | 2.3908 | 2.1709 | 2.1726 | 1.0989 | 1.1007 | 2.1718 | 2.1719 | 2.1751 | C3 | 2.5387 | 1.5230 | | 2.9712 | 3.4953 | 2.7856 | 2.1670 | 2.1662 | 1.0967 | 1.0962 | 1.0985 | F4 | 1.4001 | 2.3908 | 2.9712 | | 2.0148 | 2.0145 | 2.6380 | 3.3118 | 3.9572 | 2.6886 | 3.3805 | H5 | 1.0981 | 2.1709 | 3.4953 | 2.0148 | | 1.7909 | 2.4822 | 2.4412 | 4.3253 | 3.8250 | 3.8130 | H6 | 1.0995 | 2.1726 | 2.7856 | 2.0145 | 1.7909 | | 3.0659 | 2.5132 | 3.7974 | 3.1278 | 2.6150 | H7 | 2.1328 | 1.0989 | 2.1670 | 2.6380 | 2.4822 | 3.0659 | | 1.7632 | 2.5026 | 2.5340 | 3.0854 | H8 | 2.1139 | 1.1007 | 2.1662 | 3.3118 | 2.4412 | 2.5132 | 1.7632 | | 2.5157 | 3.0826 | 2.5190 | H9 | 3.4859 | 2.1718 | 1.0967 | 3.9572 | 4.3253 | 3.7974 | 2.5026 | 2.5157 | | 1.7738 | 1.7734 | H10 | 2.8096 | 2.1719 | 1.0962 | 2.6886 | 3.8250 | 3.1278 | 2.5340 | 3.0826 | 1.7738 | | 1.7744 | H11 | 2.8154 | 2.1751 | 1.0985 | 3.3805 | 3.8130 | 2.6150 | 3.0854 | 2.5190 | 1.7734 | 1.7744 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.616 |
|
C1 |
C2 |
H7 |
108.611 |
| C1 |
C2 |
H8 |
107.052 |
|
C2 |
C1 |
F4 |
110.387 |
| C2 |
C1 |
H5 |
111.672 |
|
C2 |
C1 |
H6 |
111.726 |
| C2 |
C3 |
H9 |
110.950 |
|
C2 |
C3 |
H10 |
110.987 |
| C2 |
C3 |
H11 |
111.105 |
|
C3 |
C2 |
H7 |
110.438 |
| C3 |
C2 |
H8 |
110.264 |
|
F4 |
C1 |
H5 |
106.885 |
| F4 |
C1 |
H6 |
106.782 |
|
H5 |
C1 |
H6 |
109.157 |
| H7 |
C2 |
H8 |
106.564 |
|
H9 |
C3 |
H10 |
107.981 |
| H9 |
C3 |
H11 |
107.776 |
|
H10 |
C3 |
H11 |
107.898 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.903 |
|
|
|
| 2 |
C |
0.347 |
|
|
|
| 3 |
C |
0.401 |
|
|
|
| 4 |
F |
-0.473 |
|
|
|
| 5 |
H |
-0.215 |
|
|
|
| 6 |
H |
-0.231 |
|
|
|
| 7 |
H |
-0.221 |
|
|
|
| 8 |
H |
-0.153 |
|
|
|
| 9 |
H |
-0.132 |
|
|
|
| 10 |
H |
-0.094 |
|
|
|
| 11 |
H |
-0.131 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.097 |
1.508 |
0.479 |
1.926 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.269 |
1.820 |
0.731 |
| y |
1.820 |
-25.621 |
-0.036 |
| z |
0.731 |
-0.036 |
-24.889 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.014 |
1.820 |
0.731 |
| y |
1.820 |
0.457 |
-0.036 |
| z |
0.731 |
-0.036 |
1.556 |
|
| Polar |
|---|
| 3z2-r2 | 3.113 |
|---|
| x2-y2 | -1.648 |
|---|
| xy | 1.820 |
|---|
| xz | 0.731 |
|---|
| yz | -0.036 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.485 |
0.233 |
0.055 |
| y |
0.233 |
6.022 |
-0.030 |
| z |
0.055 |
-0.030 |
5.704 |
<r2> (average value of r
2) Å
2
| <r2> |
92.353 |
| (<r2>)1/2 |
9.610 |
