Jump to
S1C2
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -218.279948 |
| Energy at 298.15K | -218.287670 |
| HF Energy | -218.279948 |
| Nuclear repulsion energy | 126.789297 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3066 |
3066 |
34.99 |
|
|
|
| 2 |
A' |
2994 |
2994 |
47.29 |
|
|
|
| 3 |
A' |
2980 |
2980 |
45.04 |
|
|
|
| 4 |
A' |
2975 |
2975 |
13.12 |
|
|
|
| 5 |
A' |
1463 |
1463 |
4.98 |
|
|
|
| 6 |
A' |
1454 |
1454 |
1.01 |
|
|
|
| 7 |
A' |
1442 |
1442 |
0.82 |
|
|
|
| 8 |
A' |
1373 |
1373 |
10.57 |
|
|
|
| 9 |
A' |
1364 |
1364 |
0.72 |
|
|
|
| 10 |
A' |
1285 |
1285 |
0.93 |
|
|
|
| 11 |
A' |
1107 |
1107 |
1.19 |
|
|
|
| 12 |
A' |
1020 |
1020 |
13.17 |
|
|
|
| 13 |
A' |
978 |
978 |
118.30 |
|
|
|
| 14 |
A' |
879 |
879 |
8.54 |
|
|
|
| 15 |
A' |
438 |
438 |
6.92 |
|
|
|
| 16 |
A' |
267 |
267 |
3.44 |
|
|
|
| 17 |
A" |
3060 |
3060 |
93.34 |
|
|
|
| 18 |
A" |
3039 |
3039 |
2.32 |
|
|
|
| 19 |
A" |
3021 |
3021 |
7.26 |
|
|
|
| 20 |
A" |
1450 |
1450 |
5.92 |
|
|
|
| 21 |
A" |
1277 |
1277 |
0.07 |
|
|
|
| 22 |
A" |
1227 |
1227 |
0.22 |
|
|
|
| 23 |
A" |
1148 |
1148 |
0.96 |
|
|
|
| 24 |
A" |
872 |
872 |
0.97 |
|
|
|
| 25 |
A" |
746 |
746 |
1.66 |
|
|
|
| 26 |
A" |
222 |
222 |
0.01 |
|
|
|
| 27 |
A" |
115 |
115 |
3.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20630.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 20630.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.116 |
-0.780 |
0.000 |
| C2 |
0.000 |
0.736 |
0.000 |
| C3 |
-1.470 |
1.184 |
0.000 |
| F4 |
1.480 |
-1.174 |
0.000 |
| H5 |
-0.344 |
-1.225 |
0.900 |
| H6 |
-0.344 |
-1.225 |
-0.900 |
| H7 |
0.519 |
1.134 |
-0.888 |
| H8 |
0.519 |
1.134 |
0.888 |
| H9 |
-1.543 |
2.282 |
0.000 |
| H10 |
-2.002 |
0.811 |
-0.892 |
| H11 |
-2.002 |
0.811 |
0.892 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5209 | 2.5243 | 1.4189 | 1.1046 | 1.1046 | 2.1485 | 2.1485 | 3.4830 | 2.7959 | 2.7959 |
C2 | 1.5209 | | 1.5362 | 2.4162 | 2.1853 | 2.1853 | 1.1031 | 1.1031 | 2.1841 | 2.1932 | 2.1932 | C3 | 2.5243 | 1.5362 | | 3.7757 | 2.8069 | 2.8069 | 2.1789 | 2.1789 | 1.1012 | 1.1033 | 1.1033 | F4 | 1.4189 | 2.4162 | 3.7757 | | 2.0344 | 2.0344 | 2.6530 | 2.6530 | 4.5913 | 4.1061 | 4.1061 | H5 | 1.1046 | 2.1853 | 2.8069 | 2.0344 | | 1.8001 | 3.0838 | 2.5125 | 3.8143 | 3.1791 | 2.6262 | H6 | 1.1046 | 2.1853 | 2.8069 | 2.0344 | 1.8001 | | 2.5125 | 3.0838 | 3.8143 | 2.6262 | 3.1791 | H7 | 2.1485 | 1.1031 | 2.1789 | 2.6530 | 3.0838 | 2.5125 | | 1.7762 | 2.5217 | 2.5423 | 3.1033 | H8 | 2.1485 | 1.1031 | 2.1789 | 2.6530 | 2.5125 | 3.0838 | 1.7762 | | 2.5217 | 3.1033 | 2.5423 | H9 | 3.4830 | 2.1841 | 1.1012 | 4.5913 | 3.8143 | 3.8143 | 2.5217 | 2.5217 | | 1.7804 | 1.7804 | H10 | 2.7959 | 2.1932 | 1.1033 | 4.1061 | 3.1791 | 2.6262 | 2.5423 | 3.1033 | 1.7804 | | 1.7831 | H11 | 2.7959 | 2.1932 | 1.1033 | 4.1061 | 2.6262 | 3.1791 | 3.1033 | 2.5423 | 1.7804 | 1.7831 | |
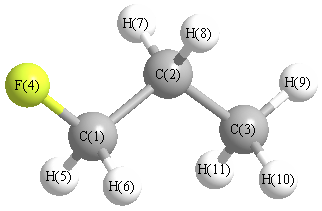 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.319 |
|
C1 |
C2 |
H7 |
108.885 |
| C1 |
C2 |
H8 |
108.885 |
|
C2 |
C1 |
F4 |
110.503 |
| C2 |
C1 |
H5 |
111.700 |
|
C2 |
C1 |
H6 |
111.700 |
| C2 |
C3 |
H9 |
110.733 |
|
C2 |
C3 |
H10 |
111.327 |
| C2 |
C3 |
H11 |
111.327 |
|
C3 |
C2 |
H7 |
110.205 |
| C3 |
C2 |
H8 |
110.205 |
|
F4 |
C1 |
H5 |
106.784 |
| F4 |
C1 |
H6 |
106.784 |
|
H5 |
C1 |
H6 |
109.132 |
| H7 |
C2 |
H8 |
107.238 |
|
H9 |
C3 |
H10 |
107.735 |
| H9 |
C3 |
H11 |
107.735 |
|
H10 |
C3 |
H11 |
107.822 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.027 |
|
|
|
| 2 |
C |
0.555 |
|
|
|
| 3 |
C |
0.635 |
|
|
|
| 4 |
F |
-0.453 |
|
|
|
| 5 |
H |
-0.328 |
|
|
|
| 6 |
H |
-0.328 |
|
|
|
| 7 |
H |
-0.251 |
|
|
|
| 8 |
H |
-0.251 |
|
|
|
| 9 |
H |
-0.227 |
|
|
|
| 10 |
H |
-0.190 |
|
|
|
| 11 |
H |
-0.190 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.854 |
0.906 |
0.000 |
2.063 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.038 |
2.344 |
0.000 |
| y |
2.344 |
-26.322 |
0.000 |
| z |
0.000 |
0.000 |
-24.936 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.409 |
2.344 |
0.000 |
| y |
2.344 |
0.665 |
0.000 |
| z |
0.000 |
0.000 |
2.744 |
|
| Polar |
|---|
| 3z2-r2 | 5.488 |
|---|
| x2-y2 | -2.717 |
|---|
| xy | 2.344 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.792 |
-0.478 |
0.000 |
| y |
-0.478 |
6.587 |
0.000 |
| z |
0.000 |
0.000 |
5.785 |
<r2> (average value of r
2) Å
2
| <r2> |
105.237 |
| (<r2>)1/2 |
10.258 |
Jump to
S1C1
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -218.280166 |
| Energy at 298.15K | |
| HF Energy | -218.280166 |
| Nuclear repulsion energy | 128.897524 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3073 |
3073 |
29.57 |
|
|
|
| 2 |
A |
3057 |
3057 |
58.83 |
|
|
|
| 3 |
A |
3040 |
3040 |
47.09 |
|
|
|
| 4 |
A |
3020 |
3020 |
16.33 |
|
|
|
| 5 |
A |
2982 |
2982 |
20.38 |
|
|
|
| 6 |
A |
2980 |
2980 |
57.69 |
|
|
|
| 7 |
A |
2973 |
2973 |
20.04 |
|
|
|
| 8 |
A |
1457 |
1457 |
2.37 |
|
|
|
| 9 |
A |
1454 |
1454 |
7.19 |
|
|
|
| 10 |
A |
1442 |
1442 |
4.36 |
|
|
|
| 11 |
A |
1424 |
1424 |
2.41 |
|
|
|
| 12 |
A |
1372 |
1372 |
6.72 |
|
|
|
| 13 |
A |
1360 |
1360 |
5.18 |
|
|
|
| 14 |
A |
1331 |
1331 |
1.33 |
|
|
|
| 15 |
A |
1264 |
1264 |
0.96 |
|
|
|
| 16 |
A |
1229 |
1229 |
0.75 |
|
|
|
| 17 |
A |
1136 |
1136 |
1.25 |
|
|
|
| 18 |
A |
1088 |
1088 |
3.15 |
|
|
|
| 19 |
A |
1044 |
1044 |
35.68 |
|
|
|
| 20 |
A |
933 |
933 |
71.81 |
|
|
|
| 21 |
A |
894 |
894 |
8.09 |
|
|
|
| 22 |
A |
853 |
853 |
6.61 |
|
|
|
| 23 |
A |
743 |
743 |
1.29 |
|
|
|
| 24 |
A |
465 |
465 |
4.14 |
|
|
|
| 25 |
A |
311 |
311 |
1.28 |
|
|
|
| 26 |
A |
203 |
203 |
1.56 |
|
|
|
| 27 |
A |
131 |
131 |
2.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20629.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 20629.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.790 |
0.520 |
0.307 |
| C2 |
-0.602 |
0.643 |
-0.294 |
| C3 |
-1.555 |
-0.485 |
0.120 |
| F4 |
1.437 |
-0.652 |
-0.171 |
| H5 |
1.433 |
1.372 |
0.027 |
| H6 |
0.753 |
0.432 |
1.407 |
| H7 |
-0.510 |
0.689 |
-1.393 |
| H8 |
-1.004 |
1.621 |
0.031 |
| H9 |
-2.552 |
-0.339 |
-0.327 |
| H10 |
-1.173 |
-1.464 |
-0.208 |
| H11 |
-1.676 |
-0.516 |
1.216 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5204 | 2.5579 | 1.4213 | 1.1035 | 1.1049 | 2.1466 | 2.1224 | 3.5078 | 2.8380 | 2.8248 |
C2 | 1.5204 | | 1.5338 | 2.4178 | 2.1848 | 2.1855 | 1.1037 | 1.1062 | 2.1838 | 2.1848 | 2.1860 | C3 | 2.5579 | 1.5338 | | 3.0107 | 3.5191 | 2.7977 | 2.1813 | 2.1782 | 1.1017 | 1.1009 | 1.1036 | F4 | 1.4213 | 2.4178 | 3.0107 | | 2.0331 | 2.0328 | 2.6609 | 3.3409 | 4.0038 | 2.7339 | 3.4107 | H5 | 1.1035 | 2.1848 | 3.5191 | 2.0331 | | 1.8034 | 2.5018 | 2.4496 | 4.3509 | 3.8587 | 3.8267 | H6 | 1.1049 | 2.1855 | 2.7977 | 2.0328 | 1.8034 | | 3.0833 | 2.5293 | 3.8112 | 3.1487 | 2.6146 | H7 | 2.1466 | 1.1037 | 2.1813 | 2.6609 | 2.5018 | 3.0833 | | 1.7722 | 2.5223 | 2.5455 | 3.1015 | H8 | 2.1224 | 1.1062 | 2.1782 | 3.3409 | 2.4496 | 2.5293 | 1.7722 | | 2.5227 | 3.0985 | 2.5340 | H9 | 3.5078 | 2.1838 | 1.1017 | 4.0038 | 4.3509 | 3.8112 | 2.5223 | 2.5227 | | 1.7832 | 1.7829 | H10 | 2.8380 | 2.1848 | 1.1009 | 2.7339 | 3.8587 | 3.1487 | 2.5455 | 3.0985 | 1.7832 | | 1.7831 | H11 | 2.8248 | 2.1860 | 1.1036 | 3.4107 | 3.8267 | 2.6146 | 3.1015 | 2.5340 | 1.7829 | 1.7831 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.752 |
|
C1 |
C2 |
H7 |
108.736 |
| C1 |
C2 |
H8 |
106.746 |
|
C2 |
C1 |
F4 |
110.507 |
| C2 |
C1 |
H5 |
111.761 |
|
C2 |
C1 |
H6 |
111.731 |
| C2 |
C3 |
H9 |
110.847 |
|
C2 |
C3 |
H10 |
110.982 |
| C2 |
C3 |
H11 |
110.913 |
|
C3 |
C2 |
H7 |
110.531 |
| C3 |
C2 |
H8 |
110.140 |
|
F4 |
C1 |
H5 |
106.595 |
| F4 |
C1 |
H6 |
106.496 |
|
H5 |
C1 |
H6 |
109.490 |
| H7 |
C2 |
H8 |
106.628 |
|
H9 |
C3 |
H10 |
108.111 |
| H9 |
C3 |
H11 |
107.894 |
|
H10 |
C3 |
H11 |
107.968 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.024 |
|
|
|
| 2 |
C |
0.593 |
|
|
|
| 3 |
C |
0.608 |
|
|
|
| 4 |
F |
-0.458 |
|
|
|
| 5 |
H |
-0.302 |
|
|
|
| 6 |
H |
-0.312 |
|
|
|
| 7 |
H |
-0.315 |
|
|
|
| 8 |
H |
-0.248 |
|
|
|
| 9 |
H |
-0.216 |
|
|
|
| 10 |
H |
-0.167 |
|
|
|
| 11 |
H |
-0.207 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.133 |
1.495 |
0.468 |
1.933 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.611 |
1.797 |
0.732 |
| y |
1.797 |
-25.848 |
-0.022 |
| z |
0.732 |
-0.022 |
-25.157 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.108 |
1.797 |
0.732 |
| y |
1.797 |
0.536 |
-0.022 |
| z |
0.732 |
-0.022 |
1.572 |
|
| Polar |
|---|
| 3z2-r2 | 3.145 |
|---|
| x2-y2 | -1.763 |
|---|
| xy | 1.797 |
|---|
| xz | 0.732 |
|---|
| yz | -0.022 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.847 |
0.236 |
0.061 |
| y |
0.236 |
6.333 |
-0.029 |
| z |
0.061 |
-0.029 |
5.980 |
<r2> (average value of r
2) Å
2
| <r2> |
93.970 |
| (<r2>)1/2 |
9.694 |
