Jump to
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -243.871019 |
| Energy at 298.15K | |
| HF Energy | -243.871019 |
| Nuclear repulsion energy | 125.134152 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4140 |
3770 |
138.22 |
|
|
|
| 2 |
A' |
3991 |
3634 |
85.65 |
|
|
|
| 3 |
A' |
3845 |
3501 |
71.09 |
|
|
|
| 4 |
A' |
1958 |
1783 |
731.18 |
|
|
|
| 5 |
A' |
1754 |
1597 |
146.02 |
|
|
|
| 6 |
A' |
1554 |
1415 |
168.17 |
|
|
|
| 7 |
A' |
1354 |
1233 |
221.84 |
|
|
|
| 8 |
A' |
1175 |
1070 |
44.53 |
|
|
|
| 9 |
A' |
1050 |
956 |
23.90 |
|
|
|
| 10 |
A' |
642 |
585 |
45.15 |
|
|
|
| 11 |
A' |
530 |
483 |
6.79 |
|
|
|
| 12 |
A" |
877 |
799 |
50.85 |
|
|
|
| 13 |
A" |
620 |
564 |
64.55 |
|
|
|
| 14 |
A" |
538 |
489 |
69.57 |
|
|
|
| 15 |
A" |
122i |
111i |
245.19 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11952.6 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 10884.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.125 |
0.000 |
| O2 |
-0.032 |
1.319 |
0.000 |
| N3 |
1.117 |
-0.626 |
0.000 |
| O4 |
-1.098 |
-0.635 |
0.000 |
| H5 |
1.997 |
-0.165 |
0.000 |
| H6 |
1.066 |
-1.618 |
0.000 |
| H7 |
-1.846 |
-0.056 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.1950 | 1.3463 | 1.3350 | 2.0175 | 2.0425 | 1.8547 |
O2 | 1.1950 | | 2.2597 | 2.2259 | 2.5135 | 3.1354 | 2.2760 | N3 | 1.3463 | 2.2597 | | 2.2152 | 0.9928 | 0.9927 | 3.0178 | O4 | 1.3350 | 2.2259 | 2.2152 | | 3.1298 | 2.3764 | 0.9463 | H5 | 2.0175 | 2.5135 | 0.9928 | 3.1298 | | 1.7253 | 3.8441 | H6 | 2.0425 | 3.1354 | 0.9927 | 2.3764 | 1.7253 | | 3.3044 | H7 | 1.8547 | 2.2760 | 3.0178 | 0.9463 | 3.8441 | 3.3044 | |
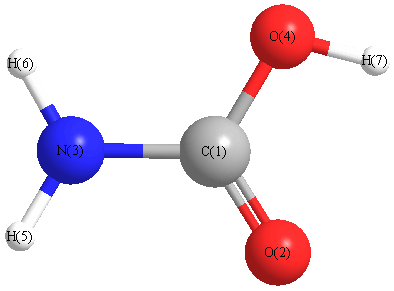 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.408 |
|
C1 |
N3 |
H6 |
120.926 |
| C1 |
O4 |
H7 |
107.562 |
|
O2 |
C1 |
N3 |
125.438 |
| O2 |
C1 |
O4 |
123.146 |
|
N3 |
C1 |
O4 |
111.416 |
| H5 |
N3 |
H6 |
120.666 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.264 |
|
|
|
| 2 |
O |
-0.637 |
|
|
|
| 3 |
N |
-0.139 |
|
|
|
| 4 |
O |
-0.562 |
|
|
|
| 5 |
H |
-0.033 |
|
|
|
| 6 |
H |
-0.031 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.658 |
-2.616 |
0.000 |
2.697 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.554 |
-2.584 |
0.000 |
| y |
-2.584 |
-26.770 |
0.000 |
| z |
0.000 |
0.000 |
-24.166 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.914 |
-2.584 |
0.000 |
| y |
-2.584 |
-6.910 |
0.000 |
| z |
0.000 |
0.000 |
-3.004 |
|
| Polar |
|---|
| 3z2-r2 | -6.008 |
|---|
| x2-y2 | 11.216 |
|---|
| xy | -2.584 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.414 |
-0.264 |
0.000 |
| y |
-0.264 |
4.546 |
0.000 |
| z |
0.000 |
0.000 |
2.992 |
<r2> (average value of r
2) Å
2
| <r2> |
63.397 |
| (<r2>)1/2 |
7.962 |
Jump to
S1C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -243.871029 |
| Energy at 298.15K | -243.875878 |
| HF Energy | -243.871029 |
| Nuclear repulsion energy | 125.113337 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4140 |
3770 |
138.05 |
|
|
|
| 2 |
A |
3980 |
3624 |
80.78 |
|
|
|
| 3 |
A |
3837 |
3494 |
66.11 |
|
|
|
| 4 |
A |
1960 |
1784 |
719.25 |
|
|
|
| 5 |
A |
1755 |
1598 |
140.53 |
|
|
|
| 6 |
A |
1553 |
1414 |
167.82 |
|
|
|
| 7 |
A |
1356 |
1235 |
212.00 |
|
|
|
| 8 |
A |
1181 |
1076 |
54.03 |
|
|
|
| 9 |
A |
1050 |
956 |
23.38 |
|
|
|
| 10 |
A |
877 |
799 |
55.78 |
|
|
|
| 11 |
A |
643 |
586 |
48.48 |
|
|
|
| 12 |
A |
613 |
558 |
74.28 |
|
|
|
| 13 |
A |
541 |
492 |
41.48 |
|
|
|
| 14 |
A |
523 |
476 |
22.84 |
|
|
|
| 15 |
A |
214 |
195 |
253.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12110.9 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 11028.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.040 |
0.118 |
-0.002 |
| O2 |
-0.447 |
1.241 |
0.004 |
| N3 |
1.260 |
-0.242 |
-0.032 |
| O4 |
-0.841 |
-0.949 |
0.002 |
| H5 |
1.939 |
0.474 |
0.083 |
| H6 |
1.517 |
-1.192 |
0.102 |
| H7 |
-1.734 |
-0.636 |
0.004 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.1946 | 1.3488 | 1.3347 | 2.0122 | 2.0372 | 1.8548 |
O2 | 1.1946 | | 2.2614 | 2.2255 | 2.5074 | 3.1284 | 2.2763 | N3 | 1.3488 | 2.2614 | | 2.2170 | 0.9936 | 0.9935 | 3.0200 | O4 | 1.3347 | 2.2255 | 2.2170 | | 3.1242 | 2.3724 | 0.9463 | H5 | 2.0122 | 2.5074 | 0.9936 | 3.1242 | | 1.7191 | 3.8380 | H6 | 2.0372 | 3.1284 | 0.9935 | 2.3724 | 1.7191 | | 3.2995 | H7 | 1.8548 | 2.2763 | 3.0200 | 0.9463 | 3.8380 | 3.2995 | |
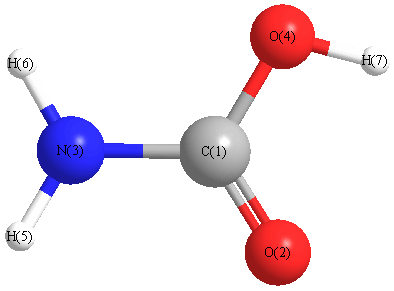 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.619 |
|
C1 |
N3 |
H6 |
120.101 |
| C1 |
O4 |
H7 |
107.592 |
|
O2 |
C1 |
N3 |
125.418 |
| O2 |
C1 |
O4 |
123.164 |
|
N3 |
C1 |
O4 |
111.413 |
| H5 |
N3 |
H6 |
119.795 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.265 |
|
|
|
| 2 |
O |
-0.634 |
|
|
|
| 3 |
N |
-0.159 |
|
|
|
| 4 |
O |
-0.561 |
|
|
|
| 5 |
H |
-0.026 |
|
|
|
| 6 |
H |
-0.025 |
|
|
|
| 7 |
H |
0.140 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.388 |
-2.249 |
0.396 |
2.672 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.263 |
1.322 |
0.784 |
| y |
1.322 |
-27.218 |
-0.228 |
| z |
0.784 |
-0.228 |
-24.124 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.408 |
1.322 |
0.784 |
| y |
1.322 |
-7.524 |
-0.228 |
| z |
0.784 |
-0.228 |
-2.883 |
|
| Polar |
|---|
| 3z2-r2 | -5.767 |
|---|
| x2-y2 | 11.955 |
|---|
| xy | 1.322 |
|---|
| xz | 0.784 |
|---|
| yz | -0.228 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.582 |
-0.248 |
-0.013 |
| y |
-0.248 |
4.378 |
0.007 |
| z |
-0.013 |
0.007 |
2.993 |
<r2> (average value of r
2) Å
2
| <r2> |
63.410 |
| (<r2>)1/2 |
7.963 |
