Jump to
S1C2
Energy calculated at CCSD=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -244.595633 |
| Energy at 298.15K | |
| HF Energy | -243.867873 |
| Nuclear repulsion energy | 123.011197 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3832 |
3832 |
87.56 |
|
|
|
| 2 |
A' |
3801 |
3801 |
68.85 |
|
|
|
| 3 |
A' |
3655 |
3655 |
62.31 |
|
|
|
| 4 |
A' |
1837 |
1837 |
590.24 |
|
|
|
| 5 |
A' |
1637 |
1637 |
114.51 |
|
|
|
| 6 |
A' |
1450 |
1450 |
130.57 |
|
|
|
| 7 |
A' |
1260 |
1260 |
209.01 |
|
|
|
| 8 |
A' |
1091 |
1091 |
33.98 |
|
|
|
| 9 |
A' |
964 |
964 |
41.34 |
|
|
|
| 10 |
A' |
589 |
589 |
34.30 |
|
|
|
| 11 |
A' |
491 |
491 |
5.76 |
|
|
|
| 12 |
A" |
789 |
789 |
21.67 |
|
|
|
| 13 |
A" |
587 |
587 |
57.92 |
|
|
|
| 14 |
A" |
486 |
486 |
63.02 |
|
|
|
| 15 |
A" |
224i |
224i |
218.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11121.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11121.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.131 |
0.000 |
| O2 |
-0.062 |
1.347 |
0.000 |
| N3 |
1.143 |
-0.610 |
0.000 |
| O4 |
-1.097 |
-0.683 |
0.000 |
| H5 |
2.026 |
-0.127 |
0.000 |
| H6 |
1.107 |
-1.617 |
0.000 |
| H7 |
-1.859 |
-0.087 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2178 | 1.3620 | 1.3660 | 2.0427 | 2.0686 | 1.8721 |
O2 | 1.2178 | | 2.2982 | 2.2786 | 2.5560 | 3.1859 | 2.2997 | N3 | 1.3620 | 2.2982 | | 2.2412 | 1.0071 | 1.0073 | 3.0473 | O4 | 1.3660 | 2.2786 | 2.2412 | | 3.1727 | 2.3938 | 0.9671 | H5 | 2.0427 | 2.5560 | 1.0071 | 3.1727 | | 1.7508 | 3.8859 | H6 | 2.0686 | 3.1859 | 1.0073 | 2.3938 | 1.7508 | | 3.3372 | H7 | 1.8721 | 2.2997 | 3.0473 | 0.9671 | 3.8859 | 3.3372 | |
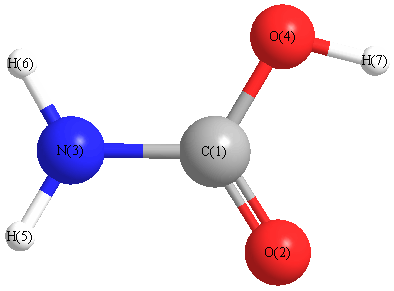 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.371 |
|
C1 |
N3 |
H6 |
120.911 |
| C1 |
O4 |
H7 |
105.446 |
|
O2 |
C1 |
N3 |
125.869 |
| O2 |
C1 |
O4 |
123.645 |
|
N3 |
C1 |
O4 |
110.486 |
| H5 |
N3 |
H6 |
120.718 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -244.595777 |
| Energy at 298.15K | -244.600570 |
| HF Energy | -243.867784 |
| Nuclear repulsion energy | 122.949920 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3831 |
3831 |
87.47 |
|
|
|
| 2 |
A |
3766 |
3766 |
57.85 |
|
|
|
| 3 |
A |
3630 |
3630 |
48.77 |
|
|
|
| 4 |
A |
1840 |
1840 |
559.07 |
|
|
|
| 5 |
A |
1638 |
1638 |
102.65 |
|
|
|
| 6 |
A |
1447 |
1447 |
129.20 |
|
|
|
| 7 |
A |
1264 |
1264 |
183.63 |
|
|
|
| 8 |
A |
1106 |
1106 |
60.44 |
|
|
|
| 9 |
A |
965 |
965 |
39.34 |
|
|
|
| 10 |
A |
787 |
787 |
30.72 |
|
|
|
| 11 |
A |
592 |
592 |
45.86 |
|
|
|
| 12 |
A |
572 |
572 |
72.74 |
|
|
|
| 13 |
A |
500 |
500 |
20.11 |
|
|
|
| 14 |
A |
469 |
469 |
25.58 |
|
|
|
| 15 |
A |
316 |
316 |
241.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11359.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11359.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.040 |
0.124 |
-0.002 |
| O2 |
-0.466 |
1.264 |
0.007 |
| N3 |
1.279 |
-0.237 |
-0.054 |
| O4 |
-0.840 |
-0.982 |
0.002 |
| H5 |
1.949 |
0.492 |
0.140 |
| H6 |
1.529 |
-1.189 |
0.171 |
| H7 |
-1.747 |
-0.647 |
0.006 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2168 | 1.3689 | 1.3651 | 2.0277 | 2.0536 | 1.8730 |
O2 | 1.2168 | | 2.3029 | 2.2772 | 2.5387 | 3.1663 | 2.3009 | N3 | 1.3689 | 2.3029 | | 2.2463 | 1.0094 | 1.0096 | 3.0541 | O4 | 1.3651 | 2.2772 | 2.2463 | | 3.1572 | 2.3832 | 0.9672 | H5 | 2.0277 | 2.5387 | 1.0094 | 3.1572 | | 1.7335 | 3.8697 | H6 | 2.0536 | 3.1663 | 1.0096 | 2.3832 | 1.7335 | | 3.3242 | H7 | 1.8730 | 2.3009 | 3.0541 | 0.9672 | 3.8697 | 3.3242 | |
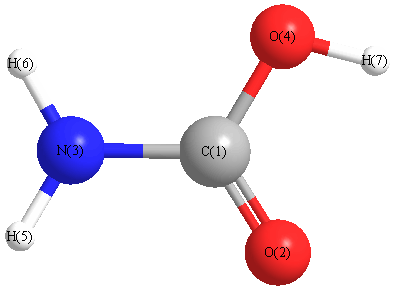 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.173 |
|
C1 |
N3 |
H6 |
118.618 |
| C1 |
O4 |
H7 |
105.580 |
|
O2 |
C1 |
N3 |
125.803 |
| O2 |
C1 |
O4 |
123.674 |
|
N3 |
C1 |
O4 |
110.499 |
| H5 |
N3 |
H6 |
118.325 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
