Jump to
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -356.611148 |
| Energy at 298.15K | -356.616432 |
| Nuclear repulsion energy | 234.889319 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4108 |
3741 |
138.90 |
|
|
|
| 2 |
A' |
3966 |
3612 |
100.17 |
|
|
|
| 3 |
A' |
3821 |
3480 |
81.29 |
|
|
|
| 4 |
A' |
1998 |
1820 |
17.87 |
|
|
|
| 5 |
A' |
1956 |
1781 |
913.36 |
|
|
|
| 6 |
A' |
1752 |
1596 |
115.01 |
|
|
|
| 7 |
A' |
1563 |
1423 |
17.31 |
|
|
|
| 8 |
A' |
1425 |
1298 |
73.53 |
|
|
|
| 9 |
A' |
1314 |
1196 |
375.48 |
|
|
|
| 10 |
A' |
1192 |
1086 |
6.25 |
|
|
|
| 11 |
A' |
855 |
778 |
9.39 |
|
|
|
| 12 |
A' |
667 |
607 |
94.39 |
|
|
|
| 13 |
A' |
574 |
522 |
0.02 |
|
|
|
| 14 |
A' |
448 |
408 |
2.82 |
|
|
|
| 15 |
A' |
293 |
267 |
17.62 |
|
|
|
| 16 |
A" |
935 |
852 |
4.75 |
|
|
|
| 17 |
A" |
698 |
635 |
153.78 |
|
|
|
| 18 |
A" |
643 |
586 |
32.97 |
|
|
|
| 19 |
A" |
481 |
438 |
2.63 |
|
|
|
| 20 |
A" |
327 |
298 |
230.82 |
|
|
|
| 21 |
A" |
55 |
50 |
3.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14536.3 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 13236.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.756 |
0.000 |
| C2 |
-0.054 |
-0.783 |
0.000 |
| O3 |
-1.087 |
-1.375 |
0.000 |
| O4 |
1.025 |
1.357 |
0.000 |
| O5 |
-1.191 |
1.306 |
0.000 |
| N6 |
1.171 |
-1.330 |
0.000 |
| H7 |
1.254 |
-2.322 |
0.000 |
| H8 |
1.982 |
-0.754 |
0.000 |
| H9 |
-1.091 |
2.249 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5394 | 2.3923 | 1.1881 | 1.3118 | 2.3927 | 3.3235 | 2.4914 | 1.8493 |
C2 | 1.5394 | | 1.1914 | 2.3957 | 2.3782 | 1.3420 | 2.0199 | 2.0356 | 3.2042 | O3 | 2.3923 | 1.1914 | | 3.4531 | 2.6832 | 2.2591 | 2.5256 | 3.1312 | 3.6242 | O4 | 1.1881 | 2.3957 | 3.4531 | | 2.2165 | 2.6910 | 3.6855 | 2.3173 | 2.2966 | O5 | 1.3118 | 2.3782 | 2.6832 | 2.2165 | | 3.5401 | 4.3750 | 3.7828 | 0.9483 | N6 | 2.3927 | 1.3420 | 2.2591 | 2.6910 | 3.5401 | | 0.9948 | 0.9945 | 4.2346 | H7 | 3.3235 | 2.0199 | 2.5256 | 3.6855 | 4.3750 | 0.9948 | | 1.7284 | 5.1375 | H8 | 2.4914 | 2.0356 | 3.1312 | 2.3173 | 3.7828 | 0.9945 | 1.7284 | | 4.2966 | H9 | 1.8493 | 3.2042 | 3.6242 | 2.2966 | 0.9483 | 4.2346 | 5.1375 | 4.2966 | |
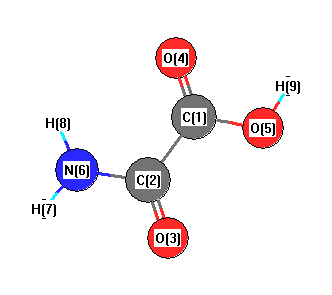 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
121.820 |
|
C1 |
C2 |
N6 |
112.100 |
| C1 |
O5 |
H9 |
108.754 |
|
C2 |
C1 |
O4 |
122.365 |
| C2 |
C1 |
O5 |
112.803 |
|
C2 |
N6 |
H7 |
118.874 |
| C2 |
N6 |
H8 |
120.477 |
|
O3 |
C2 |
N6 |
126.079 |
| O4 |
C1 |
O5 |
124.832 |
|
H7 |
N6 |
H8 |
120.649 |
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.088 |
|
|
|
| 2 |
C |
0.844 |
|
|
|
| 3 |
O |
-0.684 |
|
|
|
| 4 |
O |
-0.695 |
|
|
|
| 5 |
O |
-0.595 |
|
|
|
| 6 |
N |
-0.173 |
|
|
|
| 7 |
H |
0.004 |
|
|
|
| 8 |
H |
0.014 |
|
|
|
| 9 |
H |
0.197 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.156 |
0.678 |
0.000 |
2.260 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.856 |
-8.028 |
-0.002 |
| y |
-8.028 |
-40.528 |
0.002 |
| z |
-0.002 |
0.002 |
-33.760 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
12.288 |
-8.028 |
-0.002 |
| y |
-8.028 |
-11.220 |
0.002 |
| z |
-0.002 |
0.002 |
-1.069 |
|
| Polar |
|---|
| 3z2-r2 | -2.137 |
|---|
| x2-y2 | 15.672 |
|---|
| xy | -8.028 |
|---|
| xz | -0.002 |
|---|
| yz | 0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
140.186 |
| (<r2>)1/2 |
11.840 |
Jump to
S1C1
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -356.616438 |
| Energy at 298.15K | -356.621963 |
| Nuclear repulsion energy | 236.113100 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4030 |
3670 |
173.65 |
|
|
|
| 2 |
A' |
3956 |
3602 |
104.12 |
|
|
|
| 3 |
A' |
3812 |
3471 |
90.07 |
|
|
|
| 4 |
A' |
2031 |
1850 |
269.04 |
|
|
|
| 5 |
A' |
1932 |
1759 |
616.52 |
|
|
|
| 6 |
A' |
1756 |
1599 |
78.49 |
|
|
|
| 7 |
A' |
1583 |
1442 |
156.22 |
|
|
|
| 8 |
A' |
1449 |
1320 |
496.81 |
|
|
|
| 9 |
A' |
1315 |
1198 |
19.96 |
|
|
|
| 10 |
A' |
1196 |
1089 |
0.25 |
|
|
|
| 11 |
A' |
883 |
804 |
8.73 |
|
|
|
| 12 |
A' |
683 |
622 |
25.64 |
|
|
|
| 13 |
A' |
588 |
536 |
3.57 |
|
|
|
| 14 |
A' |
440 |
401 |
6.12 |
|
|
|
| 15 |
A' |
295 |
269 |
41.60 |
|
|
|
| 16 |
A" |
935 |
852 |
0.45 |
|
|
|
| 17 |
A" |
714 |
650 |
126.69 |
|
|
|
| 18 |
A" |
683 |
622 |
10.50 |
|
|
|
| 19 |
A" |
516 |
470 |
89.18 |
|
|
|
| 20 |
A" |
393 |
358 |
178.41 |
|
|
|
| 21 |
A" |
107 |
97 |
7.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14648.7 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 13339.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.003 |
-0.791 |
0.000 |
| C2 |
0.000 |
0.747 |
0.000 |
| O3 |
-1.051 |
1.329 |
0.000 |
| O4 |
1.003 |
-1.423 |
0.000 |
| O5 |
-1.202 |
-1.305 |
0.000 |
| N6 |
1.214 |
1.294 |
0.000 |
| H7 |
1.308 |
2.285 |
0.000 |
| H8 |
2.019 |
0.707 |
0.000 |
| H9 |
-1.839 |
-0.596 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
O4 |
O5 |
N6 |
H7 |
H8 |
H9 |
| C1 | | 1.5386 | 2.3678 | 1.1829 | 1.3097 | 2.4118 | 3.3415 | 2.5122 | 1.8517 |
C2 | 1.5386 | | 1.2011 | 2.3909 | 2.3783 | 1.3317 | 2.0183 | 2.0192 | 2.2774 | O3 | 2.3678 | 1.2011 | | 3.4340 | 2.6383 | 2.2654 | 2.5445 | 3.1319 | 2.0806 | O4 | 1.1829 | 2.3909 | 3.4340 | | 2.2082 | 2.7253 | 3.7205 | 2.3604 | 2.9593 | O5 | 1.3097 | 2.3783 | 2.6383 | 2.2082 | | 3.5487 | 4.3799 | 3.7979 | 0.9523 | N6 | 2.4118 | 1.3317 | 2.2654 | 2.7253 | 3.5487 | | 0.9953 | 0.9956 | 3.5909 | H7 | 3.3415 | 2.0183 | 2.5445 | 3.7205 | 4.3799 | 0.9953 | | 1.7304 | 4.2661 | H8 | 2.5122 | 2.0192 | 3.1319 | 2.3604 | 3.7979 | 0.9956 | 1.7304 | | 4.0719 | H9 | 1.8517 | 2.2774 | 2.0806 | 2.9593 | 0.9523 | 3.5909 | 4.2661 | 4.0719 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O3 |
119.080 |
|
C1 |
C2 |
N6 |
114.136 |
| C1 |
O5 |
H9 |
108.863 |
|
C2 |
C1 |
O4 |
122.382 |
| C2 |
C1 |
O5 |
112.979 |
|
C2 |
N6 |
H7 |
119.604 |
| C2 |
N6 |
H8 |
119.669 |
|
O3 |
C2 |
N6 |
126.784 |
| O4 |
C1 |
O5 |
124.639 |
|
H7 |
N6 |
H8 |
120.727 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.095 |
|
|
|
| 2 |
C |
0.817 |
|
|
|
| 3 |
O |
-0.744 |
|
|
|
| 4 |
O |
-0.635 |
|
|
|
| 5 |
O |
-0.606 |
|
|
|
| 6 |
N |
-0.142 |
|
|
|
| 7 |
H |
-0.007 |
|
|
|
| 8 |
H |
0.014 |
|
|
|
| 9 |
H |
0.208 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.357 |
2.462 |
0.000 |
2.812 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.102 |
7.814 |
0.000 |
| y |
7.814 |
-38.372 |
0.000 |
| z |
0.000 |
0.000 |
-33.714 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.941 |
7.814 |
0.000 |
| y |
7.814 |
-5.964 |
0.000 |
| z |
0.000 |
0.000 |
1.023 |
|
| Polar |
|---|
| 3z2-r2 | 2.046 |
|---|
| x2-y2 | 7.270 |
|---|
| xy | 7.814 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.192 |
-0.180 |
0.000 |
| y |
-0.180 |
6.096 |
0.000 |
| z |
0.000 |
0.000 |
3.962 |
<r2> (average value of r
2) Å
2
| <r2> |
138.273 |
| (<r2>)1/2 |
11.759 |
