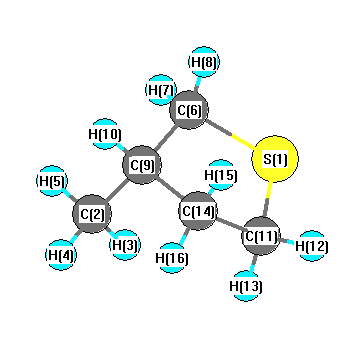
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3151 |
3020 |
14.09 |
|
|
|
| 2 |
A |
3146 |
3014 |
13.17 |
|
|
|
| 3 |
A |
3140 |
3009 |
28.27 |
|
|
|
| 4 |
A |
3137 |
3006 |
25.54 |
|
|
|
| 5 |
A |
3118 |
2988 |
24.11 |
|
|
|
| 6 |
A |
3090 |
2961 |
7.83 |
|
|
|
| 7 |
A |
3087 |
2958 |
61.34 |
|
|
|
| 8 |
A |
3070 |
2942 |
36.10 |
|
|
|
| 9 |
A |
3058 |
2931 |
6.54 |
|
|
|
| 10 |
A |
3055 |
2928 |
25.83 |
|
|
|
| 11 |
A |
1494 |
1431 |
3.39 |
|
|
|
| 12 |
A |
1492 |
1430 |
3.41 |
|
|
|
| 13 |
A |
1474 |
1413 |
6.47 |
|
|
|
| 14 |
A |
1470 |
1408 |
4.82 |
|
|
|
| 15 |
A |
1463 |
1402 |
4.12 |
|
|
|
| 16 |
A |
1401 |
1343 |
11.76 |
|
|
|
| 17 |
A |
1362 |
1305 |
3.50 |
|
|
|
| 18 |
A |
1337 |
1281 |
1.03 |
|
|
|
| 19 |
A |
1310 |
1255 |
2.51 |
|
|
|
| 20 |
A |
1287 |
1233 |
3.81 |
|
|
|
| 21 |
A |
1251 |
1199 |
7.82 |
|
|
|
| 22 |
A |
1229 |
1178 |
7.02 |
|
|
|
| 23 |
A |
1182 |
1133 |
1.32 |
|
|
|
| 24 |
A |
1128 |
1081 |
0.16 |
|
|
|
| 25 |
A |
1078 |
1033 |
0.34 |
|
|
|
| 26 |
A |
1065 |
1020 |
0.59 |
|
|
|
| 27 |
A |
1042 |
998 |
0.44 |
|
|
|
| 28 |
A |
1006 |
964 |
1.83 |
|
|
|
| 29 |
A |
963 |
923 |
0.30 |
|
|
|
| 30 |
A |
941 |
902 |
0.34 |
|
|
|
| 31 |
A |
899 |
862 |
2.32 |
|
|
|
| 32 |
A |
858 |
823 |
1.09 |
|
|
|
| 33 |
A |
814 |
780 |
1.49 |
|
|
|
| 34 |
A |
729 |
699 |
5.06 |
|
|
|
| 35 |
A |
701 |
672 |
2.25 |
|
|
|
| 36 |
A |
614 |
588 |
0.81 |
|
|
|
| 37 |
A |
496 |
476 |
0.13 |
|
|
|
| 38 |
A |
378 |
362 |
0.16 |
|
|
|
| 39 |
A |
354 |
339 |
0.16 |
|
|
|
| 40 |
A |
230 |
221 |
0.06 |
|
|
|
| 41 |
A |
189 |
181 |
0.20 |
|
|
|
| 42 |
A |
112 |
107 |
2.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31199.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 29898.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.323 |
|
|
|
| 2 |
C |
0.555 |
|
|
|
| 3 |
H |
-0.049 |
|
|
|
| 4 |
H |
-0.152 |
|
|
|
| 5 |
H |
-0.156 |
|
|
|
| 6 |
C |
0.373 |
|
|
|
| 7 |
H |
-0.190 |
|
|
|
| 8 |
H |
-0.228 |
|
|
|
| 9 |
C |
-0.197 |
|
|
|
| 10 |
H |
-0.352 |
|
|
|
| 11 |
C |
0.169 |
|
|
|
| 12 |
H |
-0.158 |
|
|
|
| 13 |
H |
-0.202 |
|
|
|
| 14 |
C |
0.801 |
|
|
|
| 15 |
H |
-0.286 |
|
|
|
| 16 |
H |
-0.250 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.911 |
0.813 |
0.732 |
2.202 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.085 |
-1.960 |
-0.079 |
| y |
-1.960 |
-43.602 |
-1.211 |
| z |
-0.079 |
-1.211 |
-46.132 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.218 |
-1.960 |
-0.079 |
| y |
-1.960 |
4.006 |
-1.211 |
| z |
-0.079 |
-1.211 |
0.212 |
|
| Polar |
|---|
| 3z2-r2 | 0.424 |
|---|
| x2-y2 | -5.483 |
|---|
| xy | -1.960 |
|---|
| xz | -0.079 |
|---|
| yz | -1.211 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.594 |
0.094 |
0.238 |
| y |
0.094 |
11.964 |
-0.140 |
| z |
0.238 |
-0.140 |
10.586 |
<r2> (average value of r
2) Å
2
| <r2> |
194.264 |
| (<r2>)1/2 |
13.938 |