return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSCH3 (Thioacetone)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-515.281485
Energy at 298.15K-515.287322
HF Energy-514.627567
Nuclear repulsion energy150.925846
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3177 3104        
2 A1 3048 2977        
3 A1 1538 1502        
4 A1 1445 1412        
5 A1 1332 1301        
6 A1 1036 1013        
7 A1 710 693        
8 A1 377 368        
9 A2 3108 3036        
10 A2 1513 1478        
11 A2 951 929        
12 A2 106 103        
13 B1 3113 3041        
14 B1 1531 1496        
15 B1 1103 1078        
16 B1 442 432        
17 B1 171 167        
18 B2 3176 3102        
19 B2 3043 2972        
20 B2 1511 1476        
21 B2 1434 1401        
22 B2 1245 1216        
23 B2 960 938        
24 B2 393 384        

Unscaled Zero Point Vibrational Energy (zpe) 18230.5 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 17809.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.28666 0.15941 0.10658

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -0.261
S2 0.000 0.000 1.394
C3 0.000 1.280 -1.078
C4 0.000 -1.280 -1.078
H5 0.000 2.170 -0.431
H6 0.000 -2.170 -0.431
H7 0.890 1.301 -1.737
H8 -0.890 1.301 -1.737
H9 -0.890 -1.301 -1.737
H10 0.890 -1.301 -1.737

Atom - Atom Distances (Å)
  C1 S2 C3 C4 H5 H6 H7 H8 H9 H10
C11.65511.51801.51802.17682.17682.15922.15922.15922.1592
S21.65512.78332.78332.83552.83553.50533.50533.50533.5053
C31.51802.78332.55931.10063.50991.10801.10802.80792.8079
C41.51802.78332.55933.50991.10062.80792.80791.10801.1080
H52.17682.83551.10063.50994.34041.80411.80413.81373.8137
H62.17682.83553.50991.10064.34043.81373.81371.80411.8041
H72.15923.50531.10802.80791.80413.81371.78033.15202.6010
H82.15923.50531.10802.80791.80413.81371.78032.60103.1520
H92.15923.50532.80791.10803.81371.80413.15202.60101.7803
H102.15923.50532.80791.10803.81371.80412.60103.15201.7803

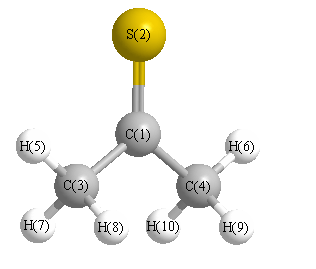
picture of Thioacetone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 111.471 C1 C3 H7 109.634
C1 C3 H8 109.634 C1 C4 H6 111.471
C1 C4 H9 109.634 C1 C4 H10 109.634
S2 C1 C3 122.543 S2 C1 C4 122.543
C3 C1 C4 114.915 H5 C3 H7 109.541
H5 C3 H8 109.541 H6 C4 H9 109.541
H6 C4 H10 109.541 H7 C3 H8 106.911
H9 C4 H10 106.911
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability