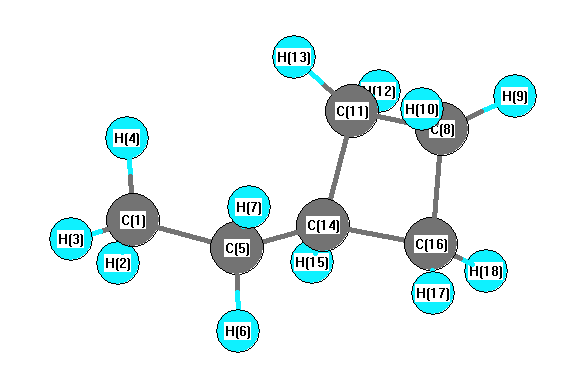
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3150 |
3018 |
79.43 |
|
|
|
| 2 |
A |
3135 |
3004 |
36.05 |
|
|
|
| 3 |
A |
3132 |
3001 |
10.27 |
|
|
|
| 4 |
A |
3128 |
2998 |
43.02 |
|
|
|
| 5 |
A |
3123 |
2993 |
11.90 |
|
|
|
| 6 |
A |
3090 |
2961 |
16.58 |
|
|
|
| 7 |
A |
3083 |
2955 |
66.94 |
|
|
|
| 8 |
A |
3075 |
2947 |
33.97 |
|
|
|
| 9 |
A |
3073 |
2944 |
63.32 |
|
|
|
| 10 |
A |
3060 |
2933 |
22.25 |
|
|
|
| 11 |
A |
3050 |
2923 |
25.64 |
|
|
|
| 12 |
A |
3038 |
2911 |
25.10 |
|
|
|
| 13 |
A |
1502 |
1440 |
2.63 |
|
|
|
| 14 |
A |
1483 |
1421 |
6.17 |
|
|
|
| 15 |
A |
1477 |
1416 |
7.23 |
|
|
|
| 16 |
A |
1471 |
1410 |
2.22 |
|
|
|
| 17 |
A |
1471 |
1409 |
1.01 |
|
|
|
| 18 |
A |
1462 |
1401 |
2.45 |
|
|
|
| 19 |
A |
1396 |
1338 |
3.23 |
|
|
|
| 20 |
A |
1388 |
1330 |
2.41 |
|
|
|
| 21 |
A |
1332 |
1276 |
3.59 |
|
|
|
| 22 |
A |
1297 |
1243 |
0.51 |
|
|
|
| 23 |
A |
1280 |
1227 |
2.18 |
|
|
|
| 24 |
A |
1273 |
1220 |
0.83 |
|
|
|
| 25 |
A |
1253 |
1201 |
0.02 |
|
|
|
| 26 |
A |
1244 |
1192 |
0.02 |
|
|
|
| 27 |
A |
1239 |
1187 |
0.07 |
|
|
|
| 28 |
A |
1191 |
1141 |
0.64 |
|
|
|
| 29 |
A |
1184 |
1134 |
0.06 |
|
|
|
| 30 |
A |
1132 |
1085 |
0.74 |
|
|
|
| 31 |
A |
1098 |
1052 |
0.06 |
|
|
|
| 32 |
A |
1041 |
997 |
0.20 |
|
|
|
| 33 |
A |
1006 |
964 |
0.45 |
|
|
|
| 34 |
A |
993 |
952 |
2.46 |
|
|
|
| 35 |
A |
974 |
933 |
0.62 |
|
|
|
| 36 |
A |
947 |
908 |
1.42 |
|
|
|
| 37 |
A |
933 |
894 |
1.09 |
|
|
|
| 38 |
A |
864 |
828 |
0.55 |
|
|
|
| 39 |
A |
817 |
783 |
0.38 |
|
|
|
| 40 |
A |
770 |
737 |
0.32 |
|
|
|
| 41 |
A |
754 |
723 |
1.68 |
|
|
|
| 42 |
A |
674 |
646 |
2.84 |
|
|
|
| 43 |
A |
420 |
402 |
0.09 |
|
|
|
| 44 |
A |
363 |
348 |
0.08 |
|
|
|
| 45 |
A |
243 |
233 |
0.03 |
|
|
|
| 46 |
A |
208 |
199 |
0.02 |
|
|
|
| 47 |
A |
132 |
127 |
0.02 |
|
|
|
| 48 |
A |
111 |
106 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36778.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 35244.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.451 |
|
|
|
| 2 |
H |
-0.117 |
|
|
|
| 3 |
H |
-0.128 |
|
|
|
| 4 |
H |
-0.149 |
|
|
|
| 5 |
C |
0.587 |
|
|
|
| 6 |
H |
-0.273 |
|
|
|
| 7 |
H |
-0.253 |
|
|
|
| 8 |
C |
0.605 |
|
|
|
| 9 |
H |
-0.401 |
|
|
|
| 10 |
H |
-0.392 |
|
|
|
| 11 |
C |
1.007 |
|
|
|
| 12 |
H |
-0.381 |
|
|
|
| 13 |
H |
-0.448 |
|
|
|
| 14 |
C |
0.215 |
|
|
|
| 15 |
H |
-0.619 |
|
|
|
| 16 |
C |
1.061 |
|
|
|
| 17 |
H |
-0.406 |
|
|
|
| 18 |
H |
-0.360 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.021 |
-0.047 |
0.044 |
0.068 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.724 |
-0.378 |
-0.601 |
| y |
-0.378 |
-40.699 |
-0.033 |
| z |
-0.601 |
-0.033 |
-39.508 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.621 |
-0.378 |
-0.601 |
| y |
-0.378 |
-0.583 |
-0.033 |
| z |
-0.601 |
-0.033 |
1.203 |
|
| Polar |
|---|
| 3z2-r2 | 2.407 |
|---|
| x2-y2 | -0.025 |
|---|
| xy | -0.378 |
|---|
| xz | -0.601 |
|---|
| yz | -0.033 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.087 |
0.096 |
0.135 |
| y |
0.096 |
10.494 |
0.042 |
| z |
0.135 |
0.042 |
9.678 |
<r2> (average value of r
2) Å
2
| <r2> |
193.593 |
| (<r2>)1/2 |
13.914 |