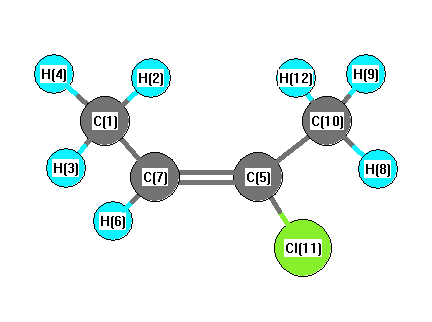
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3199 |
3066 |
7.41 |
|
|
|
| 2 |
A' |
3170 |
3038 |
14.64 |
|
|
|
| 3 |
A' |
3160 |
3028 |
12.09 |
|
|
|
| 4 |
A' |
3061 |
2933 |
13.61 |
|
|
|
| 5 |
A' |
3049 |
2922 |
31.15 |
|
|
|
| 6 |
A' |
1759 |
1686 |
13.29 |
|
|
|
| 7 |
A' |
1470 |
1409 |
7.75 |
|
|
|
| 8 |
A' |
1460 |
1399 |
5.20 |
|
|
|
| 9 |
A' |
1404 |
1345 |
10.83 |
|
|
|
| 10 |
A' |
1395 |
1337 |
0.03 |
|
|
|
| 11 |
A' |
1340 |
1284 |
0.42 |
|
|
|
| 12 |
A' |
1157 |
1109 |
26.67 |
|
|
|
| 13 |
A' |
1084 |
1039 |
47.40 |
|
|
|
| 14 |
A' |
1014 |
972 |
9.21 |
|
|
|
| 15 |
A' |
925 |
887 |
14.24 |
|
|
|
| 16 |
A' |
687 |
658 |
20.50 |
|
|
|
| 17 |
A' |
443 |
425 |
8.02 |
|
|
|
| 18 |
A' |
348 |
333 |
0.44 |
|
|
|
| 19 |
A' |
267 |
256 |
0.38 |
|
|
|
| 20 |
A" |
3131 |
3001 |
7.82 |
|
|
|
| 21 |
A" |
3110 |
2980 |
13.90 |
|
|
|
| 22 |
A" |
1454 |
1393 |
12.89 |
|
|
|
| 23 |
A" |
1445 |
1385 |
3.35 |
|
|
|
| 24 |
A" |
1049 |
1006 |
1.10 |
|
|
|
| 25 |
A" |
1046 |
1002 |
0.75 |
|
|
|
| 26 |
A" |
849 |
813 |
16.24 |
|
|
|
| 27 |
A" |
446 |
428 |
4.38 |
|
|
|
| 28 |
A" |
233 |
223 |
0.60 |
|
|
|
| 29 |
A" |
165 |
158 |
0.44 |
|
|
|
| 30 |
A" |
113 |
109 |
0.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21715.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 20810.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.129 |
|
|
|
| 2 |
H |
-0.261 |
|
|
|
| 3 |
H |
0.031 |
|
|
|
| 4 |
H |
0.031 |
|
|
|
| 5 |
C |
-0.097 |
|
|
|
| 6 |
H |
-0.521 |
|
|
|
| 7 |
C |
0.800 |
|
|
|
| 8 |
H |
0.050 |
|
|
|
| 9 |
H |
0.050 |
|
|
|
| 10 |
C |
0.238 |
|
|
|
| 11 |
Cl |
-0.280 |
|
|
|
| 12 |
H |
-0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.174 |
0.171 |
0.000 |
2.181 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.709 |
0.274 |
0.000 |
| y |
0.274 |
-35.621 |
0.000 |
| z |
0.000 |
0.000 |
-39.553 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.122 |
0.274 |
0.000 |
| y |
0.274 |
3.510 |
0.000 |
| z |
0.000 |
0.000 |
-2.388 |
|
| Polar |
|---|
| 3z2-r2 | -4.776 |
|---|
| x2-y2 | -3.088 |
|---|
| xy | 0.274 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.963 |
1.444 |
0.000 |
| y |
1.444 |
10.416 |
0.000 |
| z |
0.000 |
0.000 |
7.225 |
<r2> (average value of r
2) Å
2
| <r2> |
183.917 |
| (<r2>)1/2 |
13.562 |