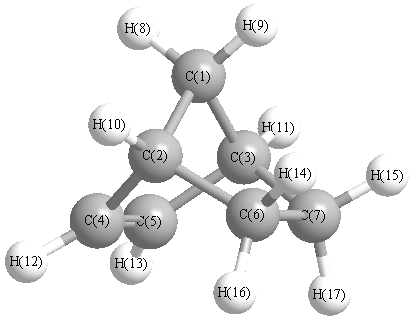
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3244 |
3109 |
16.05 |
|
|
|
| 2 |
A' |
3145 |
3014 |
31.92 |
|
|
|
| 3 |
A' |
3143 |
3012 |
50.54 |
|
|
|
| 4 |
A' |
3139 |
3008 |
8.09 |
|
|
|
| 5 |
A' |
3087 |
2958 |
55.96 |
|
|
|
| 6 |
A' |
3072 |
2943 |
41.49 |
|
|
|
| 7 |
A' |
1657 |
1588 |
4.04 |
|
|
|
| 8 |
A' |
1505 |
1442 |
1.08 |
|
|
|
| 9 |
A' |
1479 |
1417 |
5.58 |
|
|
|
| 10 |
A' |
1324 |
1268 |
0.51 |
|
|
|
| 11 |
A' |
1310 |
1256 |
1.72 |
|
|
|
| 12 |
A' |
1188 |
1138 |
0.30 |
|
|
|
| 13 |
A' |
1141 |
1094 |
5.41 |
|
|
|
| 14 |
A' |
1108 |
1062 |
1.64 |
|
|
|
| 15 |
A' |
1037 |
993 |
1.28 |
|
|
|
| 16 |
A' |
986 |
945 |
0.65 |
|
|
|
| 17 |
A' |
968 |
928 |
0.40 |
|
|
|
| 18 |
A' |
928 |
889 |
0.85 |
|
|
|
| 19 |
A' |
900 |
862 |
6.65 |
|
|
|
| 20 |
A' |
821 |
786 |
0.60 |
|
|
|
| 21 |
A' |
784 |
751 |
6.15 |
|
|
|
| 22 |
A' |
728 |
698 |
45.26 |
|
|
|
| 23 |
A' |
476 |
456 |
1.48 |
|
|
|
| 24 |
A' |
379 |
364 |
3.00 |
|
|
|
| 25 |
A" |
3218 |
3083 |
8.98 |
|
|
|
| 26 |
A" |
3137 |
3006 |
73.43 |
|
|
|
| 27 |
A" |
3126 |
2996 |
3.34 |
|
|
|
| 28 |
A" |
3074 |
2946 |
28.71 |
|
|
|
| 29 |
A" |
1479 |
1418 |
1.26 |
|
|
|
| 30 |
A" |
1366 |
1309 |
9.48 |
|
|
|
| 31 |
A" |
1299 |
1245 |
3.62 |
|
|
|
| 32 |
A" |
1278 |
1225 |
0.41 |
|
|
|
| 33 |
A" |
1264 |
1211 |
1.03 |
|
|
|
| 34 |
A" |
1234 |
1182 |
0.43 |
|
|
|
| 35 |
A" |
1194 |
1144 |
0.13 |
|
|
|
| 36 |
A" |
1127 |
1080 |
0.40 |
|
|
|
| 37 |
A" |
1043 |
1000 |
0.68 |
|
|
|
| 38 |
A" |
963 |
923 |
0.00 |
|
|
|
| 39 |
A" |
956 |
916 |
0.15 |
|
|
|
| 40 |
A" |
930 |
891 |
4.91 |
|
|
|
| 41 |
A" |
852 |
817 |
4.65 |
|
|
|
| 42 |
A" |
808 |
774 |
1.71 |
|
|
|
| 43 |
A" |
675 |
647 |
0.62 |
|
|
|
| 44 |
A" |
485 |
465 |
0.16 |
|
|
|
| 45 |
A" |
253 |
243 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33653.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 32249.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.996 |
|
|
|
| 2 |
C |
0.502 |
|
|
|
| 3 |
C |
0.502 |
|
|
|
| 4 |
C |
0.465 |
|
|
|
| 5 |
C |
0.465 |
|
|
|
| 6 |
C |
0.595 |
|
|
|
| 7 |
C |
0.595 |
|
|
|
| 8 |
H |
-0.435 |
|
|
|
| 9 |
H |
-0.335 |
|
|
|
| 10 |
H |
-0.518 |
|
|
|
| 11 |
H |
-0.518 |
|
|
|
| 12 |
H |
-0.493 |
|
|
|
| 13 |
H |
-0.493 |
|
|
|
| 14 |
H |
-0.327 |
|
|
|
| 15 |
H |
-0.327 |
|
|
|
| 16 |
H |
-0.338 |
|
|
|
| 17 |
H |
-0.338 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.215 |
-0.323 |
0.000 |
0.388 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.633 |
-0.261 |
0.000 |
| y |
-0.261 |
-45.429 |
0.000 |
| z |
0.000 |
0.000 |
-42.177 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.170 |
-0.261 |
0.000 |
| y |
-0.261 |
-3.025 |
0.000 |
| z |
0.000 |
0.000 |
1.854 |
|
| Polar |
|---|
| 3z2-r2 | 3.709 |
|---|
| x2-y2 | 2.797 |
|---|
| xy | -0.261 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.068 |
-0.556 |
0.000 |
| y |
-0.556 |
11.184 |
0.000 |
| z |
0.000 |
0.000 |
11.873 |
<r2> (average value of r
2) Å
2
| <r2> |
164.601 |
| (<r2>)1/2 |
12.830 |