Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -193.085597 |
| Energy at 298.15K | -193.092786 |
| HF Energy | -193.085597 |
| Nuclear repulsion energy | 128.247136 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3112 |
2983 |
27.65 |
|
|
|
| 2 |
A1 |
3057 |
2930 |
0.27 |
|
|
|
| 3 |
A1 |
1532 |
1468 |
0.00 |
|
|
|
| 4 |
A1 |
1481 |
1419 |
4.22 |
|
|
|
| 5 |
A1 |
1368 |
1311 |
1.26 |
|
|
|
| 6 |
A1 |
1072 |
1028 |
7.40 |
|
|
|
| 7 |
A1 |
954 |
914 |
32.09 |
|
|
|
| 8 |
A1 |
815 |
781 |
5.63 |
|
|
|
| 9 |
A2 |
3102 |
2973 |
0.00 |
|
|
|
| 10 |
A2 |
1229 |
1178 |
0.00 |
|
|
|
| 11 |
A2 |
1158 |
1110 |
0.00 |
|
|
|
| 12 |
A2 |
834 |
799 |
0.00 |
|
|
|
| 13 |
B1 |
3175 |
3042 |
30.46 |
|
|
|
| 14 |
B1 |
3101 |
2971 |
68.09 |
|
|
|
| 15 |
B1 |
1192 |
1142 |
0.08 |
|
|
|
| 16 |
B1 |
1143 |
1096 |
1.96 |
|
|
|
| 17 |
B1 |
766 |
734 |
0.29 |
|
|
|
| 18 |
B1 |
36 |
34 |
3.67 |
|
|
|
| 19 |
B2 |
3048 |
2921 |
159.89 |
|
|
|
| 20 |
B2 |
1503 |
1440 |
0.44 |
|
|
|
| 21 |
B2 |
1302 |
1248 |
1.21 |
|
|
|
| 22 |
B2 |
1249 |
1197 |
7.54 |
|
|
|
| 23 |
B2 |
1066 |
1021 |
99.73 |
|
|
|
| 24 |
B2 |
965 |
925 |
5.28 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19129.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 18331.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.068 |
| C2 |
0.000 |
0.000 |
-1.072 |
| C3 |
0.000 |
1.034 |
0.063 |
| C4 |
0.000 |
-1.034 |
0.063 |
| H5 |
0.893 |
0.000 |
-1.704 |
| H6 |
-0.893 |
0.000 |
-1.704 |
| H7 |
0.896 |
1.667 |
0.136 |
| H8 |
-0.896 |
1.667 |
0.136 |
| H9 |
-0.896 |
-1.667 |
0.136 |
| H10 |
0.896 |
-1.667 |
0.136 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1400 | 1.4417 | 1.4417 | 2.9123 | 2.9123 | 2.1092 | 2.1092 | 2.1092 | 2.1092 |
C2 | 2.1400 | | 1.5359 | 1.5359 | 1.0940 | 1.0940 | 2.2454 | 2.2454 | 2.2454 | 2.2454 | C3 | 1.4417 | 1.5359 | | 2.0684 | 2.2341 | 2.2341 | 1.0992 | 1.0992 | 2.8468 | 2.8468 | C4 | 1.4417 | 1.5359 | 2.0684 | | 2.2341 | 2.2341 | 2.8468 | 2.8468 | 1.0992 | 1.0992 | H5 | 2.9123 | 1.0940 | 2.2341 | 2.2341 | | 1.7860 | 2.4832 | 3.0605 | 3.0605 | 2.4832 | H6 | 2.9123 | 1.0940 | 2.2341 | 2.2341 | 1.7860 | | 3.0605 | 2.4832 | 2.4832 | 3.0605 | H7 | 2.1092 | 2.2454 | 1.0992 | 2.8468 | 2.4832 | 3.0605 | | 1.7918 | 3.7849 | 3.3339 | H8 | 2.1092 | 2.2454 | 1.0992 | 2.8468 | 3.0605 | 2.4832 | 1.7918 | | 3.3339 | 3.7849 | H9 | 2.1092 | 2.2454 | 2.8468 | 1.0992 | 3.0605 | 2.4832 | 3.7849 | 3.3339 | | 1.7918 | H10 | 2.1092 | 2.2454 | 2.8468 | 1.0992 | 2.4832 | 3.0605 | 3.3339 | 3.7849 | 1.7918 | |
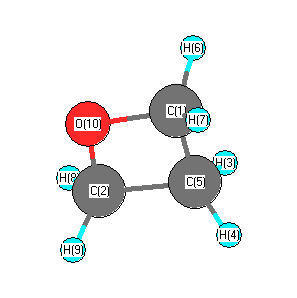 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.839 |
|
O1 |
C3 |
H7 |
111.508 |
| O1 |
C3 |
H8 |
111.508 |
|
O1 |
C4 |
C2 |
91.839 |
| O1 |
C4 |
H9 |
111.508 |
|
O1 |
C4 |
H10 |
111.508 |
| C2 |
C3 |
H7 |
115.894 |
|
C2 |
C3 |
H8 |
115.894 |
| C2 |
C4 |
H9 |
115.894 |
|
C2 |
C4 |
H10 |
115.894 |
| C3 |
O1 |
C4 |
91.672 |
|
C3 |
C2 |
C4 |
84.651 |
| C3 |
C2 |
H5 |
115.285 |
|
C3 |
C2 |
H6 |
115.285 |
| C4 |
C2 |
H5 |
115.285 |
|
C4 |
C2 |
H6 |
115.285 |
| H5 |
C2 |
H6 |
109.419 |
|
H7 |
C3 |
H8 |
109.178 |
| H9 |
C4 |
H10 |
109.178 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.736 |
|
|
|
| 2 |
C |
0.560 |
|
|
|
| 3 |
C |
1.054 |
|
|
|
| 4 |
C |
1.054 |
|
|
|
| 5 |
H |
-0.307 |
|
|
|
| 6 |
H |
-0.307 |
|
|
|
| 7 |
H |
-0.329 |
|
|
|
| 8 |
H |
-0.329 |
|
|
|
| 9 |
H |
-0.329 |
|
|
|
| 10 |
H |
-0.329 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.958 |
1.958 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.176 |
0.000 |
0.000 |
| y |
0.000 |
-23.134 |
0.000 |
| z |
0.000 |
0.000 |
-28.045 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.413 |
0.000 |
0.000 |
| y |
0.000 |
2.977 |
0.000 |
| z |
0.000 |
0.000 |
-4.390 |
|
| Polar |
|---|
| 3z2-r2 | -8.780 |
|---|
| x2-y2 | -1.042 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.367 |
0.000 |
0.000 |
| y |
0.000 |
6.630 |
0.000 |
| z |
0.000 |
0.000 |
5.816 |
<r2> (average value of r
2) Å
2
| <r2> |
66.394 |
| (<r2>)1/2 |
8.148 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -193.085597 |
| Energy at 298.15K | -193.092786 |
| HF Energy | -193.085597 |
| Nuclear repulsion energy | 128.246970 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3174 |
3042 |
30.52 |
|
|
|
| 2 |
A' |
3112 |
2983 |
27.67 |
|
|
|
| 3 |
A' |
3101 |
2971 |
68.04 |
|
|
|
| 4 |
A' |
3057 |
2930 |
0.27 |
|
|
|
| 5 |
A' |
1532 |
1468 |
0.00 |
|
|
|
| 6 |
A' |
1481 |
1419 |
4.22 |
|
|
|
| 7 |
A' |
1368 |
1311 |
1.26 |
|
|
|
| 8 |
A' |
1192 |
1142 |
0.08 |
|
|
|
| 9 |
A' |
1143 |
1096 |
1.96 |
|
|
|
| 10 |
A' |
1072 |
1028 |
7.40 |
|
|
|
| 11 |
A' |
954 |
914 |
32.08 |
|
|
|
| 12 |
A' |
815 |
781 |
5.62 |
|
|
|
| 13 |
A' |
766 |
734 |
0.30 |
|
|
|
| 14 |
A' |
36 |
35 |
3.67 |
|
|
|
| 15 |
A" |
3102 |
2973 |
0.00 |
|
|
|
| 16 |
A" |
3048 |
2921 |
159.88 |
|
|
|
| 17 |
A" |
1503 |
1440 |
0.44 |
|
|
|
| 18 |
A" |
1302 |
1248 |
1.21 |
|
|
|
| 19 |
A" |
1249 |
1197 |
7.53 |
|
|
|
| 20 |
A" |
1229 |
1178 |
0.00 |
|
|
|
| 21 |
A" |
1158 |
1110 |
0.00 |
|
|
|
| 22 |
A" |
1066 |
1021 |
99.75 |
|
|
|
| 23 |
A" |
965 |
925 |
5.27 |
|
|
|
| 24 |
A" |
834 |
799 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19129.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 18332.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.000 |
-1.068 |
0.000 |
| C2 |
0.000 |
1.072 |
0.000 |
| C3 |
0.000 |
-0.063 |
1.034 |
| C4 |
0.000 |
-0.063 |
-1.034 |
| H5 |
0.893 |
1.705 |
0.000 |
| H6 |
-0.893 |
1.704 |
0.000 |
| H7 |
0.896 |
-0.136 |
1.667 |
| H8 |
-0.896 |
-0.136 |
1.667 |
| H9 |
0.896 |
-0.136 |
-1.667 |
| H10 |
-0.896 |
-0.136 |
-1.667 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1399 | 1.4417 | 1.4417 | 2.9125 | 2.9121 | 2.1093 | 2.1093 | 2.1093 | 2.1093 |
C2 | 2.1399 | | 1.5359 | 1.5359 | 1.0941 | 1.0940 | 2.2453 | 2.2455 | 2.2453 | 2.2455 | C3 | 1.4417 | 1.5359 | | 2.0685 | 2.2342 | 2.2342 | 1.0992 | 1.0992 | 2.8467 | 2.8470 | C4 | 1.4417 | 1.5359 | 2.0685 | | 2.2342 | 2.2342 | 2.8467 | 2.8470 | 1.0992 | 1.0992 | H5 | 2.9125 | 1.0941 | 2.2342 | 2.2342 | | 1.7859 | 2.4832 | 3.0605 | 2.4832 | 3.0605 | H6 | 2.9121 | 1.0940 | 2.2342 | 2.2342 | 1.7859 | | 3.0605 | 2.4833 | 3.0605 | 2.4833 | H7 | 2.1093 | 2.2453 | 1.0992 | 2.8467 | 2.4832 | 3.0605 | | 1.7918 | 3.3334 | 3.7849 | H8 | 2.1093 | 2.2455 | 1.0992 | 2.8470 | 3.0605 | 2.4833 | 1.7918 | | 3.7849 | 3.3345 | H9 | 2.1093 | 2.2453 | 2.8467 | 1.0992 | 2.4832 | 3.0605 | 3.3334 | 3.7849 | | 1.7918 | H10 | 2.1093 | 2.2455 | 2.8470 | 1.0992 | 3.0605 | 2.4833 | 3.7849 | 3.3345 | 1.7918 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.834 |
|
O1 |
C3 |
H7 |
111.514 |
| O1 |
C3 |
H8 |
111.512 |
|
O1 |
C4 |
C2 |
91.834 |
| O1 |
C4 |
H9 |
111.514 |
|
O1 |
C4 |
H10 |
111.512 |
| C2 |
C3 |
H7 |
115.885 |
|
C2 |
C3 |
H8 |
115.895 |
| C2 |
C4 |
H9 |
115.885 |
|
C2 |
C4 |
H10 |
115.895 |
| C3 |
O1 |
C4 |
91.678 |
|
C3 |
C2 |
C4 |
84.654 |
| C3 |
C2 |
H5 |
115.292 |
|
C3 |
C2 |
H6 |
115.286 |
| C4 |
C2 |
H5 |
115.292 |
|
C4 |
C2 |
H6 |
115.286 |
| H5 |
C2 |
H6 |
109.406 |
|
H7 |
C3 |
H8 |
109.180 |
| H9 |
C4 |
H10 |
109.180 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.736 |
|
|
|
| 2 |
C |
0.560 |
|
|
|
| 3 |
C |
1.054 |
|
|
|
| 4 |
C |
1.054 |
|
|
|
| 5 |
H |
-0.307 |
|
|
|
| 6 |
H |
-0.307 |
|
|
|
| 7 |
H |
-0.329 |
|
|
|
| 8 |
H |
-0.329 |
|
|
|
| 9 |
H |
-0.329 |
|
|
|
| 10 |
H |
-0.329 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.001 |
1.958 |
0.000 |
1.958 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.177 |
-0.001 |
0.000 |
| y |
-0.001 |
-28.045 |
0.000 |
| z |
0.000 |
0.000 |
-23.133 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.413 |
-0.001 |
0.000 |
| y |
-0.001 |
-4.390 |
0.000 |
| z |
0.000 |
0.000 |
2.977 |
|
| Polar |
|---|
| 3z2-r2 | 5.954 |
|---|
| x2-y2 | 3.868 |
|---|
| xy | -0.001 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.367 |
0.000 |
0.000 |
| y |
0.000 |
5.816 |
0.000 |
| z |
0.000 |
0.000 |
6.630 |
<r2> (average value of r
2) Å
2
| <r2> |
66.394 |
| (<r2>)1/2 |
8.148 |
