Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -192.629513 |
| Energy at 298.15K | |
| HF Energy | -191.931947 |
| Nuclear repulsion energy | 126.571592 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3103 |
3031 |
|
|
|
|
| 2 |
A1 |
3054 |
2983 |
|
|
|
|
| 3 |
A1 |
1592 |
1555 |
|
|
|
|
| 4 |
A1 |
1538 |
1502 |
|
|
|
|
| 5 |
A1 |
1401 |
1368 |
|
|
|
|
| 6 |
A1 |
1062 |
1037 |
|
|
|
|
| 7 |
A1 |
936 |
915 |
|
|
|
|
| 8 |
A1 |
809 |
790 |
|
|
|
|
| 9 |
A2 |
3097 |
3026 |
|
|
|
|
| 10 |
A2 |
1255 |
1226 |
|
|
|
|
| 11 |
A2 |
1174 |
1147 |
|
|
|
|
| 12 |
A2 |
857 |
838 |
|
|
|
|
| 13 |
B1 |
3161 |
3088 |
|
|
|
|
| 14 |
B1 |
3096 |
3025 |
|
|
|
|
| 15 |
B1 |
1214 |
1186 |
|
|
|
|
| 16 |
B1 |
1158 |
1131 |
|
|
|
|
| 17 |
B1 |
782 |
764 |
|
|
|
|
| 18 |
B1 |
83i |
81i |
|
|
|
|
| 19 |
B2 |
3045 |
2975 |
|
|
|
|
| 20 |
B2 |
1563 |
1527 |
|
|
|
|
| 21 |
B2 |
1327 |
1296 |
|
|
|
|
| 22 |
B2 |
1285 |
1256 |
|
|
|
|
| 23 |
B2 |
1049 |
1025 |
|
|
|
|
| 24 |
B2 |
970 |
947 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19221.9 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 18777.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.087 |
| C2 |
0.000 |
0.000 |
-1.084 |
| C3 |
0.000 |
1.052 |
0.059 |
| C4 |
0.000 |
-1.052 |
0.059 |
| H5 |
0.900 |
0.000 |
-1.718 |
| H6 |
-0.900 |
0.000 |
-1.718 |
| H7 |
0.904 |
1.682 |
0.135 |
| H8 |
-0.904 |
1.682 |
0.135 |
| H9 |
-0.904 |
-1.682 |
0.135 |
| H10 |
0.904 |
-1.682 |
0.135 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1702 | 1.4704 | 1.4704 | 2.9453 | 2.9453 | 2.1340 | 2.1340 | 2.1340 | 2.1340 |
C2 | 2.1702 | | 1.5528 | 1.5528 | 1.1012 | 1.1012 | 2.2654 | 2.2654 | 2.2654 | 2.2654 | C3 | 1.4704 | 1.5528 | | 2.1033 | 2.2524 | 2.2524 | 1.1051 | 1.1051 | 2.8806 | 2.8806 | C4 | 1.4704 | 1.5528 | 2.1033 | | 2.2524 | 2.2524 | 2.8806 | 2.8806 | 1.1051 | 1.1051 | H5 | 2.9453 | 1.1012 | 2.2524 | 2.2524 | | 1.8005 | 2.5023 | 3.0852 | 3.0852 | 2.5023 | H6 | 2.9453 | 1.1012 | 2.2524 | 2.2524 | 1.8005 | | 3.0852 | 2.5023 | 2.5023 | 3.0852 | H7 | 2.1340 | 2.2654 | 1.1051 | 2.8806 | 2.5023 | 3.0852 | | 1.8087 | 3.8198 | 3.3644 | H8 | 2.1340 | 2.2654 | 1.1051 | 2.8806 | 3.0852 | 2.5023 | 1.8087 | | 3.3644 | 3.8198 | H9 | 2.1340 | 2.2654 | 2.8806 | 1.1051 | 3.0852 | 2.5023 | 3.8198 | 3.3644 | | 1.8087 | H10 | 2.1340 | 2.2654 | 2.8806 | 1.1051 | 2.5023 | 3.0852 | 3.3644 | 3.8198 | 1.8087 | |
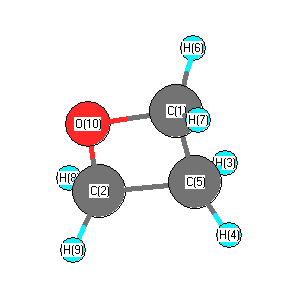 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.710 |
|
O1 |
C3 |
H7 |
111.110 |
| O1 |
C3 |
H8 |
111.110 |
|
O1 |
C4 |
C2 |
91.710 |
| O1 |
C4 |
H9 |
111.110 |
|
O1 |
C4 |
H10 |
111.110 |
| C2 |
C3 |
H7 |
115.906 |
|
C2 |
C3 |
H8 |
115.906 |
| C2 |
C4 |
H9 |
115.906 |
|
C2 |
C4 |
H10 |
115.906 |
| C3 |
O1 |
C4 |
91.321 |
|
C3 |
C2 |
C4 |
85.260 |
| C3 |
C2 |
H5 |
115.071 |
|
C3 |
C2 |
H6 |
115.071 |
| C4 |
C2 |
H5 |
115.071 |
|
C4 |
C2 |
H6 |
115.071 |
| H5 |
C2 |
H6 |
109.671 |
|
H7 |
C3 |
H8 |
109.843 |
| H9 |
C4 |
H10 |
109.843 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -192.629685 |
| Energy at 298.15K | -192.636949 |
| HF Energy | -191.931502 |
| Nuclear repulsion energy | 126.779003 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3161 |
3088 |
|
|
|
|
| 2 |
A' |
3106 |
3035 |
|
|
|
|
| 3 |
A' |
3098 |
3027 |
|
|
|
|
| 4 |
A' |
3046 |
2975 |
|
|
|
|
| 5 |
A' |
1589 |
1553 |
|
|
|
|
| 6 |
A' |
1535 |
1500 |
|
|
|
|
| 7 |
A' |
1398 |
1366 |
|
|
|
|
| 8 |
A' |
1221 |
1193 |
|
|
|
|
| 9 |
A' |
1168 |
1141 |
|
|
|
|
| 10 |
A' |
1064 |
1040 |
|
|
|
|
| 11 |
A' |
934 |
913 |
|
|
|
|
| 12 |
A' |
861 |
841 |
|
|
|
|
| 13 |
A' |
728 |
711 |
|
|
|
|
| 14 |
A' |
108 |
105 |
|
|
|
|
| 15 |
A" |
3104 |
3033 |
|
|
|
|
| 16 |
A" |
3038 |
2968 |
|
|
|
|
| 17 |
A" |
1560 |
1524 |
|
|
|
|
| 18 |
A" |
1328 |
1297 |
|
|
|
|
| 19 |
A" |
1290 |
1260 |
|
|
|
|
| 20 |
A" |
1245 |
1216 |
|
|
|
|
| 21 |
A" |
1168 |
1141 |
|
|
|
|
| 22 |
A" |
1037 |
1014 |
|
|
|
|
| 23 |
A" |
971 |
948 |
|
|
|
|
| 24 |
A" |
855 |
835 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19306.7 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 18860.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.227 |
-1.061 |
0.000 |
| C2 |
-0.062 |
1.081 |
0.000 |
| C3 |
-0.062 |
-0.066 |
1.044 |
| C4 |
-0.062 |
-0.066 |
-1.044 |
| H5 |
0.872 |
1.665 |
0.000 |
| H6 |
-0.932 |
1.757 |
0.000 |
| H7 |
0.725 |
-0.056 |
1.818 |
| H8 |
-1.047 |
-0.259 |
1.510 |
| H9 |
0.725 |
-0.056 |
-1.818 |
| H10 |
-1.047 |
-0.259 |
-1.510 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1606 | 1.4709 | 1.4709 | 2.8011 | 3.0461 | 2.1362 | 2.1325 | 2.1362 | 2.1325 |
C2 | 2.1606 | | 1.5510 | 1.5510 | 1.1015 | 1.1017 | 2.2840 | 2.2466 | 2.2840 | 2.2466 | C3 | 1.4709 | 1.5510 | | 2.0890 | 2.2270 | 2.2737 | 1.1036 | 1.1072 | 2.9689 | 2.7451 | C4 | 1.4709 | 1.5510 | 2.0890 | | 2.2270 | 2.2737 | 2.9689 | 2.7451 | 1.1036 | 1.1072 | H5 | 2.8011 | 1.1015 | 2.2270 | 2.2270 | | 1.8058 | 2.5080 | 3.1092 | 2.5080 | 3.1092 | H6 | 3.0461 | 1.1017 | 2.2737 | 2.2737 | 1.8058 | | 3.0555 | 2.5213 | 3.0555 | 2.5213 | H7 | 2.1362 | 2.2840 | 1.1036 | 2.9689 | 2.5080 | 3.0555 | | 1.8106 | 3.6364 | 3.7765 | H8 | 2.1325 | 2.2466 | 1.1072 | 2.7451 | 3.1092 | 2.5213 | 1.8106 | | 3.7765 | 3.0204 | H9 | 2.1362 | 2.2840 | 2.9689 | 1.1036 | 2.5080 | 3.0555 | 3.6364 | 3.7765 | | 1.8106 | H10 | 2.1325 | 2.2466 | 2.7451 | 1.1072 | 3.1092 | 2.5213 | 3.7765 | 3.0204 | 1.8106 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.242 |
|
O1 |
C3 |
H7 |
111.354 |
| O1 |
C3 |
H8 |
110.827 |
|
O1 |
C4 |
C2 |
91.242 |
| O1 |
C4 |
H9 |
111.354 |
|
O1 |
C4 |
H10 |
110.827 |
| C2 |
C3 |
H7 |
117.738 |
|
C2 |
C3 |
H8 |
114.338 |
| C2 |
C4 |
H9 |
117.738 |
|
C2 |
C4 |
H10 |
114.338 |
| C3 |
O1 |
C4 |
90.489 |
|
C3 |
C2 |
C4 |
84.662 |
| C3 |
C2 |
H5 |
113.101 |
|
C3 |
C2 |
H6 |
116.976 |
| C4 |
C2 |
H5 |
113.101 |
|
C4 |
C2 |
H6 |
116.976 |
| H5 |
C2 |
H6 |
110.094 |
|
H7 |
C3 |
H8 |
109.965 |
| H9 |
C4 |
H10 |
109.965 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
