Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.284858 |
| Energy at 298.15K | -133.289046 |
| HF Energy | -133.284858 |
| Nuclear repulsion energy | 64.810842 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3494 |
3348 |
1.22 |
|
|
|
| 2 |
A' |
3302 |
3164 |
3.12 |
|
|
|
| 3 |
A' |
3182 |
3049 |
3.54 |
|
|
|
| 4 |
A' |
3086 |
2957 |
40.28 |
|
|
|
| 5 |
A' |
1511 |
1448 |
8.80 |
|
|
|
| 6 |
A' |
1457 |
1397 |
1.81 |
|
|
|
| 7 |
A' |
1361 |
1304 |
29.97 |
|
|
|
| 8 |
A' |
1255 |
1202 |
6.58 |
|
|
|
| 9 |
A' |
1122 |
1076 |
20.23 |
|
|
|
| 10 |
A' |
995 |
954 |
10.34 |
|
|
|
| 11 |
A' |
485 |
465 |
12.44 |
|
|
|
| 12 |
A" |
1042 |
998 |
0.47 |
|
|
|
| 13 |
A" |
837 |
803 |
13.28 |
|
|
|
| 14 |
A" |
681 |
652 |
100.83 |
|
|
|
| 15 |
A" |
524 |
503 |
4.66 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12166.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 11659.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.417 |
0.000 |
| C2 |
1.153 |
-0.377 |
0.000 |
| N3 |
-1.192 |
-0.136 |
0.000 |
| H4 |
0.134 |
1.509 |
0.000 |
| H5 |
2.145 |
0.069 |
0.000 |
| H6 |
1.057 |
-1.461 |
0.000 |
| H7 |
-1.912 |
0.590 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3999 | 1.3141 | 1.1000 | 2.1733 | 2.1556 | 1.9194 |
C2 | 1.3999 | | 2.3571 | 2.1434 | 1.0877 | 1.0887 | 3.2134 | N3 | 1.3141 | 2.3571 | | 2.1132 | 3.3433 | 2.6104 | 1.0223 | H4 | 1.1000 | 2.1434 | 2.1132 | | 2.4734 | 3.1104 | 2.2430 | H5 | 2.1733 | 1.0877 | 3.3433 | 2.4734 | | 1.8774 | 4.0901 | H6 | 2.1556 | 1.0887 | 2.6104 | 3.1104 | 1.8774 | | 3.6085 | H7 | 1.9194 | 3.2134 | 1.0223 | 2.2430 | 4.0901 | 3.6085 | |
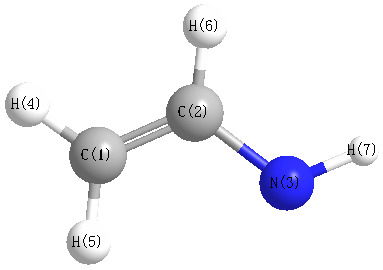 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.258 |
|
C1 |
C2 |
H6 |
119.512 |
| C1 |
N3 |
H7 |
109.849 |
|
C2 |
C1 |
N3 |
120.535 |
| C2 |
C1 |
H4 |
117.543 |
|
N3 |
C1 |
H4 |
121.922 |
| H5 |
C2 |
H6 |
119.230 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.899 |
|
|
|
| 2 |
C |
0.458 |
|
|
|
| 3 |
N |
-0.417 |
|
|
|
| 4 |
H |
-0.379 |
|
|
|
| 5 |
H |
-0.298 |
|
|
|
| 6 |
H |
-0.303 |
|
|
|
| 7 |
H |
0.040 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.152 |
1.578 |
0.000 |
1.954 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.156 |
-2.868 |
0.000 |
| y |
-2.868 |
-17.892 |
0.000 |
| z |
0.000 |
0.000 |
-20.489 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.034 |
-2.868 |
0.000 |
| y |
-2.868 |
0.930 |
0.000 |
| z |
0.000 |
0.000 |
-2.965 |
|
| Polar |
|---|
| 3z2-r2 | -5.930 |
|---|
| x2-y2 | 0.736 |
|---|
| xy | -2.868 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.380 |
-0.334 |
0.000 |
| y |
-0.334 |
5.017 |
0.000 |
| z |
0.000 |
0.000 |
3.749 |
<r2> (average value of r
2) Å
2
| <r2> |
45.665 |
| (<r2>)1/2 |
6.758 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -133.283822 |
| Energy at 298.15K | -133.287992 |
| HF Energy | -133.283822 |
| Nuclear repulsion energy | 64.821190 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3441 |
3298 |
4.88 |
|
|
|
| 2 |
A' |
3282 |
3146 |
6.92 |
|
|
|
| 3 |
A' |
3167 |
3034 |
3.86 |
|
|
|
| 4 |
A' |
3141 |
3010 |
18.94 |
|
|
|
| 5 |
A' |
1493 |
1431 |
6.04 |
|
|
|
| 6 |
A' |
1451 |
1390 |
2.89 |
|
|
|
| 7 |
A' |
1384 |
1326 |
10.33 |
|
|
|
| 8 |
A' |
1245 |
1193 |
28.11 |
|
|
|
| 9 |
A' |
1148 |
1101 |
31.79 |
|
|
|
| 10 |
A' |
983 |
942 |
0.67 |
|
|
|
| 11 |
A' |
490 |
470 |
8.19 |
|
|
|
| 12 |
A" |
1076 |
1031 |
52.06 |
|
|
|
| 13 |
A" |
838 |
803 |
61.51 |
|
|
|
| 14 |
A" |
688 |
659 |
7.27 |
|
|
|
| 15 |
A" |
484 |
464 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12155.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 11648.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.450 |
0.000 |
| C2 |
1.128 |
-0.382 |
0.000 |
| N3 |
-1.256 |
0.065 |
0.000 |
| H4 |
0.167 |
1.534 |
0.000 |
| H5 |
2.135 |
0.032 |
0.000 |
| H6 |
1.018 |
-1.467 |
0.000 |
| H7 |
-1.295 |
-0.961 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4014 | 1.3138 | 1.0963 | 2.1756 | 2.1700 | 1.9155 |
C2 | 1.4014 | | 2.4253 | 2.1429 | 1.0885 | 1.0905 | 2.4917 | N3 | 1.3138 | 2.4253 | | 2.0449 | 3.3911 | 2.7411 | 1.0263 | H4 | 1.0963 | 2.1429 | 2.0449 | | 2.4760 | 3.1184 | 2.8914 | H5 | 2.1756 | 1.0885 | 3.3911 | 2.4760 | | 1.8689 | 3.5712 | H6 | 2.1700 | 1.0905 | 2.7411 | 3.1184 | 1.8689 | | 2.3677 | H7 | 1.9155 | 2.4917 | 1.0263 | 2.8914 | 3.5712 | 2.3677 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.293 |
|
C1 |
C2 |
H6 |
120.594 |
| C1 |
N3 |
H7 |
109.269 |
|
C2 |
C1 |
N3 |
126.537 |
| C2 |
C1 |
H4 |
117.659 |
|
N3 |
C1 |
H4 |
115.805 |
| H5 |
C2 |
H6 |
118.113 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.764 |
|
|
|
| 2 |
C |
0.619 |
|
|
|
| 3 |
N |
-0.397 |
|
|
|
| 4 |
H |
-0.294 |
|
|
|
| 5 |
H |
-0.317 |
|
|
|
| 6 |
H |
-0.373 |
|
|
|
| 7 |
H |
-0.002 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.219 |
-1.023 |
0.000 |
2.443 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.076 |
2.131 |
0.000 |
| y |
2.131 |
-16.362 |
0.000 |
| z |
0.000 |
0.000 |
-20.472 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.659 |
2.131 |
0.000 |
| y |
2.131 |
4.412 |
0.000 |
| z |
0.000 |
0.000 |
-1.753 |
|
| Polar |
|---|
| 3z2-r2 | -3.506 |
|---|
| x2-y2 | -4.714 |
|---|
| xy | 2.131 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.208 |
-0.253 |
0.000 |
| y |
-0.253 |
5.043 |
0.000 |
| z |
0.000 |
0.000 |
3.771 |
<r2> (average value of r
2) Å
2
| <r2> |
45.578 |
| (<r2>)1/2 |
6.751 |
