Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.950544 |
| Energy at 298.15K | -132.954721 |
| HF Energy | -132.487061 |
| Nuclear repulsion energy | 64.488847 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3426 |
3347 |
|
|
|
|
| 2 |
A' |
3307 |
3231 |
|
|
|
|
| 3 |
A' |
3198 |
3124 |
|
|
|
|
| 4 |
A' |
3089 |
3018 |
|
|
|
|
| 5 |
A' |
1542 |
1506 |
|
|
|
|
| 6 |
A' |
1485 |
1450 |
|
|
|
|
| 7 |
A' |
1393 |
1361 |
|
|
|
|
| 8 |
A' |
1252 |
1223 |
|
|
|
|
| 9 |
A' |
1094 |
1069 |
|
|
|
|
| 10 |
A' |
1020 |
996 |
|
|
|
|
| 11 |
A' |
498 |
486 |
|
|
|
|
| 12 |
A" |
1102 |
1077 |
|
|
|
|
| 13 |
A" |
804 |
785 |
|
|
|
|
| 14 |
A" |
722 |
705 |
|
|
|
|
| 15 |
A" |
545 |
533 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12237.5 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 11954.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.422 |
0.000 |
| C2 |
1.156 |
-0.394 |
0.000 |
| N3 |
-1.196 |
-0.123 |
0.000 |
| H4 |
0.149 |
1.517 |
0.000 |
| H5 |
2.158 |
0.044 |
0.000 |
| H6 |
1.042 |
-1.483 |
0.000 |
| H7 |
-1.911 |
0.617 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4148 | 1.3145 | 1.1055 | 2.1908 | 2.1709 | 1.9212 |
C2 | 1.4148 | | 2.3673 | 2.1605 | 1.0936 | 1.0946 | 3.2294 | N3 | 1.3145 | 2.3673 | | 2.1214 | 3.3581 | 2.6183 | 1.0295 | H4 | 1.1055 | 2.1605 | 2.1214 | | 2.4918 | 3.1302 | 2.2480 | H5 | 2.1908 | 1.0936 | 3.3581 | 2.4918 | | 1.8908 | 4.1094 | H6 | 2.1709 | 1.0946 | 2.6183 | 3.1302 | 1.8908 | | 3.6235 | H7 | 1.9212 | 3.2294 | 1.0295 | 2.2480 | 4.1094 | 3.6235 | |
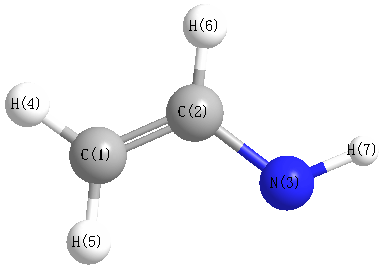 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.183 |
|
C1 |
C2 |
H6 |
119.250 |
| C1 |
N3 |
H7 |
109.494 |
|
C2 |
C1 |
N3 |
120.267 |
| C2 |
C1 |
H4 |
117.493 |
|
N3 |
C1 |
H4 |
122.240 |
| H5 |
C2 |
H6 |
119.567 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -132.949653 |
| Energy at 298.15K | -132.953866 |
| HF Energy | -132.486096 |
| Nuclear repulsion energy | 64.487631 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3367 |
3289 |
|
|
|
|
| 2 |
A' |
3286 |
3210 |
|
|
|
|
| 3 |
A' |
3177 |
3104 |
|
|
|
|
| 4 |
A' |
3152 |
3080 |
|
|
|
|
| 5 |
A' |
1533 |
1498 |
|
|
|
|
| 6 |
A' |
1472 |
1438 |
|
|
|
|
| 7 |
A' |
1398 |
1366 |
|
|
|
|
| 8 |
A' |
1240 |
1211 |
|
|
|
|
| 9 |
A' |
1120 |
1094 |
|
|
|
|
| 10 |
A' |
1009 |
986 |
|
|
|
|
| 11 |
A' |
498 |
487 |
|
|
|
|
| 12 |
A" |
1133 |
1107 |
|
|
|
|
| 13 |
A" |
818 |
799 |
|
|
|
|
| 14 |
A" |
713 |
697 |
|
|
|
|
| 15 |
A" |
509 |
497 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12212.9 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 11930.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.454 |
0.000 |
| C2 |
1.132 |
-0.396 |
0.000 |
| N3 |
-1.260 |
0.077 |
0.000 |
| H4 |
0.179 |
1.541 |
0.000 |
| H5 |
2.147 |
0.010 |
0.000 |
| H6 |
1.003 |
-1.486 |
0.000 |
| H7 |
-1.296 |
-0.956 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4155 | 1.3155 | 1.1016 | 2.1926 | 2.1840 | 1.9158 |
C2 | 1.4155 | | 2.4383 | 2.1591 | 1.0939 | 1.0971 | 2.4916 | N3 | 1.3155 | 2.4383 | | 2.0529 | 3.4082 | 2.7507 | 1.0343 | H4 | 1.1016 | 2.1591 | 2.0529 | | 2.4938 | 3.1372 | 2.9007 | H5 | 2.1926 | 1.0939 | 3.4082 | 2.4938 | | 1.8832 | 3.5766 | H6 | 2.1840 | 1.0971 | 2.7507 | 3.1372 | 1.8832 | | 2.3596 | H7 | 1.9158 | 2.4916 | 1.0343 | 2.9007 | 3.5766 | 2.3596 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.266 |
|
C1 |
C2 |
H6 |
120.208 |
| C1 |
N3 |
H7 |
108.642 |
|
C2 |
C1 |
N3 |
126.421 |
| C2 |
C1 |
H4 |
117.592 |
|
N3 |
C1 |
H4 |
115.987 |
| H5 |
C2 |
H6 |
118.526 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
