Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -168.205148 |
| Energy at 298.15K | |
| HF Energy | -167.648153 |
| Nuclear repulsion energy | 59.260059 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3520 |
3438 |
|
|
|
|
| 2 |
Σ |
2269 |
2216 |
|
|
|
|
| 3 |
Σ |
1281 |
1251 |
|
|
|
|
| 4 |
Π |
534 |
521 |
|
|
|
|
| 4 |
Π |
534 |
521 |
|
|
|
|
| 5 |
Π |
456i |
446i |
|
|
|
|
| 5 |
Π |
457i |
446i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3611.5 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 3528.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.205 |
| N2 |
0.000 |
0.000 |
-0.015 |
| C3 |
0.000 |
0.000 |
-1.208 |
| H4 |
0.000 |
0.000 |
-2.282 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2199 | 2.4126 | 3.4869 |
N2 | 1.2199 | | 1.1927 | 2.2670 | C3 | 2.4126 | 1.1927 | | 1.0743 | H4 | 3.4869 | 2.2670 | 1.0743 | |
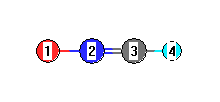 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
180.000 |
|
N2 |
C3 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -168.206912 |
| Energy at 298.15K | -168.207707 |
| HF Energy | -167.640594 |
| Nuclear repulsion energy | 59.118417 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3440 |
3360 |
|
|
|
|
| 2 |
A' |
2233 |
2181 |
|
|
|
|
| 3 |
A' |
1269 |
1240 |
|
|
|
|
| 4 |
A' |
625 |
611 |
|
|
|
|
| 5 |
A' |
497 |
485 |
|
|
|
|
| 6 |
A" |
549 |
536 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4305.9 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 4206.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.127 |
-0.408 |
0.000 |
| N2 |
0.000 |
0.049 |
0.000 |
| C3 |
1.182 |
0.306 |
0.000 |
| H4 |
1.929 |
1.087 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2166 | 2.4170 | 3.4024 |
N2 | 1.2166 | | 1.2093 | 2.1903 | C3 | 2.4170 | 1.2093 | | 1.0810 | H4 | 3.4024 | 2.1903 | 1.0810 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
170.169 |
|
N2 |
C3 |
H4 |
145.969 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
