Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -151.901104 |
| Energy at 298.15K | |
| HF Energy | -151.901104 |
| Nuclear repulsion energy | 52.391401 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3490 |
3344 |
124.18 |
50.94 |
0.23 |
0.37 |
| 2 |
Σ |
2132 |
2043 |
271.40 |
11.51 |
0.41 |
0.59 |
| 3 |
Σ |
1320 |
1265 |
23.77 |
46.20 |
0.14 |
0.25 |
| 4 |
Π |
564 |
541 |
0.67 |
0.28 |
0.75 |
0.86 |
| 4 |
Π |
515 |
494 |
4.55 |
0.64 |
0.75 |
0.86 |
| 5 |
Π |
358 |
343 |
26.67 |
0.45 |
0.75 |
0.86 |
| 5 |
Π |
428i |
410i |
120.25 |
7.35 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 3975.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 3809.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.022 |
| C2 |
0.000 |
0.000 |
-1.245 |
| O3 |
0.000 |
0.000 |
1.206 |
| H4 |
0.000 |
0.000 |
-2.313 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2666 | 1.1841 | 2.3350 |
C2 | 1.2666 | | 2.4507 | 1.0684 | O3 | 1.1841 | 2.4507 | | 3.5191 | H4 | 2.3350 | 1.0684 | 3.5191 | |
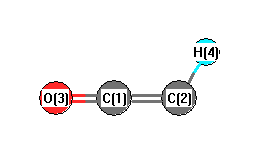 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.041 |
|
|
|
| 2 |
C |
0.686 |
|
|
|
| 3 |
O |
-0.325 |
|
|
|
| 4 |
H |
-0.402 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.178 |
2.178 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.246 |
0.000 |
0.000 |
| y |
0.000 |
-18.078 |
0.000 |
| z |
0.000 |
0.000 |
-14.868 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.226 |
0.000 |
0.000 |
| y |
0.000 |
-2.521 |
0.000 |
| z |
0.000 |
0.000 |
2.294 |
|
| Polar |
|---|
| 3z2-r2 | 4.588 |
|---|
| x2-y2 | 1.831 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.639 |
0.000 |
0.000 |
| y |
0.000 |
2.991 |
0.000 |
| z |
0.000 |
0.000 |
6.300 |
<r2> (average value of r
2) Å
2
| <r2> |
36.526 |
| (<r2>)1/2 |
6.044 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -151.904137 |
| Energy at 298.15K | |
| HF Energy | -151.904137 |
| Nuclear repulsion energy | 52.240125 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3352 |
3212 |
38.96 |
83.17 |
0.30 |
0.47 |
| 2 |
A' |
2115 |
2027 |
363.18 |
8.74 |
0.72 |
0.84 |
| 3 |
A' |
1262 |
1209 |
5.05 |
39.14 |
0.16 |
0.27 |
| 4 |
A' |
575 |
551 |
43.00 |
3.03 |
0.75 |
0.86 |
| 5 |
A' |
499 |
478 |
203.39 |
1.24 |
0.15 |
0.26 |
| 6 |
A" |
495 |
474 |
2.14 |
0.58 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4148.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 3975.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.053 |
0.000 |
| C2 |
1.065 |
-0.698 |
0.000 |
| O3 |
-1.064 |
0.544 |
0.000 |
| H4 |
2.124 |
-0.482 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.3030 | 1.1724 | 2.1907 |
C2 | 1.3030 | | 2.4652 | 1.0809 | O3 | 1.1724 | 2.4652 | | 3.3499 | H4 | 2.1907 | 1.0809 | 3.3499 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
133.325 |
|
C2 |
C1 |
O3 |
169.608 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.407 |
|
|
|
| 2 |
C |
0.132 |
|
|
|
| 3 |
O |
-0.268 |
|
|
|
| 4 |
H |
-0.271 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.609 |
0.414 |
0.000 |
1.661 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.333 |
1.008 |
0.000 |
| y |
1.008 |
-19.308 |
0.000 |
| z |
0.000 |
0.000 |
-16.411 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.527 |
1.008 |
0.000 |
| y |
1.008 |
-3.436 |
0.000 |
| z |
0.000 |
0.000 |
0.909 |
|
| Polar |
|---|
| 3z2-r2 | 1.819 |
|---|
| x2-y2 | 3.975 |
|---|
| xy | 1.008 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.401 |
-1.517 |
0.000 |
| y |
-1.517 |
4.098 |
0.000 |
| z |
0.000 |
0.000 |
3.041 |
<r2> (average value of r
2) Å
2
| <r2> |
36.552 |
| (<r2>)1/2 |
6.046 |
