Jump to
S1C2
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -151.560488 |
| Energy at 298.15K | |
| HF Energy | -151.102031 |
| Nuclear repulsion energy | 51.917438 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3510 |
3429 |
|
|
|
|
| 2 |
Σ |
2372 |
2317 |
|
|
|
|
| 3 |
Σ |
1399 |
1367 |
|
|
|
|
| 4 |
Π |
584 |
571 |
|
|
|
|
| 4 |
Π |
548 |
535 |
|
|
|
|
| 5 |
Π |
80i |
78i |
|
|
|
|
| 5 |
Π |
543i |
530i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3895.9 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 3805.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.029 |
| C2 |
0.000 |
0.000 |
-1.261 |
| O3 |
0.000 |
0.000 |
1.216 |
| H4 |
0.000 |
0.000 |
-2.335 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2897 | 1.1869 | 2.3644 |
C2 | 1.2897 | | 2.4765 | 1.0747 | O3 | 1.1869 | 2.4765 | | 3.5512 | H4 | 2.3644 | 1.0747 | 3.5512 | |
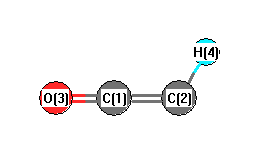 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -151.569642 |
| Energy at 298.15K | |
| HF Energy | -151.100752 |
| Nuclear repulsion energy | 51.373930 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3309 |
3232 |
|
|
|
|
| 2 |
A' |
2382 |
2327 |
|
|
|
|
| 3 |
A' |
1291 |
1261 |
|
|
|
|
| 4 |
A' |
680 |
664 |
|
|
|
|
| 5 |
A' |
609 |
595 |
|
|
|
|
| 6 |
A" |
509 |
498 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4390.2 cm
-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 4288.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.065 |
0.000 |
| C2 |
1.089 |
-0.733 |
0.000 |
| O3 |
-1.080 |
0.542 |
0.000 |
| H4 |
2.107 |
-0.330 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.3503 | 1.1808 | 2.1442 |
C2 | 1.3503 | | 2.5164 | 1.0951 | O3 | 1.1808 | 2.5164 | | 3.3049 | H4 | 2.1442 | 1.0951 | 3.3049 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
122.173 |
|
C2 |
C1 |
O3 |
167.577 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
