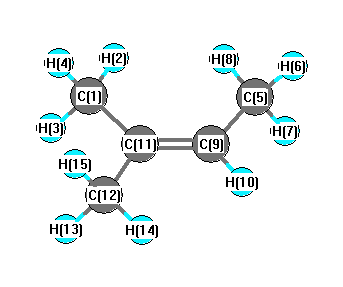
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3167 |
3035 |
44.40 |
|
|
|
| 2 |
A |
3161 |
3029 |
25.71 |
|
|
|
| 3 |
A |
3147 |
3016 |
10.42 |
|
|
|
| 4 |
A |
3145 |
3014 |
9.94 |
|
|
|
| 5 |
A |
3042 |
2916 |
15.86 |
|
|
|
| 6 |
A |
3039 |
2913 |
72.02 |
|
|
|
| 7 |
A |
3033 |
2906 |
28.75 |
|
|
|
| 8 |
A |
1768 |
1694 |
2.15 |
|
|
|
| 9 |
A |
1477 |
1415 |
9.05 |
|
|
|
| 10 |
A |
1474 |
1413 |
5.17 |
|
|
|
| 11 |
A |
1458 |
1397 |
1.68 |
|
|
|
| 12 |
A |
1410 |
1351 |
8.17 |
|
|
|
| 13 |
A |
1401 |
1343 |
0.11 |
|
|
|
| 14 |
A |
1394 |
1336 |
6.83 |
|
|
|
| 15 |
A |
1353 |
1296 |
1.62 |
|
|
|
| 16 |
A |
1246 |
1194 |
3.81 |
|
|
|
| 17 |
A |
1139 |
1091 |
10.48 |
|
|
|
| 18 |
A |
1066 |
1022 |
6.76 |
|
|
|
| 19 |
A |
969 |
929 |
3.86 |
|
|
|
| 20 |
A |
957 |
917 |
0.39 |
|
|
|
| 21 |
A |
787 |
754 |
1.80 |
|
|
|
| 22 |
A |
527 |
505 |
2.14 |
|
|
|
| 23 |
A |
383 |
367 |
0.52 |
|
|
|
| 24 |
A |
296 |
284 |
0.25 |
|
|
|
| 25 |
A |
3098 |
2968 |
17.51 |
|
|
|
| 26 |
A |
3095 |
2966 |
31.05 |
|
|
|
| 27 |
A |
3092 |
2963 |
14.19 |
|
|
|
| 28 |
A |
1467 |
1405 |
19.80 |
|
|
|
| 29 |
A |
1455 |
1394 |
1.83 |
|
|
|
| 30 |
A |
1445 |
1384 |
0.50 |
|
|
|
| 31 |
A |
1085 |
1040 |
0.03 |
|
|
|
| 32 |
A |
1046 |
1003 |
2.00 |
|
|
|
| 33 |
A |
1000 |
958 |
2.95 |
|
|
|
| 34 |
A |
818 |
784 |
12.78 |
|
|
|
| 35 |
A |
455 |
436 |
3.74 |
|
|
|
| 36 |
A |
267 |
255 |
1.88 |
|
|
|
| 37 |
A |
188 |
180 |
1.42 |
|
|
|
| 38 |
A |
122 |
117 |
0.63 |
|
|
|
| 39 |
A |
110 |
105 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29790.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28547.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.209 |
|
|
|
| 2 |
H |
-0.266 |
|
|
|
| 3 |
H |
0.041 |
|
|
|
| 4 |
H |
0.041 |
|
|
|
| 5 |
C |
0.042 |
|
|
|
| 6 |
H |
0.040 |
|
|
|
| 7 |
H |
0.040 |
|
|
|
| 8 |
H |
-0.269 |
|
|
|
| 9 |
C |
0.590 |
|
|
|
| 10 |
H |
-0.730 |
|
|
|
| 11 |
C |
0.210 |
|
|
|
| 12 |
C |
0.339 |
|
|
|
| 13 |
H |
-0.276 |
|
|
|
| 14 |
H |
-0.006 |
|
|
|
| 15 |
H |
-0.006 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.206 |
0.049 |
0.000 |
0.211 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.549 |
-0.321 |
0.000 |
| y |
-0.321 |
-32.355 |
0.000 |
| z |
0.000 |
0.000 |
-34.486 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.871 |
-0.321 |
0.000 |
| y |
-0.321 |
1.162 |
0.000 |
| z |
0.000 |
0.000 |
-2.034 |
|
| Polar |
|---|
| 3z2-r2 | -4.067 |
|---|
| x2-y2 | -0.194 |
|---|
| xy | -0.321 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.752 |
-0.949 |
0.000 |
| y |
-0.949 |
12.127 |
0.000 |
| z |
0.000 |
0.000 |
7.546 |
<r2> (average value of r
2) Å
2
| <r2> |
150.735 |
| (<r2>)1/2 |
12.277 |