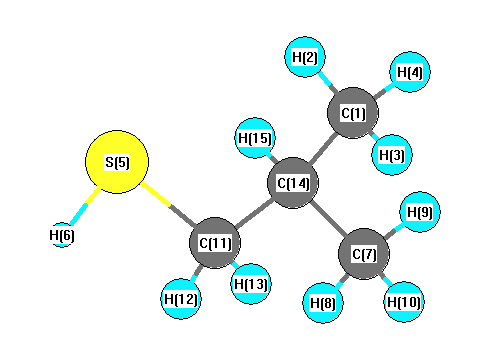
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3145 |
3014 |
28.20 |
|
|
|
| 2 |
A |
3140 |
3009 |
27.11 |
|
|
|
| 3 |
A |
3138 |
3007 |
26.82 |
|
|
|
| 4 |
A |
3130 |
2999 |
34.84 |
|
|
|
| 5 |
A |
3125 |
2995 |
8.63 |
|
|
|
| 6 |
A |
3073 |
2945 |
30.02 |
|
|
|
| 7 |
A |
3053 |
2926 |
17.06 |
|
|
|
| 8 |
A |
3049 |
2922 |
7.32 |
|
|
|
| 9 |
A |
3047 |
2920 |
24.90 |
|
|
|
| 10 |
A |
2714 |
2601 |
3.37 |
|
|
|
| 11 |
A |
1489 |
1427 |
10.84 |
|
|
|
| 12 |
A |
1485 |
1423 |
7.45 |
|
|
|
| 13 |
A |
1475 |
1414 |
4.29 |
|
|
|
| 14 |
A |
1465 |
1404 |
0.62 |
|
|
|
| 15 |
A |
1456 |
1395 |
3.77 |
|
|
|
| 16 |
A |
1406 |
1348 |
9.78 |
|
|
|
| 17 |
A |
1391 |
1333 |
8.56 |
|
|
|
| 18 |
A |
1363 |
1307 |
1.01 |
|
|
|
| 19 |
A |
1348 |
1292 |
1.57 |
|
|
|
| 20 |
A |
1266 |
1213 |
15.68 |
|
|
|
| 21 |
A |
1230 |
1179 |
5.45 |
|
|
|
| 22 |
A |
1198 |
1148 |
4.57 |
|
|
|
| 23 |
A |
1152 |
1104 |
2.62 |
|
|
|
| 24 |
A |
1081 |
1036 |
2.32 |
|
|
|
| 25 |
A |
985 |
944 |
2.01 |
|
|
|
| 26 |
A |
970 |
929 |
1.00 |
|
|
|
| 27 |
A |
925 |
886 |
1.74 |
|
|
|
| 28 |
A |
907 |
869 |
1.59 |
|
|
|
| 29 |
A |
872 |
835 |
2.24 |
|
|
|
| 30 |
A |
811 |
778 |
3.23 |
|
|
|
| 31 |
A |
738 |
707 |
1.68 |
|
|
|
| 32 |
A |
425 |
408 |
0.13 |
|
|
|
| 33 |
A |
394 |
378 |
0.96 |
|
|
|
| 34 |
A |
339 |
325 |
0.11 |
|
|
|
| 35 |
A |
248 |
238 |
0.13 |
|
|
|
| 36 |
A |
218 |
209 |
0.19 |
|
|
|
| 37 |
A |
204 |
196 |
1.62 |
|
|
|
| 38 |
A |
155 |
148 |
8.76 |
|
|
|
| 39 |
A |
73 |
70 |
9.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28841.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 27638.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.811 |
|
|
|
| 2 |
H |
-0.172 |
|
|
|
| 3 |
H |
-0.072 |
|
|
|
| 4 |
H |
-0.131 |
|
|
|
| 5 |
S |
0.130 |
|
|
|
| 6 |
H |
0.084 |
|
|
|
| 7 |
C |
0.724 |
|
|
|
| 8 |
H |
-0.151 |
|
|
|
| 9 |
H |
-0.143 |
|
|
|
| 10 |
H |
-0.049 |
|
|
|
| 11 |
C |
0.491 |
|
|
|
| 12 |
H |
-0.282 |
|
|
|
| 13 |
H |
-0.216 |
|
|
|
| 14 |
C |
-0.658 |
|
|
|
| 15 |
H |
-0.366 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.146 |
-1.059 |
0.535 |
1.649 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.557 |
2.383 |
-1.645 |
| y |
2.383 |
-40.463 |
-1.110 |
| z |
-1.645 |
-1.110 |
-41.781 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.435 |
2.383 |
-1.645 |
| y |
2.383 |
1.206 |
-1.110 |
| z |
-1.645 |
-1.110 |
-0.771 |
|
| Polar |
|---|
| 3z2-r2 | -1.542 |
|---|
| x2-y2 | -1.095 |
|---|
| xy | 2.383 |
|---|
| xz | -1.645 |
|---|
| yz | -1.110 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.914 |
-0.147 |
0.049 |
| y |
-0.147 |
10.415 |
-0.029 |
| z |
0.049 |
-0.029 |
9.432 |
<r2> (average value of r
2) Å
2
| <r2> |
201.322 |
| (<r2>)1/2 |
14.189 |