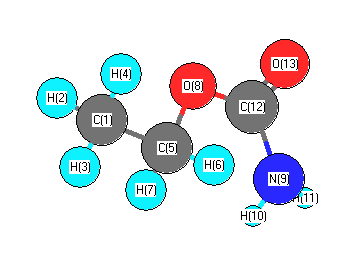
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3950 |
3597 |
53.79 |
|
|
|
| 2 |
A |
3824 |
3482 |
43.06 |
|
|
|
| 3 |
A |
3270 |
2978 |
45.18 |
|
|
|
| 4 |
A |
3254 |
2963 |
35.86 |
|
|
|
| 5 |
A |
3232 |
2943 |
13.13 |
|
|
|
| 6 |
A |
3183 |
2898 |
20.87 |
|
|
|
| 7 |
A |
3171 |
2887 |
44.88 |
|
|
|
| 8 |
A |
1955 |
1780 |
746.52 |
|
|
|
| 9 |
A |
1768 |
1610 |
153.48 |
|
|
|
| 10 |
A |
1631 |
1485 |
0.62 |
|
|
|
| 11 |
A |
1598 |
1456 |
0.35 |
|
|
|
| 12 |
A |
1578 |
1437 |
5.64 |
|
|
|
| 13 |
A |
1553 |
1414 |
14.48 |
|
|
|
| 14 |
A |
1512 |
1377 |
71.76 |
|
|
|
| 15 |
A |
1470 |
1339 |
390.99 |
|
|
|
| 16 |
A |
1413 |
1287 |
1.29 |
|
|
|
| 17 |
A |
1274 |
1160 |
5.27 |
|
|
|
| 18 |
A |
1258 |
1146 |
64.54 |
|
|
|
| 19 |
A |
1214 |
1106 |
140.24 |
|
|
|
| 20 |
A |
1188 |
1082 |
11.25 |
|
|
|
| 21 |
A |
1065 |
970 |
6.98 |
|
|
|
| 22 |
A |
935 |
851 |
12.13 |
|
|
|
| 23 |
A |
892 |
813 |
1.06 |
|
|
|
| 24 |
A |
868 |
790 |
47.91 |
|
|
|
| 25 |
A |
628 |
572 |
25.76 |
|
|
|
| 26 |
A |
600 |
546 |
8.48 |
|
|
|
| 27 |
A |
575 |
524 |
80.26 |
|
|
|
| 28 |
A |
406 |
370 |
13.35 |
|
|
|
| 29 |
A |
385 |
350 |
131.67 |
|
|
|
| 30 |
A |
280 |
255 |
0.49 |
|
|
|
| 31 |
A |
231 |
210 |
2.67 |
|
|
|
| 32 |
A |
106 |
97 |
0.60 |
|
|
|
| 33 |
A |
100 |
91 |
2.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25182.6 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 22931.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.161 |
|
|
|
| 2 |
H |
-0.023 |
|
|
|
| 3 |
H |
-0.011 |
|
|
|
| 4 |
H |
-0.017 |
|
|
|
| 5 |
C |
0.905 |
|
|
|
| 6 |
H |
-0.210 |
|
|
|
| 7 |
H |
-0.287 |
|
|
|
| 8 |
O |
-0.801 |
|
|
|
| 9 |
N |
-0.143 |
|
|
|
| 10 |
H |
0.022 |
|
|
|
| 11 |
H |
-0.028 |
|
|
|
| 12 |
C |
1.114 |
|
|
|
| 13 |
O |
-0.683 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.767 |
4.615 |
-0.622 |
5.416 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.593 |
-6.339 |
1.081 |
| y |
-6.339 |
-36.918 |
-1.472 |
| z |
1.081 |
-1.472 |
-36.396 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.064 |
-6.339 |
1.081 |
| y |
-6.339 |
-0.424 |
-1.472 |
| z |
1.081 |
-1.472 |
0.359 |
|
| Polar |
|---|
| 3z2-r2 | 0.719 |
|---|
| x2-y2 | 0.325 |
|---|
| xy | -6.339 |
|---|
| xz | 1.081 |
|---|
| yz | -1.472 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
190.388 |
| (<r2>)1/2 |
13.798 |