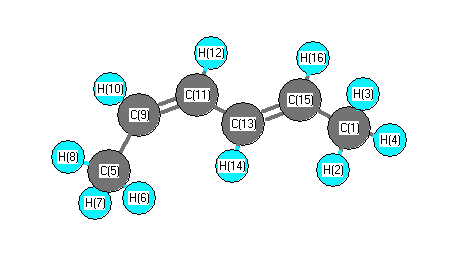
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3187 |
3054 |
51.54 |
|
|
|
| 2 |
A' |
3178 |
3046 |
6.34 |
|
|
|
| 3 |
A' |
3165 |
3033 |
6.46 |
|
|
|
| 4 |
A' |
3161 |
3029 |
21.86 |
|
|
|
| 5 |
A' |
3153 |
3022 |
2.47 |
|
|
|
| 6 |
A' |
3138 |
3007 |
6.58 |
|
|
|
| 7 |
A' |
3043 |
2916 |
32.40 |
|
|
|
| 8 |
A' |
3038 |
2911 |
42.74 |
|
|
|
| 9 |
A' |
1748 |
1675 |
4.85 |
|
|
|
| 10 |
A' |
1703 |
1632 |
5.33 |
|
|
|
| 11 |
A' |
1473 |
1412 |
12.49 |
|
|
|
| 12 |
A' |
1464 |
1403 |
17.76 |
|
|
|
| 13 |
A' |
1436 |
1376 |
4.08 |
|
|
|
| 14 |
A' |
1394 |
1336 |
3.11 |
|
|
|
| 15 |
A' |
1386 |
1328 |
3.87 |
|
|
|
| 16 |
A' |
1329 |
1274 |
1.12 |
|
|
|
| 17 |
A' |
1301 |
1246 |
1.57 |
|
|
|
| 18 |
A' |
1268 |
1215 |
0.78 |
|
|
|
| 19 |
A' |
1184 |
1134 |
0.04 |
|
|
|
| 20 |
A' |
1117 |
1070 |
0.01 |
|
|
|
| 21 |
A' |
1052 |
1008 |
0.71 |
|
|
|
| 22 |
A' |
940 |
901 |
25.31 |
|
|
|
| 23 |
A' |
933 |
895 |
15.39 |
|
|
|
| 24 |
A' |
601 |
576 |
5.52 |
|
|
|
| 25 |
A' |
467 |
448 |
0.28 |
|
|
|
| 26 |
A' |
317 |
304 |
2.30 |
|
|
|
| 27 |
A' |
150 |
144 |
0.54 |
|
|
|
| 28 |
A" |
3100 |
2971 |
16.93 |
|
|
|
| 29 |
A" |
3098 |
2969 |
16.01 |
|
|
|
| 30 |
A" |
1453 |
1392 |
12.15 |
|
|
|
| 31 |
A" |
1452 |
1392 |
1.36 |
|
|
|
| 32 |
A" |
1052 |
1008 |
0.40 |
|
|
|
| 33 |
A" |
1043 |
999 |
1.69 |
|
|
|
| 34 |
A" |
1023 |
980 |
22.73 |
|
|
|
| 35 |
A" |
981 |
940 |
15.39 |
|
|
|
| 36 |
A" |
853 |
817 |
13.01 |
|
|
|
| 37 |
A" |
724 |
694 |
18.49 |
|
|
|
| 38 |
A" |
446 |
427 |
3.84 |
|
|
|
| 39 |
A" |
219 |
210 |
0.00 |
|
|
|
| 40 |
A" |
203 |
195 |
1.21 |
|
|
|
| 41 |
A" |
126 |
121 |
0.18 |
|
|
|
| 42 |
A" |
102 |
97 |
1.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31100.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 29803.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.038 |
|
|
|
| 2 |
H |
-0.249 |
|
|
|
| 3 |
H |
0.040 |
|
|
|
| 4 |
H |
0.040 |
|
|
|
| 5 |
C |
0.083 |
|
|
|
| 6 |
H |
-0.281 |
|
|
|
| 7 |
H |
0.048 |
|
|
|
| 8 |
H |
0.048 |
|
|
|
| 9 |
C |
-0.094 |
|
|
|
| 10 |
H |
-0.608 |
|
|
|
| 11 |
C |
1.134 |
|
|
|
| 12 |
H |
-0.715 |
|
|
|
| 13 |
C |
1.936 |
|
|
|
| 14 |
H |
-0.745 |
|
|
|
| 15 |
C |
0.055 |
|
|
|
| 16 |
H |
-0.652 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.289 |
0.190 |
0.000 |
0.346 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.638 |
-0.725 |
0.000 |
| y |
-0.725 |
-34.481 |
0.000 |
| z |
0.000 |
0.000 |
-41.449 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.327 |
-0.725 |
0.000 |
| y |
-0.725 |
4.563 |
0.000 |
| z |
0.000 |
0.000 |
-5.890 |
|
| Polar |
|---|
| 3z2-r2 | -11.780 |
|---|
| x2-y2 | -2.157 |
|---|
| xy | -0.725 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.456 |
-1.191 |
0.000 |
| y |
-1.191 |
19.368 |
0.000 |
| z |
0.000 |
0.000 |
8.247 |
<r2> (average value of r
2) Å
2
| <r2> |
259.910 |
| (<r2>)1/2 |
16.122 |