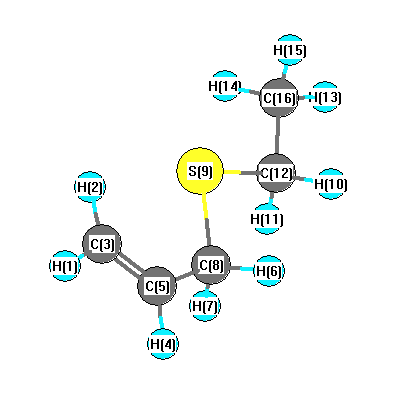
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3268 |
3132 |
13.06 |
|
|
|
| 2 |
A |
3176 |
3044 |
3.99 |
|
|
|
| 3 |
A |
3164 |
3032 |
14.09 |
|
|
|
| 4 |
A |
3144 |
3013 |
21.27 |
|
|
|
| 5 |
A |
3062 |
2934 |
26.47 |
|
|
|
| 6 |
A |
3061 |
2933 |
29.06 |
|
|
|
| 7 |
A |
3037 |
2910 |
26.73 |
|
|
|
| 8 |
A |
1730 |
1658 |
28.22 |
|
|
|
| 9 |
A |
1481 |
1419 |
2.77 |
|
|
|
| 10 |
A |
1465 |
1404 |
2.38 |
|
|
|
| 11 |
A |
1444 |
1383 |
10.60 |
|
|
|
| 12 |
A |
1437 |
1377 |
7.32 |
|
|
|
| 13 |
A |
1392 |
1334 |
4.02 |
|
|
|
| 14 |
A |
1325 |
1269 |
3.50 |
|
|
|
| 15 |
A |
1307 |
1253 |
2.41 |
|
|
|
| 16 |
A |
1268 |
1215 |
38.14 |
|
|
|
| 17 |
A |
1089 |
1044 |
1.53 |
|
|
|
| 18 |
A |
1066 |
1022 |
4.47 |
|
|
|
| 19 |
A |
1005 |
963 |
4.55 |
|
|
|
| 20 |
A |
953 |
913 |
3.38 |
|
|
|
| 21 |
A |
766 |
734 |
3.26 |
|
|
|
| 22 |
A |
705 |
676 |
1.19 |
|
|
|
| 23 |
A |
505 |
484 |
0.30 |
|
|
|
| 24 |
A |
334 |
320 |
0.65 |
|
|
|
| 25 |
A |
239 |
229 |
0.14 |
|
|
|
| 26 |
A |
121 |
116 |
0.23 |
|
|
|
| 27 |
A |
3152 |
3021 |
24.29 |
|
|
|
| 28 |
A |
3112 |
2982 |
5.14 |
|
|
|
| 29 |
A |
3079 |
2951 |
9.92 |
|
|
|
| 30 |
A |
1468 |
1407 |
9.11 |
|
|
|
| 31 |
A |
1256 |
1204 |
0.01 |
|
|
|
| 32 |
A |
1167 |
1119 |
2.36 |
|
|
|
| 33 |
A |
1036 |
993 |
0.00 |
|
|
|
| 34 |
A |
1014 |
972 |
10.15 |
|
|
|
| 35 |
A |
959 |
919 |
42.36 |
|
|
|
| 36 |
A |
906 |
868 |
1.34 |
|
|
|
| 37 |
A |
783 |
751 |
3.48 |
|
|
|
| 38 |
A |
552 |
529 |
13.63 |
|
|
|
| 39 |
A |
240 |
230 |
0.10 |
|
|
|
| 40 |
A |
150 |
144 |
0.55 |
|
|
|
| 41 |
A |
62 |
59 |
0.03 |
|
|
|
| 42 |
A |
35 |
34 |
1.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30256.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28995.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.424 |
|
|
|
| 2 |
H |
-0.377 |
|
|
|
| 3 |
C |
0.422 |
|
|
|
| 4 |
H |
-0.551 |
|
|
|
| 5 |
C |
0.970 |
|
|
|
| 6 |
H |
-0.052 |
|
|
|
| 7 |
H |
-0.052 |
|
|
|
| 8 |
C |
-0.061 |
|
|
|
| 9 |
S |
0.250 |
|
|
|
| 10 |
H |
-0.224 |
|
|
|
| 11 |
H |
-0.224 |
|
|
|
| 12 |
C |
0.187 |
|
|
|
| 13 |
H |
-0.098 |
|
|
|
| 14 |
H |
-0.108 |
|
|
|
| 15 |
H |
-0.108 |
|
|
|
| 16 |
C |
0.451 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.454 |
-1.164 |
0.000 |
1.863 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.218 |
1.460 |
0.000 |
| y |
1.460 |
-43.491 |
0.000 |
| z |
0.000 |
0.000 |
-48.634 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.845 |
1.460 |
0.000 |
| y |
1.460 |
1.935 |
0.000 |
| z |
0.000 |
0.000 |
-5.779 |
|
| Polar |
|---|
| 3z2-r2 | -11.558 |
|---|
| x2-y2 | 1.273 |
|---|
| xy | 1.460 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
15.676 |
1.508 |
0.000 |
| y |
1.508 |
13.051 |
0.000 |
| z |
0.000 |
0.000 |
9.705 |
<r2> (average value of r
2) Å
2
| <r2> |
295.039 |
| (<r2>)1/2 |
17.177 |