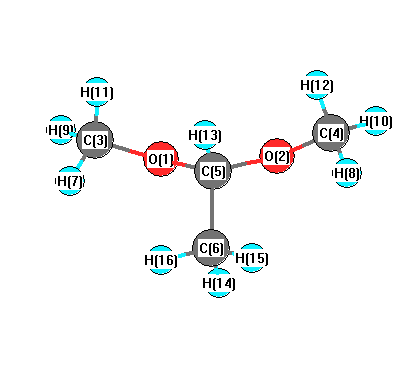
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3168 |
3036 |
17.51 |
|
|
|
| 2 |
A |
3160 |
3028 |
21.89 |
|
|
|
| 3 |
A |
3071 |
2943 |
76.76 |
|
|
|
| 4 |
A |
3065 |
2938 |
24.70 |
|
|
|
| 5 |
A |
3006 |
2881 |
62.36 |
|
|
|
| 6 |
A |
2932 |
2809 |
58.93 |
|
|
|
| 7 |
A |
1484 |
1422 |
1.75 |
|
|
|
| 8 |
A |
1483 |
1421 |
7.54 |
|
|
|
| 9 |
A |
1464 |
1402 |
18.01 |
|
|
|
| 10 |
A |
1450 |
1389 |
3.63 |
|
|
|
| 11 |
A |
1396 |
1338 |
60.08 |
|
|
|
| 12 |
A |
1356 |
1300 |
1.05 |
|
|
|
| 13 |
A |
1263 |
1210 |
23.91 |
|
|
|
| 14 |
A |
1185 |
1136 |
34.37 |
|
|
|
| 15 |
A |
1167 |
1118 |
0.99 |
|
|
|
| 16 |
A |
1125 |
1078 |
8.24 |
|
|
|
| 17 |
A |
1089 |
1044 |
42.33 |
|
|
|
| 18 |
A |
851 |
815 |
6.08 |
|
|
|
| 19 |
A |
534 |
512 |
6.46 |
|
|
|
| 20 |
A |
447 |
429 |
9.79 |
|
|
|
| 21 |
A |
251 |
241 |
2.75 |
|
|
|
| 22 |
A |
174 |
167 |
1.84 |
|
|
|
| 23 |
A |
52 |
50 |
3.84 |
|
|
|
| 24 |
A |
3160 |
3028 |
18.57 |
|
|
|
| 25 |
A |
3150 |
3018 |
31.36 |
|
|
|
| 26 |
A |
3067 |
2939 |
13.47 |
|
|
|
| 27 |
A |
3004 |
2879 |
69.46 |
|
|
|
| 28 |
A |
1487 |
1425 |
5.66 |
|
|
|
| 29 |
A |
1471 |
1410 |
3.16 |
|
|
|
| 30 |
A |
1464 |
1403 |
9.05 |
|
|
|
| 31 |
A |
1451 |
1391 |
0.73 |
|
|
|
| 32 |
A |
1372 |
1314 |
47.05 |
|
|
|
| 33 |
A |
1210 |
1159 |
199.65 |
|
|
|
| 34 |
A |
1170 |
1121 |
193.09 |
|
|
|
| 35 |
A |
1164 |
1116 |
33.34 |
|
|
|
| 36 |
A |
1087 |
1042 |
12.02 |
|
|
|
| 37 |
A |
919 |
881 |
7.03 |
|
|
|
| 38 |
A |
470 |
451 |
1.90 |
|
|
|
| 39 |
A |
347 |
332 |
0.66 |
|
|
|
| 40 |
A |
229 |
219 |
0.00 |
|
|
|
| 41 |
A |
194 |
186 |
0.07 |
|
|
|
| 42 |
A |
20 |
20 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30803.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 29518.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.656 |
|
|
|
| 2 |
O |
-0.656 |
|
|
|
| 3 |
C |
0.716 |
|
|
|
| 4 |
C |
0.716 |
|
|
|
| 5 |
C |
1.242 |
|
|
|
| 6 |
C |
0.385 |
|
|
|
| 7 |
H |
-0.109 |
|
|
|
| 8 |
H |
-0.109 |
|
|
|
| 9 |
H |
-0.293 |
|
|
|
| 10 |
H |
-0.293 |
|
|
|
| 11 |
H |
-0.106 |
|
|
|
| 12 |
H |
-0.106 |
|
|
|
| 13 |
H |
-0.304 |
|
|
|
| 14 |
H |
-0.094 |
|
|
|
| 15 |
H |
-0.167 |
|
|
|
| 16 |
H |
-0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.930 |
2.232 |
0.000 |
2.418 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.686 |
-1.324 |
0.000 |
| y |
-1.324 |
-40.214 |
0.000 |
| z |
0.000 |
0.000 |
-32.734 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.212 |
-1.324 |
0.000 |
| y |
-1.324 |
-4.504 |
0.000 |
| z |
0.000 |
0.000 |
6.716 |
|
| Polar |
|---|
| 3z2-r2 | 13.431 |
|---|
| x2-y2 | 1.528 |
|---|
| xy | -1.324 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.835 |
-0.063 |
0.000 |
| y |
-0.063 |
8.213 |
0.000 |
| z |
0.000 |
0.000 |
10.923 |
<r2> (average value of r
2) Å
2
| <r2> |
198.983 |
| (<r2>)1/2 |
14.106 |