Jump to
S1C2
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -269.312951 |
| Energy at 298.15K | -269.319629 |
| HF Energy | -269.312951 |
| Nuclear repulsion energy | 222.660364 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3315 |
3177 |
0.25 |
|
|
|
| 2 |
A' |
3289 |
3152 |
0.51 |
|
|
|
| 3 |
A' |
3277 |
3140 |
4.28 |
|
|
|
| 4 |
A' |
3170 |
3038 |
9.12 |
|
|
|
| 5 |
A' |
3057 |
2929 |
29.66 |
|
|
|
| 6 |
A' |
1665 |
1595 |
15.01 |
|
|
|
| 7 |
A' |
1569 |
1504 |
22.46 |
|
|
|
| 8 |
A' |
1475 |
1413 |
5.69 |
|
|
|
| 9 |
A' |
1421 |
1362 |
6.84 |
|
|
|
| 10 |
A' |
1399 |
1340 |
1.06 |
|
|
|
| 11 |
A' |
1268 |
1215 |
14.74 |
|
|
|
| 12 |
A' |
1249 |
1197 |
2.23 |
|
|
|
| 13 |
A' |
1185 |
1136 |
19.08 |
|
|
|
| 14 |
A' |
1124 |
1078 |
3.95 |
|
|
|
| 15 |
A' |
1037 |
993 |
25.80 |
|
|
|
| 16 |
A' |
989 |
948 |
2.03 |
|
|
|
| 17 |
A' |
946 |
907 |
15.71 |
|
|
|
| 18 |
A' |
902 |
864 |
3.10 |
|
|
|
| 19 |
A' |
666 |
639 |
0.64 |
|
|
|
| 20 |
A' |
339 |
325 |
2.28 |
|
|
|
| 21 |
A" |
3123 |
2993 |
12.40 |
|
|
|
| 22 |
A" |
1453 |
1393 |
7.39 |
|
|
|
| 23 |
A" |
1051 |
1007 |
1.03 |
|
|
|
| 24 |
A" |
863 |
827 |
0.23 |
|
|
|
| 25 |
A" |
803 |
769 |
20.15 |
|
|
|
| 26 |
A" |
740 |
709 |
65.41 |
|
|
|
| 27 |
A" |
642 |
616 |
1.15 |
|
|
|
| 28 |
A" |
617 |
591 |
7.49 |
|
|
|
| 29 |
A" |
246 |
236 |
5.92 |
|
|
|
| 30 |
A" |
135 |
129 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21507.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 20610.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.035 |
-0.240 |
0.000 |
| C2 |
0.000 |
0.646 |
0.000 |
| C3 |
0.502 |
-1.491 |
0.000 |
| C4 |
-1.183 |
-0.032 |
0.000 |
| C5 |
-0.855 |
-1.427 |
0.000 |
| C6 |
0.352 |
2.086 |
0.000 |
| H7 |
1.211 |
-2.310 |
0.000 |
| H8 |
-2.172 |
0.412 |
0.000 |
| H9 |
-1.539 |
-2.269 |
0.000 |
| H10 |
-0.563 |
2.687 |
0.000 |
| H11 |
0.942 |
2.353 |
0.887 |
| H12 |
0.942 |
2.353 |
-0.887 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3626 | 1.3595 | 2.2273 | 2.2316 | 2.4247 | 2.0768 | 3.2721 | 3.2767 | 3.3345 | 2.7427 | 2.7427 |
C2 | 1.3626 | | 2.1958 | 1.3635 | 2.2432 | 1.4825 | 3.1943 | 2.1843 | 3.2962 | 2.1165 | 2.1421 | 2.1421 | C3 | 1.3595 | 2.1958 | | 2.2291 | 1.3586 | 3.5807 | 1.0824 | 3.2823 | 2.1840 | 4.3115 | 3.9700 | 3.9700 | C4 | 2.2273 | 1.3635 | 2.2291 | | 1.4336 | 2.6160 | 3.3040 | 1.0841 | 2.2650 | 2.7884 | 3.3155 | 3.3155 | C5 | 2.2316 | 2.2432 | 1.3586 | 1.4336 | | 3.7155 | 2.2460 | 2.2625 | 1.0840 | 4.1245 | 4.2793 | 4.2793 | C6 | 2.4247 | 1.4825 | 3.5807 | 2.6160 | 3.7155 | | 4.4791 | 3.0287 | 4.7479 | 1.0944 | 1.0983 | 1.0983 | H7 | 2.0768 | 3.1943 | 1.0824 | 3.3040 | 2.2460 | 4.4791 | | 4.3415 | 2.7495 | 5.3017 | 4.7542 | 4.7542 | H8 | 3.2721 | 2.1843 | 3.2823 | 1.0841 | 2.2625 | 3.0287 | 4.3415 | | 2.7548 | 2.7859 | 3.7752 | 3.7752 | H9 | 3.2767 | 3.2962 | 2.1840 | 2.2650 | 1.0840 | 4.7479 | 2.7495 | 2.7548 | | 5.0506 | 5.3204 | 5.3204 | H10 | 3.3345 | 2.1165 | 4.3115 | 2.7884 | 4.1245 | 1.0944 | 5.3017 | 2.7859 | 5.0506 | | 1.7786 | 1.7786 | H11 | 2.7427 | 2.1421 | 3.9700 | 3.3155 | 4.2793 | 1.0983 | 4.7542 | 3.7752 | 5.3204 | 1.7786 | | 1.7739 | H12 | 2.7427 | 2.1421 | 3.9700 | 3.3155 | 4.2793 | 1.0983 | 4.7542 | 3.7752 | 5.3204 | 1.7786 | 1.7739 | |
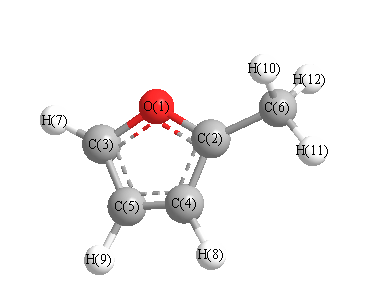 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
109.574 |
|
O1 |
C2 |
C6 |
116.850 |
| O1 |
C3 |
C5 |
110.370 |
|
O1 |
C3 |
H7 |
116.069 |
| C2 |
O1 |
C3 |
107.538 |
|
C2 |
C4 |
C5 |
106.610 |
| C2 |
C4 |
H8 |
125.983 |
|
C2 |
C6 |
H10 |
109.520 |
| C2 |
C6 |
H11 |
111.333 |
|
C2 |
C6 |
H12 |
111.333 |
| C3 |
C5 |
C6 |
73.727 |
|
C3 |
C5 |
H9 |
126.415 |
| C4 |
C2 |
C6 |
133.575 |
|
C4 |
C5 |
H9 |
127.677 |
| C5 |
C3 |
H7 |
133.561 |
|
C5 |
C4 |
H8 |
127.407 |
| H10 |
C6 |
H11 |
108.419 |
|
H10 |
C6 |
H12 |
108.419 |
| H11 |
C6 |
H12 |
107.720 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.594 |
|
|
|
| 2 |
C |
-0.174 |
|
|
|
| 3 |
C |
0.357 |
|
|
|
| 4 |
C |
1.091 |
|
|
|
| 5 |
C |
0.572 |
|
|
|
| 6 |
C |
0.187 |
|
|
|
| 7 |
H |
-0.435 |
|
|
|
| 8 |
H |
-0.508 |
|
|
|
| 9 |
H |
-0.417 |
|
|
|
| 10 |
H |
-0.079 |
|
|
|
| 11 |
H |
0.001 |
|
|
|
| 12 |
H |
0.001 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.473 |
0.437 |
0.000 |
0.644 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.358 |
-0.445 |
0.000 |
| y |
-0.445 |
-30.430 |
0.000 |
| z |
0.000 |
0.000 |
-38.182 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.052 |
-0.445 |
0.000 |
| y |
-0.445 |
5.840 |
0.000 |
| z |
0.000 |
0.000 |
-5.787 |
|
| Polar |
|---|
| 3z2-r2 | -11.575 |
|---|
| x2-y2 | -3.928 |
|---|
| xy | -0.445 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.472 |
0.327 |
0.000 |
| y |
0.327 |
11.565 |
0.000 |
| z |
0.000 |
0.000 |
6.553 |
<r2> (average value of r
2) Å
2
| <r2> |
140.864 |
| (<r2>)1/2 |
11.869 |
