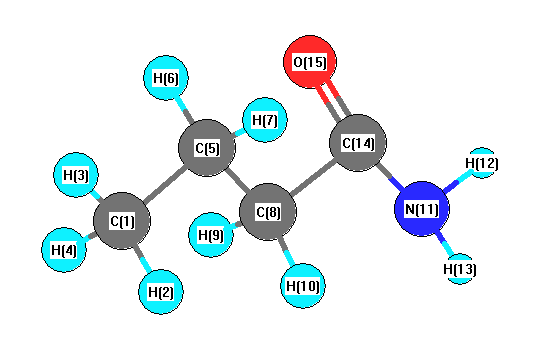
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3960 |
3606 |
49.34 |
|
|
|
| 2 |
A |
3819 |
3477 |
51.97 |
|
|
|
| 3 |
A |
3250 |
2960 |
67.88 |
|
|
|
| 4 |
A |
3233 |
2944 |
59.58 |
|
|
|
| 5 |
A |
3209 |
2922 |
22.24 |
|
|
|
| 6 |
A |
3200 |
2914 |
11.56 |
|
|
|
| 7 |
A |
3193 |
2908 |
17.57 |
|
|
|
| 8 |
A |
3163 |
2880 |
32.13 |
|
|
|
| 9 |
A |
3155 |
2873 |
27.92 |
|
|
|
| 10 |
A |
1921 |
1749 |
411.86 |
|
|
|
| 11 |
A |
1762 |
1604 |
125.98 |
|
|
|
| 12 |
A |
1608 |
1465 |
6.63 |
|
|
|
| 13 |
A |
1594 |
1452 |
6.08 |
|
|
|
| 14 |
A |
1588 |
1446 |
1.26 |
|
|
|
| 15 |
A |
1571 |
1430 |
7.50 |
|
|
|
| 16 |
A |
1554 |
1415 |
68.20 |
|
|
|
| 17 |
A |
1520 |
1384 |
6.49 |
|
|
|
| 18 |
A |
1457 |
1327 |
52.08 |
|
|
|
| 19 |
A |
1425 |
1298 |
4.19 |
|
|
|
| 20 |
A |
1354 |
1233 |
107.93 |
|
|
|
| 21 |
A |
1349 |
1229 |
1.09 |
|
|
|
| 22 |
A |
1222 |
1113 |
1.24 |
|
|
|
| 23 |
A |
1210 |
1102 |
1.35 |
|
|
|
| 24 |
A |
1162 |
1058 |
5.77 |
|
|
|
| 25 |
A |
1116 |
1017 |
2.82 |
|
|
|
| 26 |
A |
991 |
903 |
2.58 |
|
|
|
| 27 |
A |
951 |
866 |
1.60 |
|
|
|
| 28 |
A |
910 |
828 |
1.03 |
|
|
|
| 29 |
A |
802 |
730 |
6.08 |
|
|
|
| 30 |
A |
674 |
614 |
9.29 |
|
|
|
| 31 |
A |
659 |
600 |
22.11 |
|
|
|
| 32 |
A |
539 |
490 |
5.41 |
|
|
|
| 33 |
A |
462 |
421 |
2.59 |
|
|
|
| 34 |
A |
365 |
333 |
4.00 |
|
|
|
| 35 |
A |
262 |
238 |
0.18 |
|
|
|
| 36 |
A |
205 |
187 |
222.94 |
|
|
|
| 37 |
A |
193 |
176 |
10.47 |
|
|
|
| 38 |
A |
105 |
96 |
0.87 |
|
|
|
| 39 |
A |
25 |
23 |
2.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30369.0 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 27654.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.481 |
|
|
|
| 2 |
H |
-0.082 |
|
|
|
| 3 |
H |
-0.058 |
|
|
|
| 4 |
H |
-0.082 |
|
|
|
| 5 |
C |
0.262 |
|
|
|
| 6 |
H |
-0.177 |
|
|
|
| 7 |
H |
-0.148 |
|
|
|
| 8 |
C |
0.238 |
|
|
|
| 9 |
H |
-0.104 |
|
|
|
| 10 |
H |
-0.043 |
|
|
|
| 11 |
N |
-0.169 |
|
|
|
| 12 |
H |
-0.037 |
|
|
|
| 13 |
H |
-0.053 |
|
|
|
| 14 |
C |
0.715 |
|
|
|
| 15 |
O |
-0.742 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.081 |
-3.771 |
0.248 |
3.780 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.099 |
-6.485 |
0.636 |
| y |
-6.485 |
-40.108 |
-0.466 |
| z |
0.636 |
-0.466 |
-38.287 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.098 |
-6.485 |
0.636 |
| y |
-6.485 |
-3.914 |
-0.466 |
| z |
0.636 |
-0.466 |
-1.183 |
|
| Polar |
|---|
| 3z2-r2 | -2.367 |
|---|
| x2-y2 | 6.008 |
|---|
| xy | -6.485 |
|---|
| xz | 0.636 |
|---|
| yz | -0.466 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
217.714 |
| (<r2>)1/2 |
14.755 |