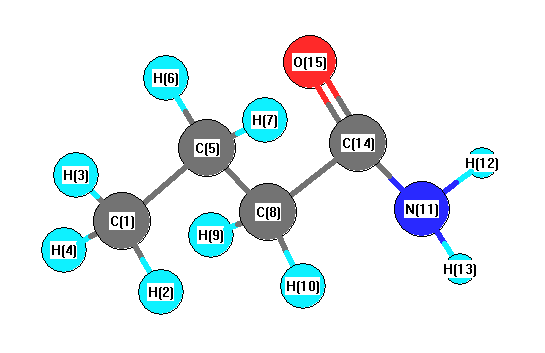
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3771 |
3614 |
37.92 |
|
|
|
| 2 |
A |
3626 |
3475 |
35.51 |
|
|
|
| 3 |
A |
3141 |
3010 |
42.59 |
|
|
|
| 4 |
A |
3138 |
3007 |
35.80 |
|
|
|
| 5 |
A |
3114 |
2984 |
6.48 |
|
|
|
| 6 |
A |
3090 |
2961 |
15.21 |
|
|
|
| 7 |
A |
3073 |
2945 |
22.42 |
|
|
|
| 8 |
A |
3049 |
2922 |
31.62 |
|
|
|
| 9 |
A |
3044 |
2917 |
13.83 |
|
|
|
| 10 |
A |
1787 |
1713 |
330.74 |
|
|
|
| 11 |
A |
1621 |
1553 |
125.84 |
|
|
|
| 12 |
A |
1489 |
1427 |
8.22 |
|
|
|
| 13 |
A |
1478 |
1416 |
7.61 |
|
|
|
| 14 |
A |
1472 |
1411 |
2.03 |
|
|
|
| 15 |
A |
1446 |
1386 |
13.85 |
|
|
|
| 16 |
A |
1424 |
1364 |
83.63 |
|
|
|
| 17 |
A |
1398 |
1339 |
2.93 |
|
|
|
| 18 |
A |
1355 |
1298 |
30.89 |
|
|
|
| 19 |
A |
1313 |
1258 |
20.54 |
|
|
|
| 20 |
A |
1256 |
1203 |
36.97 |
|
|
|
| 21 |
A |
1241 |
1189 |
3.22 |
|
|
|
| 22 |
A |
1140 |
1092 |
0.76 |
|
|
|
| 23 |
A |
1113 |
1067 |
1.26 |
|
|
|
| 24 |
A |
1085 |
1040 |
0.73 |
|
|
|
| 25 |
A |
1063 |
1019 |
3.15 |
|
|
|
| 26 |
A |
941 |
901 |
2.27 |
|
|
|
| 27 |
A |
880 |
844 |
0.73 |
|
|
|
| 28 |
A |
856 |
820 |
2.43 |
|
|
|
| 29 |
A |
746 |
715 |
5.25 |
|
|
|
| 30 |
A |
665 |
638 |
4.30 |
|
|
|
| 31 |
A |
612 |
586 |
12.29 |
|
|
|
| 32 |
A |
524 |
502 |
6.66 |
|
|
|
| 33 |
A |
426 |
408 |
2.32 |
|
|
|
| 34 |
A |
342 |
328 |
2.54 |
|
|
|
| 35 |
A |
245 |
235 |
0.38 |
|
|
|
| 36 |
A |
201 |
193 |
175.55 |
|
|
|
| 37 |
A |
182 |
175 |
11.56 |
|
|
|
| 38 |
A |
86 |
83 |
1.41 |
|
|
|
| 39 |
A |
36 |
34 |
3.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28732.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 27534.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.445 |
|
|
|
| 2 |
H |
-0.120 |
|
|
|
| 3 |
H |
-0.120 |
|
|
|
| 4 |
H |
-0.120 |
|
|
|
| 5 |
C |
0.556 |
|
|
|
| 6 |
H |
-0.268 |
|
|
|
| 7 |
H |
-0.245 |
|
|
|
| 8 |
C |
0.441 |
|
|
|
| 9 |
H |
-0.304 |
|
|
|
| 10 |
H |
-0.139 |
|
|
|
| 11 |
N |
-0.029 |
|
|
|
| 12 |
H |
-0.054 |
|
|
|
| 13 |
H |
-0.055 |
|
|
|
| 14 |
C |
0.621 |
|
|
|
| 15 |
O |
-0.610 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.081 |
-3.636 |
-0.161 |
3.640 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.488 |
-6.454 |
-0.598 |
| y |
-6.454 |
-39.549 |
-0.009 |
| z |
-0.598 |
-0.009 |
-38.176 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.375 |
-6.454 |
-0.598 |
| y |
-6.454 |
-3.717 |
-0.009 |
| z |
-0.598 |
-0.009 |
-1.658 |
|
| Polar |
|---|
| 3z2-r2 | -3.316 |
|---|
| x2-y2 | 6.061 |
|---|
| xy | -6.454 |
|---|
| xz | -0.598 |
|---|
| yz | -0.009 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.158 |
-0.308 |
-0.189 |
| y |
-0.308 |
9.531 |
-0.004 |
| z |
-0.189 |
-0.004 |
7.559 |
<r2> (average value of r
2) Å
2
| <r2> |
217.212 |
| (<r2>)1/2 |
14.738 |