Jump to
S1C2
Vibrational Frequencies calculated at M06-2X/aug-cc-pVDZ
Geometric Data calculated at M06-2X/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at M06-2X/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -323.546423 |
| Energy at 298.15K | -323.555641 |
| HF Energy | -323.546423 |
| Nuclear repulsion energy | 244.193001 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3152 |
3152 |
17.16 |
|
|
|
| 2 |
A |
3141 |
3141 |
24.02 |
|
|
|
| 3 |
A |
3135 |
3135 |
30.44 |
|
|
|
| 4 |
A |
3107 |
3107 |
10.32 |
|
|
|
| 5 |
A |
3085 |
3085 |
18.69 |
|
|
|
| 6 |
A |
3062 |
3062 |
22.33 |
|
|
|
| 7 |
A |
3056 |
3056 |
12.97 |
|
|
|
| 8 |
A |
1817 |
1817 |
236.14 |
|
|
|
| 9 |
A |
1483 |
1483 |
14.36 |
|
|
|
| 10 |
A |
1480 |
1480 |
4.37 |
|
|
|
| 11 |
A |
1467 |
1467 |
6.10 |
|
|
|
| 12 |
A |
1454 |
1454 |
1.82 |
|
|
|
| 13 |
A |
1404 |
1404 |
7.63 |
|
|
|
| 14 |
A |
1397 |
1397 |
6.46 |
|
|
|
| 15 |
A |
1358 |
1358 |
0.20 |
|
|
|
| 16 |
A |
1301 |
1301 |
22.22 |
|
|
|
| 17 |
A |
1279 |
1279 |
3.22 |
|
|
|
| 18 |
A |
1184 |
1184 |
10.94 |
|
|
|
| 19 |
A |
1126 |
1126 |
54.46 |
|
|
|
| 20 |
A |
1114 |
1114 |
7.14 |
|
|
|
| 21 |
A |
1019 |
1019 |
4.72 |
|
|
|
| 22 |
A |
940 |
940 |
302.58 |
|
|
|
| 23 |
A |
895 |
895 |
23.14 |
|
|
|
| 24 |
A |
878 |
878 |
143.50 |
|
|
|
| 25 |
A |
784 |
784 |
4.37 |
|
|
|
| 26 |
A |
654 |
654 |
6.93 |
|
|
|
| 27 |
A |
464 |
464 |
4.99 |
|
|
|
| 28 |
A |
396 |
396 |
2.53 |
|
|
|
| 29 |
A |
296 |
296 |
0.51 |
|
|
|
| 30 |
A |
270 |
270 |
0.34 |
|
|
|
| 31 |
A |
201 |
201 |
0.56 |
|
|
|
| 32 |
A |
110 |
110 |
0.68 |
|
|
|
| 33 |
A |
79 |
79 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23292.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 23292.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.179 |
-0.916 |
0.183 |
| C2 |
-1.749 |
0.460 |
-0.319 |
| C3 |
-0.401 |
0.892 |
0.224 |
| O4 |
0.576 |
-0.037 |
-0.282 |
| N5 |
1.783 |
0.206 |
0.312 |
| O6 |
2.556 |
-0.552 |
-0.146 |
| H7 |
-3.165 |
-1.182 |
-0.214 |
| H8 |
-2.240 |
-0.929 |
1.279 |
| H9 |
-1.464 |
-1.685 |
-0.130 |
| H10 |
-1.708 |
0.473 |
-1.416 |
| H11 |
-2.479 |
1.222 |
-0.014 |
| H12 |
-0.131 |
1.900 |
-0.115 |
| H13 |
-0.377 |
0.858 |
1.321 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
N5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5265 | 2.5362 | 2.9289 | 4.1199 | 4.7603 | 1.0960 | 1.0980 | 1.0958 | 2.1694 | 2.1681 | 3.4946 | 2.7729 |
C2 | 1.5265 | | 1.5160 | 2.3774 | 3.5967 | 4.4255 | 2.1710 | 2.1732 | 2.1723 | 1.0979 | 1.0984 | 2.1752 | 2.1745 | C3 | 2.5362 | 1.5160 | | 1.4393 | 2.2904 | 3.3109 | 3.4836 | 2.7947 | 2.8101 | 2.1381 | 2.1178 | 1.0971 | 1.0982 | O4 | 2.9289 | 2.3774 | 1.4393 | | 1.3673 | 2.0507 | 3.9129 | 3.3406 | 2.6270 | 2.5996 | 3.3148 | 2.0680 | 2.0686 | N5 | 4.1199 | 3.5967 | 2.2904 | 1.3673 | | 1.1754 | 5.1662 | 4.2901 | 3.7837 | 3.9038 | 4.3935 | 2.5910 | 2.4717 | O6 | 4.7603 | 4.4255 | 3.3109 | 2.0507 | 1.1754 | | 5.7562 | 5.0169 | 4.1767 | 4.5652 | 5.3399 | 3.6373 | 3.5696 | H7 | 1.0960 | 2.1710 | 3.4836 | 3.9129 | 5.1662 | 5.7562 | | 1.7753 | 1.7761 | 2.5111 | 2.5081 | 4.3259 | 3.7805 | H8 | 1.0980 | 2.1732 | 2.7947 | 3.3406 | 4.2901 | 5.0169 | 1.7753 | | 1.7775 | 3.0837 | 2.5213 | 3.7937 | 2.5813 | H9 | 1.0958 | 2.1723 | 2.8101 | 2.6270 | 3.7837 | 4.1767 | 1.7761 | 1.7775 | | 2.5236 | 3.0815 | 3.8247 | 3.1232 | H10 | 2.1694 | 1.0979 | 2.1381 | 2.5996 | 3.9038 | 4.5652 | 2.5111 | 3.0837 | 2.5236 | | 1.7665 | 2.4926 | 3.0673 | H11 | 2.1681 | 1.0984 | 2.1178 | 3.3148 | 4.3935 | 5.3399 | 2.5081 | 2.5213 | 3.0815 | 1.7665 | | 2.4460 | 2.5166 | H12 | 3.4946 | 2.1752 | 1.0971 | 2.0680 | 2.5910 | 3.6373 | 4.3259 | 3.7937 | 3.8247 | 2.4926 | 2.4460 | | 1.7915 | H13 | 2.7729 | 2.1745 | 1.0982 | 2.0686 | 2.4717 | 3.5696 | 3.7805 | 2.5813 | 3.1232 | 3.0673 | 2.5166 | 1.7915 | |
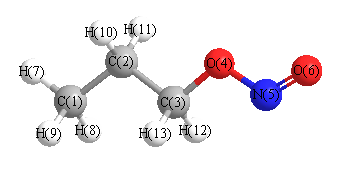 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.943 |
|
C1 |
C2 |
H10 |
110.442 |
| C1 |
C2 |
H11 |
110.311 |
|
C2 |
C1 |
H7 |
110.679 |
| C2 |
C1 |
H8 |
110.735 |
|
C2 |
C1 |
H9 |
110.795 |
| C2 |
C3 |
O4 |
107.089 |
|
C2 |
C3 |
H12 |
111.700 |
| C2 |
C3 |
H13 |
111.575 |
|
C3 |
C2 |
H10 |
108.709 |
| C3 |
C2 |
H11 |
107.123 |
|
C3 |
O4 |
N5 |
109.367 |
| O4 |
C3 |
H12 |
108.492 |
|
O4 |
C3 |
H13 |
108.472 |
| O4 |
N5 |
O6 |
107.271 |
|
H7 |
C1 |
H8 |
108.029 |
| H7 |
C1 |
H9 |
108.260 |
|
H8 |
C1 |
H9 |
108.243 |
| H10 |
C2 |
H11 |
107.090 |
|
H12 |
C3 |
H13 |
109.390 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.821 |
|
|
|
| 2 |
C |
0.811 |
|
|
|
| 3 |
C |
1.119 |
|
|
|
| 4 |
O |
-0.498 |
|
|
|
| 5 |
N |
0.449 |
|
|
|
| 6 |
O |
-0.434 |
|
|
|
| 7 |
H |
-0.262 |
|
|
|
| 8 |
H |
-0.269 |
|
|
|
| 9 |
H |
-0.256 |
|
|
|
| 10 |
H |
-0.381 |
|
|
|
| 11 |
H |
-0.305 |
|
|
|
| 12 |
H |
-0.430 |
|
|
|
| 13 |
H |
-0.365 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.673 |
-0.870 |
0.751 |
2.909 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.173 |
0.321 |
-0.824 |
| y |
0.321 |
-35.425 |
-1.146 |
| z |
-0.824 |
-1.146 |
-37.186 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.867 |
0.321 |
-0.824 |
| y |
0.321 |
2.754 |
-1.146 |
| z |
-0.824 |
-1.146 |
0.114 |
|
| Polar |
|---|
| 3z2-r2 | 0.227 |
|---|
| x2-y2 | -3.747 |
|---|
| xy | 0.321 |
|---|
| xz | -0.824 |
|---|
| yz | -1.146 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.954 |
-0.939 |
0.047 |
| y |
-0.939 |
8.116 |
0.020 |
| z |
0.047 |
0.020 |
7.015 |
<r2> (average value of r
2) Å
2
| <r2> |
186.595 |
| (<r2>)1/2 |
13.660 |
