Jump to
S1C2
Energy calculated at CCSD=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.722539 |
| Energy at 298.15K | -93.723840 |
| HF Energy | -93.411745 |
| Nuclear repulsion energy | 27.889295 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3354 |
3354 |
0.85 |
|
|
|
| 2 |
A' |
3026 |
3026 |
42.67 |
|
|
|
| 3 |
A' |
1802 |
1802 |
29.68 |
|
|
|
| 4 |
A' |
1054 |
1054 |
132.15 |
|
|
|
| 5 |
A' |
877 |
877 |
88.34 |
|
|
|
| 6 |
A" |
895 |
895 |
2.78 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5504.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5504.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
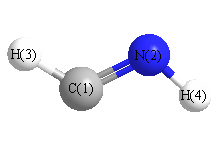
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.114 |
0.644 |
0.000 |
| N2 |
0.114 |
-0.599 |
0.000 |
| H3 |
-0.683 |
1.414 |
0.000 |
| H4 |
-0.795 |
-1.087 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2428 | 1.1074 | 1.9548 |
N2 | 1.2428 | | 2.1642 | 1.0315 | H3 | 1.1074 | 2.1642 | | 2.5028 | H4 | 1.9548 | 1.0315 | 2.5028 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
118.229 |
|
H3 |
C1 |
N2 |
134.025 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.730642 |
| Energy at 298.15K | -93.731990 |
| HF Energy | -93.417186 |
| Nuclear repulsion energy | 27.841983 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3467 |
3467 |
7.52 |
|
|
|
| 2 |
A' |
3074 |
3074 |
22.21 |
|
|
|
| 3 |
A' |
1770 |
1770 |
31.70 |
|
|
|
| 4 |
A' |
1208 |
1208 |
13.66 |
|
|
|
| 5 |
A' |
919 |
919 |
209.77 |
|
|
|
| 6 |
A" |
980 |
980 |
97.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5708.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5708.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.001 |
0.655 |
0.000 |
| N2 |
0.001 |
-0.595 |
0.000 |
| H3 |
0.907 |
1.289 |
0.000 |
| H4 |
-0.917 |
-1.054 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2505 | 1.1055 | 1.9402 |
N2 | 1.2505 | | 2.0905 | 1.0262 | H3 | 1.1055 | 2.0905 | | 2.9690 | H4 | 1.9402 | 1.0262 | 2.9690 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
116.559 |
|
H3 |
C1 |
N2 |
124.967 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
