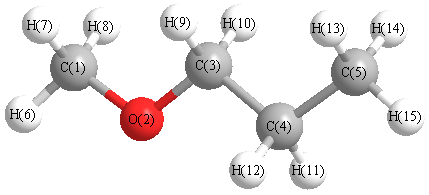
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3153 |
3022 |
28.13 |
|
|
|
| 2 |
A' |
3142 |
3011 |
31.62 |
|
|
|
| 3 |
A' |
3069 |
2941 |
26.49 |
|
|
|
| 4 |
A' |
3053 |
2926 |
29.73 |
|
|
|
| 5 |
A' |
2997 |
2872 |
83.36 |
|
|
|
| 6 |
A' |
2972 |
2848 |
44.22 |
|
|
|
| 7 |
A' |
1503 |
1441 |
6.17 |
|
|
|
| 8 |
A' |
1487 |
1425 |
4.55 |
|
|
|
| 9 |
A' |
1479 |
1418 |
6.50 |
|
|
|
| 10 |
A' |
1472 |
1411 |
2.52 |
|
|
|
| 11 |
A' |
1457 |
1397 |
0.23 |
|
|
|
| 12 |
A' |
1415 |
1356 |
13.35 |
|
|
|
| 13 |
A' |
1398 |
1339 |
3.84 |
|
|
|
| 14 |
A' |
1318 |
1264 |
3.09 |
|
|
|
| 15 |
A' |
1229 |
1178 |
51.75 |
|
|
|
| 16 |
A' |
1188 |
1138 |
134.63 |
|
|
|
| 17 |
A' |
1130 |
1082 |
10.53 |
|
|
|
| 18 |
A' |
1079 |
1034 |
3.02 |
|
|
|
| 19 |
A' |
1004 |
962 |
22.97 |
|
|
|
| 20 |
A' |
912 |
874 |
6.14 |
|
|
|
| 21 |
A' |
441 |
422 |
0.67 |
|
|
|
| 22 |
A' |
410 |
393 |
2.87 |
|
|
|
| 23 |
A' |
193 |
185 |
1.30 |
|
|
|
| 24 |
A" |
3134 |
3003 |
59.59 |
|
|
|
| 25 |
A" |
3109 |
2979 |
1.99 |
|
|
|
| 26 |
A" |
3053 |
2926 |
55.86 |
|
|
|
| 27 |
A" |
3006 |
2881 |
57.41 |
|
|
|
| 28 |
A" |
1479 |
1417 |
7.16 |
|
|
|
| 29 |
A" |
1466 |
1405 |
8.16 |
|
|
|
| 30 |
A" |
1305 |
1251 |
0.27 |
|
|
|
| 31 |
A" |
1259 |
1206 |
1.83 |
|
|
|
| 32 |
A" |
1187 |
1137 |
4.56 |
|
|
|
| 33 |
A" |
1159 |
1111 |
0.23 |
|
|
|
| 34 |
A" |
893 |
856 |
1.27 |
|
|
|
| 35 |
A" |
756 |
725 |
1.73 |
|
|
|
| 36 |
A" |
237 |
227 |
0.79 |
|
|
|
| 37 |
A" |
224 |
215 |
2.60 |
|
|
|
| 38 |
A" |
105 |
101 |
2.41 |
|
|
|
| 39 |
A" |
104 |
99 |
0.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29987.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28736.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.709 |
|
|
|
| 2 |
O |
-0.566 |
|
|
|
| 3 |
C |
0.829 |
|
|
|
| 4 |
C |
0.504 |
|
|
|
| 5 |
C |
0.391 |
|
|
|
| 6 |
H |
-0.308 |
|
|
|
| 7 |
H |
-0.082 |
|
|
|
| 8 |
H |
-0.082 |
|
|
|
| 9 |
H |
-0.294 |
|
|
|
| 10 |
H |
-0.294 |
|
|
|
| 11 |
H |
-0.218 |
|
|
|
| 12 |
H |
-0.218 |
|
|
|
| 13 |
H |
-0.116 |
|
|
|
| 14 |
H |
-0.116 |
|
|
|
| 15 |
H |
-0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.294 |
1.039 |
0.000 |
1.080 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.615 |
-2.169 |
0.000 |
| y |
-2.169 |
-33.874 |
0.000 |
| z |
0.000 |
0.000 |
-33.117 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.880 |
-2.169 |
0.000 |
| y |
-2.169 |
-1.508 |
0.000 |
| z |
0.000 |
0.000 |
-0.372 |
|
| Polar |
|---|
| 3z2-r2 | -0.744 |
|---|
| x2-y2 | 2.259 |
|---|
| xy | -2.169 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.313 |
-0.443 |
0.000 |
| y |
-0.443 |
8.138 |
0.000 |
| z |
0.000 |
0.000 |
7.622 |
<r2> (average value of r
2) Å
2
| <r2> |
180.719 |
| (<r2>)1/2 |
13.443 |