return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (Methyl propyl ether)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-233.428315
Energy at 298.15K 
HF Energy-232.166513
Nuclear repulsion energy187.039750
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3157 3084        
2 A' 3139 3067        
3 A' 3067 2997        
4 A' 3054 2984        
5 A' 3005 2936        
6 A' 2971 2902        
7 A' 1576 1540        
8 A' 1554 1518        
9 A' 1549 1513        
10 A' 1540 1505        
11 A' 1512 1477        
12 A' 1461 1428        
13 A' 1459 1425        
14 A' 1360 1328        
15 A' 1249 1220        
16 A' 1183 1156        
17 A' 1148 1122        
18 A' 1086 1061        
19 A' 995 972        
20 A' 928 907        
21 A' 446 435        
22 A' 415 406        
23 A' 196 191        
24 A" 3134 3062        
25 A" 3110 3038        
26 A" 3062 2992        
27 A" 3007 2937        
28 A" 1548 1512        
29 A" 1531 1496        
30 A" 1340 1309        
31 A" 1295 1265        
32 A" 1227 1199        
33 A" 1187 1159        
34 A" 923 901        
35 A" 779 761        
36 A" 246 241        
37 A" 237 232        
38 A" 114 111        
39 A" 100 98        

Unscaled Zero Point Vibrational Energy (zpe) 30443.4 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 29740.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.64446 0.06997 0.06634

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.334 0.658 0.000
O2 -1.284 -0.311 0.000
C3 0.000 0.324 0.000
C4 1.083 -0.753 0.000
C5 2.494 -0.137 0.000
H6 -3.286 0.104 0.000
H7 -2.282 1.303 0.903
H8 -2.282 1.303 -0.903
H9 0.098 0.975 0.899
H10 0.098 0.975 -0.899
H11 0.943 -1.394 -0.890
H12 0.943 -1.394 0.890
H13 2.654 0.494 0.894
H14 2.654 0.494 -0.894
H15 3.269 -0.924 0.000

Atom - Atom Distances (Å)
  C1 O2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.42932.35783.69734.89321.10181.11041.11042.61172.61173.96763.96765.07045.07045.8223
O21.42931.43232.40783.78182.04542.10152.10152.09032.09032.63112.63114.11764.11764.5939
C32.35781.43231.52802.53653.29382.64182.64181.11381.11382.15282.15282.80602.80603.4993
C43.69732.40781.52801.53984.45294.04554.04552.18312.18311.10591.10592.19632.19632.1927
C54.89323.78182.53651.53985.78565.06945.06942.79062.79062.18632.18631.10621.10621.1042
H61.10182.04543.29384.45295.78561.80571.80573.60803.60804.57434.57436.02026.02026.6358
H71.11042.10152.64184.04555.06941.80571.80542.40203.00254.57024.20385.00195.31506.0487
H81.11042.10152.64184.04555.06941.80571.80543.00252.40204.20384.57025.31505.00196.0487
H92.61172.09031.11382.18312.79063.60802.40203.00251.79773.08662.51512.60153.15973.8041
H102.61172.09031.11382.18312.79063.60803.00252.40201.79772.51513.08663.15972.60153.8041
H113.96762.63112.15281.10592.18634.57434.57024.20383.08662.51511.78083.11112.54822.5351
H123.96762.63112.15281.10592.18634.57434.20384.57022.51513.08661.78082.54823.11112.5351
H135.07044.11762.80602.19631.10626.02025.00195.31502.60153.15973.11112.54821.78881.7853
H145.07044.11762.80602.19631.10626.02025.31505.00193.15972.60152.54823.11111.78881.7853
H155.82234.59393.49932.19271.10426.63586.04876.04873.80413.80412.53512.53511.78531.7853

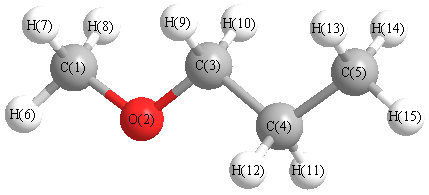
picture of Methyl propyl ether state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C3 110.960 O2 C1 H6 107.110
O2 C1 H7 111.049 O2 C1 H8 111.049
O2 C3 C4 108.807 O2 C3 H9 109.732
O2 C3 H10 109.732 C3 C4 C5 111.548
C3 C4 H11 108.580 C3 C4 H12 108.580
C4 C3 H9 110.473 C4 C3 H10 110.473
C4 C5 H13 111.155 C4 C5 H14 111.155
C4 C5 H15 110.984 C5 C4 H11 110.379
C5 C4 H12 110.379 H6 C1 H7 109.419
H6 C1 H8 109.419 H7 C1 H8 108.763
H9 C3 H10 107.612 H11 C4 H12 107.249
H13 C5 H14 107.910 H13 C5 H15 107.740
H14 C5 H15 107.740
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability