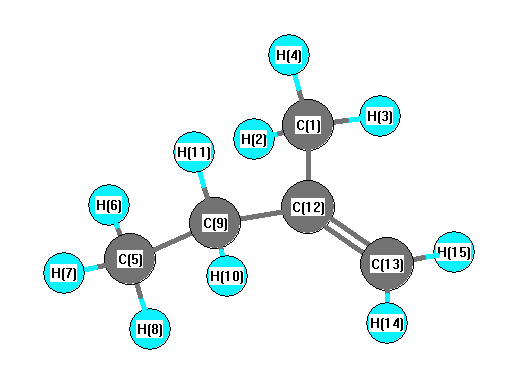
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3253 |
3117 |
17.96 |
|
|
|
| 2 |
A |
3161 |
3029 |
9.67 |
|
|
|
| 3 |
A |
3152 |
3020 |
20.29 |
|
|
|
| 4 |
A |
3145 |
3014 |
38.03 |
|
|
|
| 5 |
A |
3141 |
3010 |
32.45 |
|
|
|
| 6 |
A |
3108 |
2978 |
18.39 |
|
|
|
| 7 |
A |
3103 |
2974 |
12.50 |
|
|
|
| 8 |
A |
3059 |
2932 |
29.65 |
|
|
|
| 9 |
A |
3044 |
2917 |
27.25 |
|
|
|
| 10 |
A |
3038 |
2911 |
25.36 |
|
|
|
| 11 |
A |
1733 |
1661 |
25.41 |
|
|
|
| 12 |
A |
1488 |
1426 |
3.07 |
|
|
|
| 13 |
A |
1480 |
1418 |
7.11 |
|
|
|
| 14 |
A |
1476 |
1414 |
13.29 |
|
|
|
| 15 |
A |
1465 |
1404 |
11.47 |
|
|
|
| 16 |
A |
1453 |
1392 |
3.64 |
|
|
|
| 17 |
A |
1435 |
1375 |
0.20 |
|
|
|
| 18 |
A |
1396 |
1338 |
7.61 |
|
|
|
| 19 |
A |
1389 |
1331 |
3.31 |
|
|
|
| 20 |
A |
1349 |
1292 |
0.90 |
|
|
|
| 21 |
A |
1307 |
1252 |
1.00 |
|
|
|
| 22 |
A |
1266 |
1213 |
0.91 |
|
|
|
| 23 |
A |
1109 |
1063 |
1.11 |
|
|
|
| 24 |
A |
1093 |
1047 |
4.26 |
|
|
|
| 25 |
A |
1047 |
1004 |
0.65 |
|
|
|
| 26 |
A |
1027 |
984 |
0.81 |
|
|
|
| 27 |
A |
992 |
951 |
0.98 |
|
|
|
| 28 |
A |
953 |
913 |
1.79 |
|
|
|
| 29 |
A |
928 |
889 |
45.41 |
|
|
|
| 30 |
A |
800 |
767 |
1.50 |
|
|
|
| 31 |
A |
783 |
750 |
1.33 |
|
|
|
| 32 |
A |
720 |
690 |
0.29 |
|
|
|
| 33 |
A |
538 |
516 |
8.06 |
|
|
|
| 34 |
A |
434 |
416 |
1.67 |
|
|
|
| 35 |
A |
401 |
384 |
1.00 |
|
|
|
| 36 |
A |
264 |
253 |
0.28 |
|
|
|
| 37 |
A |
219 |
210 |
0.61 |
|
|
|
| 38 |
A |
181 |
174 |
0.20 |
|
|
|
| 39 |
A |
66 |
64 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29997.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28746.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.178 |
|
|
|
| 2 |
H |
-0.036 |
|
|
|
| 3 |
H |
-0.235 |
|
|
|
| 4 |
H |
-0.006 |
|
|
|
| 5 |
C |
0.445 |
|
|
|
| 6 |
H |
-0.168 |
|
|
|
| 7 |
H |
-0.151 |
|
|
|
| 8 |
H |
-0.138 |
|
|
|
| 9 |
C |
0.505 |
|
|
|
| 10 |
H |
-0.302 |
|
|
|
| 11 |
H |
-0.199 |
|
|
|
| 12 |
C |
0.302 |
|
|
|
| 13 |
C |
0.891 |
|
|
|
| 14 |
H |
-0.540 |
|
|
|
| 15 |
H |
-0.546 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.411 |
0.389 |
0.136 |
0.582 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.243 |
0.680 |
-0.873 |
| y |
0.680 |
-32.634 |
0.162 |
| z |
-0.873 |
0.162 |
-34.739 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.444 |
0.680 |
-0.873 |
| y |
0.680 |
1.357 |
0.162 |
| z |
-0.873 |
0.162 |
-1.800 |
|
| Polar |
|---|
| 3z2-r2 | -3.601 |
|---|
| x2-y2 | -0.609 |
|---|
| xy | 0.680 |
|---|
| xz | -0.873 |
|---|
| yz | 0.162 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.813 |
-0.865 |
-0.404 |
| y |
-0.865 |
10.129 |
0.266 |
| z |
-0.404 |
0.266 |
8.050 |
<r2> (average value of r
2) Å
2
| <r2> |
143.347 |
| (<r2>)1/2 |
11.973 |