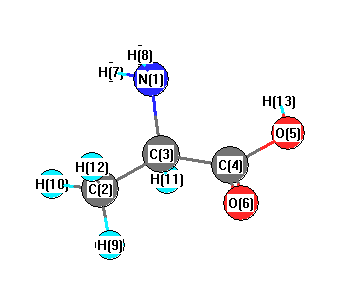
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3638 |
3487 |
12.70 |
|
|
|
| 2 |
A |
3544 |
3397 |
0.43 |
|
|
|
| 3 |
A |
3452 |
3308 |
313.33 |
|
|
|
| 4 |
A |
3176 |
3044 |
5.64 |
|
|
|
| 5 |
A |
3142 |
3011 |
24.04 |
|
|
|
| 6 |
A |
3079 |
2950 |
10.31 |
|
|
|
| 7 |
A |
3060 |
2932 |
15.10 |
|
|
|
| 8 |
A |
1862 |
1784 |
337.14 |
|
|
|
| 9 |
A |
1649 |
1581 |
33.69 |
|
|
|
| 10 |
A |
1479 |
1417 |
26.92 |
|
|
|
| 11 |
A |
1471 |
1409 |
2.91 |
|
|
|
| 12 |
A |
1443 |
1383 |
377.82 |
|
|
|
| 13 |
A |
1394 |
1336 |
14.73 |
|
|
|
| 14 |
A |
1379 |
1322 |
3.45 |
|
|
|
| 15 |
A |
1306 |
1252 |
8.12 |
|
|
|
| 16 |
A |
1248 |
1196 |
20.54 |
|
|
|
| 17 |
A |
1222 |
1171 |
7.46 |
|
|
|
| 18 |
A |
1165 |
1116 |
10.09 |
|
|
|
| 19 |
A |
1095 |
1049 |
36.18 |
|
|
|
| 20 |
A |
1016 |
973 |
0.53 |
|
|
|
| 21 |
A |
941 |
902 |
26.77 |
|
|
|
| 22 |
A |
912 |
874 |
61.18 |
|
|
|
| 23 |
A |
864 |
828 |
108.77 |
|
|
|
| 24 |
A |
815 |
781 |
5.22 |
|
|
|
| 25 |
A |
742 |
711 |
7.20 |
|
|
|
| 26 |
A |
576 |
552 |
2.97 |
|
|
|
| 27 |
A |
534 |
512 |
1.76 |
|
|
|
| 28 |
A |
405 |
388 |
4.29 |
|
|
|
| 29 |
A |
354 |
339 |
11.97 |
|
|
|
| 30 |
A |
280 |
268 |
13.06 |
|
|
|
| 31 |
A |
269 |
257 |
16.59 |
|
|
|
| 32 |
A |
223 |
214 |
0.22 |
|
|
|
| 33 |
A |
58 |
56 |
0.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23895.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 22899.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.347 |
|
|
|
| 2 |
C |
0.712 |
|
|
|
| 3 |
C |
-0.111 |
|
|
|
| 4 |
C |
0.848 |
|
|
|
| 5 |
O |
-0.480 |
|
|
|
| 6 |
O |
-0.589 |
|
|
|
| 7 |
H |
0.062 |
|
|
|
| 8 |
H |
0.049 |
|
|
|
| 9 |
H |
-0.115 |
|
|
|
| 10 |
H |
-0.074 |
|
|
|
| 11 |
H |
-0.083 |
|
|
|
| 12 |
H |
-0.108 |
|
|
|
| 13 |
H |
0.236 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
5.117 |
2.025 |
0.359 |
5.515 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.764 |
-0.321 |
-0.142 |
| y |
-0.321 |
-37.773 |
0.373 |
| z |
-0.142 |
0.373 |
-33.866 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.945 |
-0.321 |
-0.142 |
| y |
-0.321 |
-0.958 |
0.373 |
| z |
-0.142 |
0.373 |
4.902 |
|
| Polar |
|---|
| 3z2-r2 | 9.805 |
|---|
| x2-y2 | -1.991 |
|---|
| xy | -0.321 |
|---|
| xz | -0.142 |
|---|
| yz | 0.373 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.657 |
0.188 |
0.061 |
| y |
0.188 |
8.855 |
-0.087 |
| z |
0.061 |
-0.087 |
6.503 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |