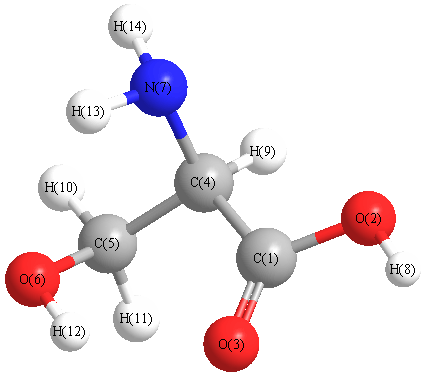
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3813 |
3654 |
65.91 |
|
|
|
| 2 |
A |
3794 |
3636 |
88.97 |
|
|
|
| 3 |
A |
3629 |
3477 |
7.42 |
|
|
|
| 4 |
A |
3535 |
3387 |
2.25 |
|
|
|
| 5 |
A |
3146 |
3015 |
17.02 |
|
|
|
| 6 |
A |
3081 |
2953 |
13.30 |
|
|
|
| 7 |
A |
3020 |
2894 |
47.51 |
|
|
|
| 8 |
A |
1813 |
1737 |
282.62 |
|
|
|
| 9 |
A |
1651 |
1582 |
33.42 |
|
|
|
| 10 |
A |
1485 |
1423 |
5.52 |
|
|
|
| 11 |
A |
1431 |
1372 |
32.31 |
|
|
|
| 12 |
A |
1401 |
1342 |
43.83 |
|
|
|
| 13 |
A |
1386 |
1328 |
35.62 |
|
|
|
| 14 |
A |
1370 |
1313 |
6.87 |
|
|
|
| 15 |
A |
1312 |
1258 |
6.43 |
|
|
|
| 16 |
A |
1242 |
1190 |
20.17 |
|
|
|
| 17 |
A |
1194 |
1144 |
21.42 |
|
|
|
| 18 |
A |
1176 |
1127 |
197.39 |
|
|
|
| 19 |
A |
1146 |
1098 |
14.29 |
|
|
|
| 20 |
A |
1117 |
1071 |
115.32 |
|
|
|
| 21 |
A |
1037 |
994 |
34.95 |
|
|
|
| 22 |
A |
998 |
956 |
2.71 |
|
|
|
| 23 |
A |
860 |
824 |
130.53 |
|
|
|
| 24 |
A |
816 |
782 |
45.39 |
|
|
|
| 25 |
A |
749 |
718 |
20.04 |
|
|
|
| 26 |
A |
625 |
599 |
88.53 |
|
|
|
| 27 |
A |
586 |
561 |
11.97 |
|
|
|
| 28 |
A |
534 |
512 |
179.72 |
|
|
|
| 29 |
A |
512 |
491 |
8.54 |
|
|
|
| 30 |
A |
449 |
430 |
8.81 |
|
|
|
| 31 |
A |
295 |
283 |
5.49 |
|
|
|
| 32 |
A |
284 |
273 |
12.48 |
|
|
|
| 33 |
A |
264 |
253 |
25.23 |
|
|
|
| 34 |
A |
208 |
199 |
3.30 |
|
|
|
| 35 |
A |
151 |
145 |
4.58 |
|
|
|
| 36 |
A |
31 |
30 |
1.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25069.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 24024.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.844 |
|
|
|
| 2 |
O |
-0.518 |
|
|
|
| 3 |
O |
-0.584 |
|
|
|
| 4 |
C |
0.100 |
|
|
|
| 5 |
C |
1.176 |
|
|
|
| 6 |
O |
-0.506 |
|
|
|
| 7 |
N |
-0.170 |
|
|
|
| 8 |
H |
0.184 |
|
|
|
| 9 |
H |
-0.227 |
|
|
|
| 10 |
H |
-0.238 |
|
|
|
| 11 |
H |
-0.172 |
|
|
|
| 12 |
H |
0.128 |
|
|
|
| 13 |
H |
0.000 |
|
|
|
| 14 |
H |
-0.016 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.059 |
0.698 |
0.541 |
3.184 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.667 |
-2.083 |
-1.385 |
| y |
-2.083 |
-44.462 |
1.711 |
| z |
-1.385 |
1.711 |
-40.885 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.006 |
-2.083 |
-1.385 |
| y |
-2.083 |
-3.186 |
1.711 |
| z |
-1.385 |
1.711 |
2.180 |
|
| Polar |
|---|
| 3z2-r2 | 4.360 |
|---|
| x2-y2 | 2.795 |
|---|
| xy | -2.083 |
|---|
| xz | -1.385 |
|---|
| yz | 1.711 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.487 |
0.015 |
-0.222 |
| y |
0.015 |
9.439 |
-0.154 |
| z |
-0.222 |
-0.154 |
7.307 |
<r2> (average value of r
2) Å
2
| <r2> |
215.117 |
| (<r2>)1/2 |
14.667 |