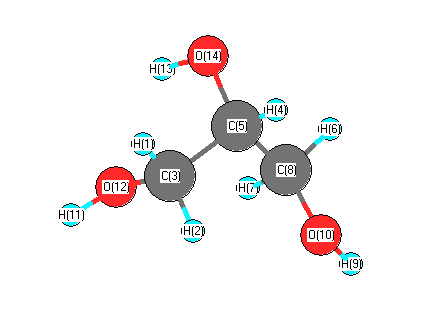
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3908 |
3745 |
51.61 |
|
|
|
| 2 |
A |
3901 |
3738 |
41.59 |
|
|
|
| 3 |
A |
3822 |
3663 |
50.31 |
|
|
|
| 4 |
A |
3108 |
2978 |
25.53 |
|
|
|
| 5 |
A |
3102 |
2973 |
48.46 |
|
|
|
| 6 |
A |
3084 |
2956 |
18.88 |
|
|
|
| 7 |
A |
3047 |
2920 |
33.53 |
|
|
|
| 8 |
A |
3037 |
2910 |
48.43 |
|
|
|
| 9 |
A |
1500 |
1437 |
8.92 |
|
|
|
| 10 |
A |
1490 |
1428 |
11.27 |
|
|
|
| 11 |
A |
1447 |
1386 |
0.71 |
|
|
|
| 12 |
A |
1446 |
1385 |
18.04 |
|
|
|
| 13 |
A |
1407 |
1349 |
36.92 |
|
|
|
| 14 |
A |
1361 |
1304 |
4.39 |
|
|
|
| 15 |
A |
1317 |
1262 |
42.34 |
|
|
|
| 16 |
A |
1281 |
1227 |
12.98 |
|
|
|
| 17 |
A |
1245 |
1193 |
10.86 |
|
|
|
| 18 |
A |
1220 |
1169 |
1.71 |
|
|
|
| 19 |
A |
1190 |
1141 |
26.55 |
|
|
|
| 20 |
A |
1148 |
1100 |
41.49 |
|
|
|
| 21 |
A |
1122 |
1075 |
90.23 |
|
|
|
| 22 |
A |
1091 |
1045 |
99.41 |
|
|
|
| 23 |
A |
1069 |
1024 |
56.88 |
|
|
|
| 24 |
A |
1004 |
962 |
19.41 |
|
|
|
| 25 |
A |
946 |
907 |
7.95 |
|
|
|
| 26 |
A |
837 |
802 |
13.26 |
|
|
|
| 27 |
A |
650 |
623 |
25.85 |
|
|
|
| 28 |
A |
485 |
465 |
5.95 |
|
|
|
| 29 |
A |
434 |
416 |
109.49 |
|
|
|
| 30 |
A |
402 |
385 |
2.15 |
|
|
|
| 31 |
A |
271 |
259 |
4.54 |
|
|
|
| 32 |
A |
252 |
241 |
17.38 |
|
|
|
| 33 |
A |
227 |
218 |
98.69 |
|
|
|
| 34 |
A |
217 |
208 |
82.26 |
|
|
|
| 35 |
A |
154 |
148 |
27.21 |
|
|
|
| 36 |
A |
92 |
89 |
4.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26155.5 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 25064.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.223 |
|
|
|
| 2 |
H |
-0.207 |
|
|
|
| 3 |
C |
1.006 |
|
|
|
| 4 |
H |
-0.360 |
|
|
|
| 5 |
C |
0.542 |
|
|
|
| 6 |
H |
-0.157 |
|
|
|
| 7 |
H |
-0.210 |
|
|
|
| 8 |
C |
0.915 |
|
|
|
| 9 |
H |
0.117 |
|
|
|
| 10 |
O |
-0.509 |
|
|
|
| 11 |
H |
0.125 |
|
|
|
| 12 |
O |
-0.577 |
|
|
|
| 13 |
H |
0.146 |
|
|
|
| 14 |
O |
-0.609 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.138 |
-1.932 |
-0.555 |
2.015 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.469 |
4.185 |
-3.193 |
| y |
4.185 |
-38.976 |
-0.388 |
| z |
-3.193 |
-0.388 |
-36.235 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.136 |
4.185 |
-3.193 |
| y |
4.185 |
-5.624 |
-0.388 |
| z |
-3.193 |
-0.388 |
-1.512 |
|
| Polar |
|---|
| 3z2-r2 | -3.025 |
|---|
| x2-y2 | 8.507 |
|---|
| xy | 4.185 |
|---|
| xz | -3.193 |
|---|
| yz | -0.388 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.524 |
0.030 |
-0.023 |
| y |
0.030 |
7.839 |
-0.186 |
| z |
-0.023 |
-0.186 |
7.483 |
<r2> (average value of r
2) Å
2
| <r2> |
174.460 |
| (<r2>)1/2 |
13.208 |