Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -551.464215 |
| Energy at 298.15K | |
| HF Energy | -551.464215 |
| Nuclear repulsion energy | 353.755281 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is D4h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
1.015 |
0.000 |
| C2 |
1.015 |
0.000 |
0.000 |
| C3 |
0.000 |
-1.015 |
0.000 |
| C4 |
-1.015 |
0.000 |
0.000 |
| F5 |
0.000 |
2.327 |
0.000 |
| F6 |
2.327 |
0.000 |
0.000 |
| F7 |
0.000 |
-2.327 |
0.000 |
| F8 |
-2.327 |
0.000 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.4353 | 2.0298 | 1.4353 | 1.3121 | 2.5386 | 3.3418 | 2.5386 |
C2 | 1.4353 | | 1.4353 | 2.0298 | 2.5386 | 1.3121 | 2.5386 | 3.3418 | C3 | 2.0298 | 1.4353 | | 1.4353 | 3.3418 | 2.5386 | 1.3121 | 2.5386 | C4 | 1.4353 | 2.0298 | 1.4353 | | 2.5386 | 3.3418 | 2.5386 | 1.3121 | F5 | 1.3121 | 2.5386 | 3.3418 | 2.5386 | | 3.2908 | 4.6539 | 3.2908 | F6 | 2.5386 | 1.3121 | 2.5386 | 3.3418 | 3.2908 | | 3.2908 | 4.6539 | F7 | 3.3418 | 2.5386 | 1.3121 | 2.5386 | 4.6539 | 3.2908 | | 3.2908 | F8 | 2.5386 | 3.3418 | 2.5386 | 1.3121 | 3.2908 | 4.6539 | 3.2908 | |
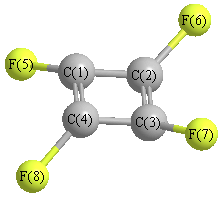 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
135.000 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
135.000 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
135.000 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
135.000 |
| C3 |
C2 |
F6 |
135.000 |
|
C3 |
C4 |
F8 |
135.000 |
| C4 |
C1 |
F5 |
135.000 |
|
C4 |
C3 |
F7 |
135.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.458 |
|
|
|
| 2 |
C |
0.407 |
|
|
|
| 3 |
C |
0.458 |
|
|
|
| 4 |
C |
0.407 |
|
|
|
| 5 |
F |
-0.468 |
|
|
|
| 6 |
F |
-0.397 |
|
|
|
| 7 |
F |
-0.468 |
|
|
|
| 8 |
F |
-0.397 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.412 |
0.000 |
0.000 |
| y |
0.000 |
-49.285 |
0.000 |
| z |
0.000 |
0.000 |
-41.053 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.756 |
0.000 |
0.000 |
| y |
0.000 |
-9.552 |
0.000 |
| z |
0.000 |
0.000 |
2.796 |
|
| Polar |
|---|
| 3z2-r2 | 5.591 |
|---|
| x2-y2 | 10.872 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.707 |
0.000 |
0.000 |
| y |
0.000 |
8.359 |
0.000 |
| z |
0.000 |
0.000 |
5.010 |
<r2> (average value of r
2) Å
2
| <r2> |
246.453 |
| (<r2>)1/2 |
15.699 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -551.543779 |
| Energy at 298.15K | -551.543846 |
| HF Energy | -551.543779 |
| Nuclear repulsion energy | 355.608399 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1861 |
1783 |
0.00 |
|
|
|
| 2 |
Ag |
1242 |
1190 |
0.00 |
|
|
|
| 3 |
Ag |
743 |
712 |
0.00 |
|
|
|
| 4 |
Ag |
357 |
342 |
0.00 |
|
|
|
| 5 |
Ag |
205 |
197 |
0.00 |
|
|
|
| 6 |
Au |
1323 |
1268 |
562.21 |
|
|
|
| 7 |
Au |
938 |
899 |
62.80 |
|
|
|
| 8 |
Au |
626 |
600 |
83.84 |
|
|
|
| 9 |
Au |
251 |
241 |
0.11 |
|
|
|
| 10 |
Au |
151 |
145 |
0.10 |
|
|
|
| 11 |
Bg |
1333 |
1277 |
0.00 |
|
|
|
| 12 |
Bg |
763 |
731 |
0.00 |
|
|
|
| 13 |
Bg |
491 |
471 |
0.00 |
|
|
|
| 14 |
Bg |
343 |
329 |
0.00 |
|
|
|
| 15 |
Bu |
1822 |
1746 |
276.80 |
|
|
|
| 16 |
Bu |
996 |
954 |
212.36 |
|
|
|
| 17 |
Bu |
314 |
301 |
4.75 |
|
|
|
| 18 |
Bu |
176 |
169 |
3.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6967.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 6677.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.135 |
0.758 |
0.667 |
| C2 |
-0.135 |
-0.758 |
0.667 |
| C3 |
-0.135 |
-0.758 |
-0.667 |
| C4 |
0.135 |
0.758 |
-0.667 |
| F5 |
-0.135 |
1.665 |
1.584 |
| F6 |
0.135 |
-1.665 |
1.584 |
| F7 |
0.135 |
-1.665 |
-1.584 |
| F8 |
-0.135 |
1.665 |
-1.584 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.5402 | 2.0381 | 1.3347 | 1.3170 | 2.5904 | 3.3073 | 2.4419 |
C2 | 1.5402 | | 1.3347 | 2.0381 | 2.5904 | 1.3170 | 2.4419 | 3.3073 | C3 | 2.0381 | 1.3347 | | 1.5402 | 3.3073 | 2.4419 | 1.3170 | 2.5904 | C4 | 1.3347 | 2.0381 | 1.5402 | | 2.4419 | 3.3073 | 2.5904 | 1.3170 | F5 | 1.3170 | 2.5904 | 3.3073 | 2.4419 | | 3.3402 | 4.6035 | 3.1677 | F6 | 2.5904 | 1.3170 | 2.4419 | 3.3073 | 3.3402 | | 3.1677 | 4.6035 | F7 | 3.3073 | 2.4419 | 1.3170 | 2.5904 | 4.6035 | 3.1677 | | 3.3402 | F8 | 2.4419 | 3.3073 | 2.5904 | 1.3170 | 3.1677 | 4.6035 | 3.3402 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
129.913 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
134.097 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
129.913 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
134.097 |
| C3 |
C2 |
F6 |
134.097 |
|
C3 |
C4 |
F8 |
129.913 |
| C4 |
C1 |
F5 |
134.097 |
|
C4 |
C3 |
F7 |
129.913 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.448 |
|
|
|
| 2 |
C |
0.448 |
|
|
|
| 3 |
C |
0.448 |
|
|
|
| 4 |
C |
0.448 |
|
|
|
| 5 |
F |
-0.448 |
|
|
|
| 6 |
F |
-0.448 |
|
|
|
| 7 |
F |
-0.448 |
|
|
|
| 8 |
F |
-0.448 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.178 |
0.064 |
0.000 |
| y |
0.064 |
-44.495 |
0.000 |
| z |
0.000 |
0.000 |
-43.769 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.954 |
0.064 |
0.000 |
| y |
0.064 |
-2.022 |
0.000 |
| z |
0.000 |
0.000 |
-0.932 |
|
| Polar |
|---|
| 3z2-r2 | -1.864 |
|---|
| x2-y2 | 3.317 |
|---|
| xy | 0.064 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.007 |
0.161 |
0.000 |
| y |
0.161 |
7.567 |
0.000 |
| z |
0.000 |
0.000 |
8.722 |
<r2> (average value of r
2) Å
2
| <r2> |
242.598 |
| (<r2>)1/2 |
15.576 |
