Jump to
S1C2
S1C3
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -224.032250 |
| Energy at 298.15K | -224.038611 |
| HF Energy | -224.032250 |
| Nuclear repulsion energy | 124.945967 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3920 |
3569 |
60.98 |
|
|
|
| 2 |
A |
3801 |
3461 |
6.75 |
|
|
|
| 3 |
A |
1937 |
1763 |
579.60 |
|
|
|
| 4 |
A |
1758 |
1601 |
1.23 |
|
|
|
| 5 |
A |
1280 |
1165 |
8.56 |
|
|
|
| 6 |
A |
1024 |
933 |
10.31 |
|
|
|
| 7 |
A |
594 |
541 |
59.58 |
|
|
|
| 8 |
A |
512 |
466 |
3.58 |
|
|
|
| 9 |
A |
393 |
358 |
88.95 |
|
|
|
| 10 |
B |
3920 |
3570 |
31.23 |
|
|
|
| 11 |
B |
3796 |
3457 |
69.48 |
|
|
|
| 12 |
B |
1767 |
1609 |
218.83 |
|
|
|
| 13 |
B |
1525 |
1388 |
225.96 |
|
|
|
| 14 |
B |
1145 |
1043 |
25.64 |
|
|
|
| 15 |
B |
867 |
790 |
89.61 |
|
|
|
| 16 |
B |
625 |
569 |
62.83 |
|
|
|
| 17 |
B |
572 |
521 |
208.39 |
|
|
|
| 18 |
B |
465 |
424 |
135.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14950.2 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 13613.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.142 |
| O2 |
0.000 |
0.000 |
1.341 |
| N3 |
-0.120 |
1.149 |
-0.602 |
| N4 |
0.120 |
-1.149 |
-0.602 |
| H5 |
0.000 |
1.983 |
-0.070 |
| H6 |
0.300 |
1.161 |
-1.506 |
| H7 |
0.000 |
-1.983 |
-0.070 |
| H8 |
-0.300 |
-1.161 |
-1.506 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.1995 | 1.3739 | 1.3739 | 1.9946 | 2.0382 | 1.9946 | 2.0382 |
O2 | 1.1995 | | 2.2609 | 2.2609 | 2.4343 | 3.0897 | 2.4343 | 3.0897 | N3 | 1.3739 | 2.2609 | | 2.3099 | 0.9971 | 0.9968 | 3.1791 | 2.4871 | N4 | 1.3739 | 2.2609 | 2.3099 | | 3.1791 | 2.4871 | 0.9971 | 0.9968 | H5 | 1.9946 | 2.4343 | 0.9971 | 3.1791 | | 1.6814 | 3.9666 | 3.4700 | H6 | 2.0382 | 3.0897 | 0.9968 | 2.4871 | 1.6814 | | 3.4700 | 2.3992 | H7 | 1.9946 | 2.4343 | 3.1791 | 0.9971 | 3.9666 | 3.4700 | | 1.6814 | H8 | 2.0382 | 3.0897 | 2.4871 | 0.9968 | 3.4700 | 2.3992 | 1.6814 | |
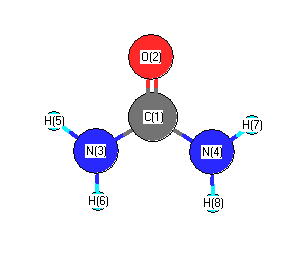 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
113.592 |
|
C1 |
N3 |
H6 |
117.695 |
| C1 |
N4 |
H7 |
113.592 |
|
C1 |
N4 |
H8 |
117.695 |
| O2 |
C1 |
N3 |
122.794 |
|
O2 |
C1 |
N4 |
122.794 |
| N3 |
C1 |
N4 |
114.412 |
|
H5 |
N3 |
H6 |
114.970 |
| H7 |
N4 |
H8 |
114.970 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.167 |
|
|
|
| 2 |
O |
-0.667 |
|
|
|
| 3 |
N |
-0.207 |
|
|
|
| 4 |
N |
-0.207 |
|
|
|
| 5 |
H |
0.003 |
|
|
|
| 6 |
H |
-0.046 |
|
|
|
| 7 |
H |
0.003 |
|
|
|
| 8 |
H |
-0.046 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.986 |
3.986 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.841 |
2.445 |
0.000 |
| y |
2.445 |
-17.365 |
0.000 |
| z |
0.000 |
0.000 |
-25.780 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.269 |
2.445 |
0.000 |
| y |
2.445 |
8.446 |
0.000 |
| z |
0.000 |
0.000 |
-4.177 |
|
| Polar |
|---|
| 3z2-r2 | -8.354 |
|---|
| x2-y2 | -8.477 |
|---|
| xy | 2.445 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.575 |
-0.086 |
0.000 |
| y |
-0.086 |
5.069 |
0.000 |
| z |
0.000 |
0.000 |
5.314 |
<r2> (average value of r
2) Å
2
| <r2> |
67.921 |
| (<r2>)1/2 |
8.241 |
Jump to
S1C1
S1C3
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -224.030817 |
| Energy at 298.15K | -224.036543 |
| HF Energy | -224.030817 |
| Nuclear repulsion energy | 125.025370 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3956 |
3602 |
70.21 |
|
|
|
| 2 |
A' |
3826 |
3484 |
10.35 |
|
|
|
| 3 |
A' |
1918 |
1747 |
617.30 |
|
|
|
| 4 |
A' |
1765 |
1608 |
7.59 |
|
|
|
| 5 |
A' |
1265 |
1152 |
8.02 |
|
|
|
| 6 |
A' |
1034 |
942 |
13.45 |
|
|
|
| 7 |
A' |
857 |
780 |
39.15 |
|
|
|
| 8 |
A' |
574 |
523 |
22.98 |
|
|
|
| 9 |
A' |
515 |
469 |
9.86 |
|
|
|
| 10 |
A' |
355 |
324 |
421.19 |
|
|
|
| 11 |
A" |
3953 |
3599 |
33.67 |
|
|
|
| 12 |
A" |
3818 |
3477 |
80.18 |
|
|
|
| 13 |
A" |
1760 |
1603 |
252.31 |
|
|
|
| 14 |
A" |
1527 |
1390 |
231.79 |
|
|
|
| 15 |
A" |
1107 |
1008 |
18.62 |
|
|
|
| 16 |
A" |
619 |
564 |
26.68 |
|
|
|
| 17 |
A" |
440 |
400 |
36.73 |
|
|
|
| 18 |
A" |
94 |
85 |
41.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14691.8 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 13378.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.010 |
0.139 |
0.000 |
| O2 |
0.031 |
1.340 |
0.000 |
| N3 |
0.031 |
-0.593 |
1.156 |
| N4 |
0.031 |
-0.593 |
-1.156 |
| H5 |
-0.139 |
-0.076 |
1.989 |
| H6 |
-0.228 |
-1.553 |
1.157 |
| H7 |
-0.139 |
-0.076 |
-1.989 |
| H8 |
-0.228 |
-1.553 |
-1.157 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2019 | 1.3678 | 1.3678 | 2.0063 | 2.0631 | 2.0063 | 2.0631 |
O2 | 1.2019 | | 2.2522 | 2.2522 | 2.4479 | 3.1266 | 2.4479 | 3.1266 | N3 | 1.3678 | 2.2522 | | 2.3113 | 0.9952 | 0.9943 | 3.1915 | 2.5174 | N4 | 1.3678 | 2.2522 | 2.3113 | | 3.1915 | 2.5174 | 0.9952 | 0.9943 | H5 | 2.0063 | 2.4479 | 0.9952 | 3.1915 | | 1.6973 | 3.9784 | 3.4767 | H6 | 2.0631 | 3.1266 | 0.9943 | 2.5174 | 1.6973 | | 3.4767 | 2.3141 | H7 | 2.0063 | 2.4479 | 3.1915 | 0.9952 | 3.9784 | 3.4767 | | 1.6973 | H8 | 2.0631 | 3.1266 | 2.5174 | 0.9943 | 3.4767 | 2.3141 | 1.6973 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
115.315 |
|
C1 |
N3 |
H6 |
120.908 |
| C1 |
N4 |
H7 |
115.315 |
|
C1 |
N4 |
H8 |
120.908 |
| O2 |
C1 |
N3 |
122.304 |
|
O2 |
C1 |
N4 |
122.304 |
| N3 |
C1 |
N4 |
115.320 |
|
H5 |
N3 |
H6 |
117.108 |
| H7 |
N4 |
H8 |
117.108 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.262 |
|
|
|
| 2 |
O |
-0.684 |
|
|
|
| 3 |
N |
-0.219 |
|
|
|
| 4 |
N |
-0.219 |
|
|
|
| 5 |
H |
-0.012 |
|
|
|
| 6 |
H |
-0.058 |
|
|
|
| 7 |
H |
-0.012 |
|
|
|
| 8 |
H |
-0.058 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.418 |
-4.348 |
0.000 |
4.573 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.902 |
1.473 |
0.000 |
| y |
1.473 |
-25.168 |
0.000 |
| z |
0.000 |
0.000 |
-17.316 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.661 |
1.473 |
0.000 |
| y |
1.473 |
-3.559 |
0.000 |
| z |
0.000 |
0.000 |
8.220 |
|
| Polar |
|---|
| 3z2-r2 | 16.439 |
|---|
| x2-y2 | -0.734 |
|---|
| xy | 1.473 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.563 |
0.039 |
0.000 |
| y |
0.039 |
5.304 |
0.000 |
| z |
0.000 |
0.000 |
5.095 |
<r2> (average value of r
2) Å
2
| <r2> |
67.931 |
| (<r2>)1/2 |
8.242 |
Jump to
S1C1
S1C2
Energy calculated at HF/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -224.030450 |
| Energy at 298.15K | |
| HF Energy | -224.030450 |
| Nuclear repulsion energy | 125.142225 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3988 |
3631 |
86.23 |
|
|
|
| 2 |
A1 |
3848 |
3504 |
10.48 |
|
|
|
| 3 |
A1 |
1905 |
1735 |
653.07 |
|
|
|
| 4 |
A1 |
1758 |
1601 |
0.50 |
|
|
|
| 5 |
A1 |
1242 |
1131 |
4.30 |
|
|
|
| 6 |
A1 |
1042 |
949 |
12.70 |
|
|
|
| 7 |
A1 |
515 |
469 |
3.45 |
|
|
|
| 8 |
A2 |
432 |
394 |
0.00 |
|
|
|
| 9 |
A2 |
374i |
341i |
0.00 |
|
|
|
| 10 |
B1 |
857 |
781 |
19.03 |
|
|
|
| 11 |
B1 |
602 |
548 |
8.60 |
|
|
|
| 12 |
B1 |
258i |
235i |
494.59 |
|
|
|
| 13 |
B2 |
3985 |
3629 |
38.18 |
|
|
|
| 14 |
B2 |
3840 |
3496 |
101.02 |
|
|
|
| 15 |
B2 |
1760 |
1603 |
296.21 |
|
|
|
| 16 |
B2 |
1530 |
1393 |
250.71 |
|
|
|
| 17 |
B2 |
1085 |
988 |
13.79 |
|
|
|
| 18 |
B2 |
613 |
558 |
19.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14184.4 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 12916.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.139 |
| O2 |
0.000 |
0.000 |
1.343 |
| N3 |
0.000 |
1.151 |
-0.589 |
| N4 |
0.000 |
-1.151 |
-0.589 |
| H5 |
0.000 |
2.008 |
-0.087 |
| H6 |
0.000 |
1.168 |
-1.581 |
| H7 |
0.000 |
-2.008 |
-0.087 |
| H8 |
0.000 |
-1.168 |
-1.581 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2038 | 1.3618 | 1.3618 | 2.0206 | 2.0787 | 2.0206 | 2.0787 |
O2 | 1.2038 | | 2.2485 | 2.2485 | 2.4648 | 3.1481 | 2.4648 | 3.1481 | N3 | 1.3618 | 2.2485 | | 2.3021 | 0.9932 | 0.9922 | 3.1987 | 2.5220 | N4 | 1.3618 | 2.2485 | 2.3021 | | 3.1987 | 2.5220 | 0.9932 | 0.9922 | H5 | 2.0206 | 2.4648 | 0.9932 | 3.1987 | | 1.7143 | 4.0160 | 3.5096 | H6 | 2.0787 | 3.1481 | 0.9922 | 2.5220 | 1.7143 | | 3.5096 | 2.3354 | H7 | 2.0206 | 2.4648 | 3.1987 | 0.9932 | 4.0160 | 3.5096 | | 1.7143 | H8 | 2.0787 | 3.1481 | 2.5220 | 0.9922 | 3.5096 | 2.3354 | 1.7143 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.328 |
|
C1 |
N3 |
H6 |
123.264 |
| C1 |
N4 |
H7 |
117.328 |
|
C1 |
N4 |
H8 |
123.264 |
| O2 |
C1 |
N3 |
122.303 |
|
O2 |
C1 |
N4 |
122.303 |
| N3 |
C1 |
N4 |
115.394 |
|
H5 |
N3 |
H6 |
119.408 |
| H7 |
N4 |
H8 |
119.408 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.268 |
|
|
|
| 2 |
O |
-0.693 |
|
|
|
| 3 |
N |
-0.180 |
|
|
|
| 4 |
N |
-0.180 |
|
|
|
| 5 |
H |
-0.032 |
|
|
|
| 6 |
H |
-0.076 |
|
|
|
| 7 |
H |
-0.032 |
|
|
|
| 8 |
H |
-0.076 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.612 |
4.612 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.244 |
0.000 |
0.000 |
| y |
0.000 |
-16.719 |
0.000 |
| z |
0.000 |
0.000 |
-24.817 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.476 |
0.000 |
0.000 |
| y |
0.000 |
8.811 |
0.000 |
| z |
0.000 |
0.000 |
-3.336 |
|
| Polar |
|---|
| 3z2-r2 | -6.671 |
|---|
| x2-y2 | -9.525 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.562 |
0.000 |
0.000 |
| y |
0.000 |
5.098 |
0.000 |
| z |
0.000 |
0.000 |
5.294 |
<r2> (average value of r
2) Å
2
| <r2> |
67.858 |
| (<r2>)1/2 |
8.238 |
