Jump to
S1C2
S1C3
Energy calculated at QCISD(T)/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -224.763087 |
| Energy at 298.15K | -224.769245 |
| HF Energy | -224.027918 |
| Nuclear repulsion energy | 122.514988 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3662 |
3580 |
|
|
|
|
| 2 |
A |
3543 |
3465 |
|
|
|
|
| 3 |
A |
1773 |
1734 |
|
|
|
|
| 4 |
A |
1618 |
1582 |
|
|
|
|
| 5 |
A |
1180 |
1154 |
|
|
|
|
| 6 |
A |
937 |
916 |
|
|
|
|
| 7 |
A |
611 |
598 |
|
|
|
|
| 8 |
A |
466 |
456 |
|
|
|
|
| 9 |
A |
373 |
365 |
|
|
|
|
| 10 |
B |
3662 |
3580 |
|
|
|
|
| 11 |
B |
3541 |
3462 |
|
|
|
|
| 12 |
B |
1626 |
1590 |
|
|
|
|
| 13 |
B |
1402 |
1371 |
|
|
|
|
| 14 |
B |
1059 |
1035 |
|
|
|
|
| 15 |
B |
769 |
752 |
|
|
|
|
| 16 |
B |
575 |
562 |
|
|
|
|
| 17 |
B |
542 |
530 |
|
|
|
|
| 18 |
B |
432 |
422 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13884.7 cm
-1
Scaled (by 0.9778) Zero Point Vibrational Energy (zpe) 13576.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.146 |
| O2 |
0.000 |
0.000 |
1.375 |
| N3 |
0.000 |
1.171 |
-0.623 |
| N4 |
0.000 |
-1.171 |
-0.623 |
| H5 |
0.241 |
1.989 |
-0.069 |
| H6 |
0.502 |
1.118 |
-1.506 |
| H7 |
-0.241 |
-1.989 |
-0.069 |
| H8 |
-0.502 |
-1.118 |
-1.506 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2293 | 1.4012 | 1.4012 | 2.0150 | 2.0573 | 2.0150 | 2.0573 |
O2 | 1.2293 | | 2.3164 | 2.3164 | 2.4698 | 3.1313 | 2.4698 | 3.1313 | N3 | 1.4012 | 2.3164 | | 2.3425 | 1.0167 | 1.0171 | 3.2174 | 2.5048 | N4 | 1.4012 | 2.3164 | 2.3425 | | 3.2174 | 2.5048 | 1.0167 | 1.0171 | H5 | 2.0150 | 2.4698 | 1.0167 | 3.2174 | | 1.7003 | 4.0069 | 3.5032 | H6 | 2.0573 | 3.1313 | 1.0171 | 2.5048 | 1.7003 | | 3.5032 | 2.4521 | H7 | 2.0150 | 2.4698 | 3.2174 | 1.0167 | 4.0069 | 3.5032 | | 1.7003 | H8 | 2.0573 | 3.1313 | 2.5048 | 1.0171 | 3.5032 | 2.4521 | 1.7003 | |
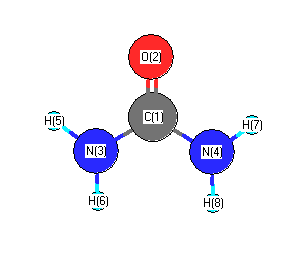 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
111.904 |
|
C1 |
N3 |
H6 |
115.663 |
| C1 |
N4 |
H7 |
111.904 |
|
C1 |
N4 |
H8 |
115.663 |
| O2 |
C1 |
N3 |
123.295 |
|
O2 |
C1 |
N4 |
123.295 |
| N3 |
C1 |
N4 |
113.410 |
|
H5 |
N3 |
H6 |
113.445 |
| H7 |
N4 |
H8 |
113.445 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at QCISD(T)/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -224.761157 |
| Energy at 298.15K | -224.766942 |
| HF Energy | -224.026435 |
| Nuclear repulsion energy | 122.566719 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3691 |
3609 |
|
|
|
|
| 2 |
A' |
3566 |
3487 |
|
|
|
|
| 3 |
A' |
1762 |
1723 |
|
|
|
|
| 4 |
A' |
1631 |
1595 |
|
|
|
|
| 5 |
A' |
1174 |
1148 |
|
|
|
|
| 6 |
A' |
945 |
924 |
|
|
|
|
| 7 |
A' |
754 |
737 |
|
|
|
|
| 8 |
A' |
525 |
514 |
|
|
|
|
| 9 |
A' |
482 |
471 |
|
|
|
|
| 10 |
A' |
435 |
425 |
|
|
|
|
| 11 |
A" |
3688 |
3606 |
|
|
|
|
| 12 |
A" |
3559 |
3480 |
|
|
|
|
| 13 |
A" |
1619 |
1583 |
|
|
|
|
| 14 |
A" |
1407 |
1376 |
|
|
|
|
| 15 |
A" |
1024 |
1001 |
|
|
|
|
| 16 |
A" |
561 |
549 |
|
|
|
|
| 17 |
A" |
431 |
422 |
|
|
|
|
| 18 |
A" |
192 |
188 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13722.0 cm
-1
Scaled (by 0.9778) Zero Point Vibrational Energy (zpe) 13417.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.008 |
0.141 |
0.000 |
| O2 |
0.045 |
1.372 |
0.000 |
| N3 |
0.045 |
-0.612 |
1.175 |
| N4 |
0.045 |
-0.612 |
-1.175 |
| H5 |
-0.189 |
-0.072 |
2.002 |
| H6 |
-0.326 |
-1.557 |
1.160 |
| H7 |
-0.189 |
-0.072 |
-2.002 |
| H8 |
-0.326 |
-1.557 |
-1.160 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2310 | 1.3962 | 1.3962 | 2.0229 | 2.0837 | 2.0229 | 2.0837 |
O2 | 1.2310 | | 2.3056 | 2.3056 | 2.4793 | 3.1718 | 2.4793 | 3.1718 | N3 | 1.3962 | 2.3056 | | 2.3499 | 1.0149 | 1.0149 | 3.2308 | 2.5463 | N4 | 1.3962 | 2.3056 | 2.3499 | | 3.2308 | 2.5463 | 1.0149 | 1.0149 | H5 | 2.0229 | 2.4793 | 1.0149 | 3.2308 | | 1.7121 | 4.0038 | 3.4962 | H6 | 2.0837 | 3.1718 | 1.0149 | 2.5463 | 1.7121 | | 3.4962 | 2.3207 | H7 | 2.0229 | 2.4793 | 3.2308 | 1.0149 | 4.0038 | 3.4962 | | 1.7121 | H8 | 2.0837 | 3.1718 | 2.5463 | 1.0149 | 3.4962 | 2.3207 | 1.7121 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
113.119 |
|
C1 |
N3 |
H6 |
118.731 |
| C1 |
N4 |
H7 |
113.119 |
|
C1 |
N4 |
H8 |
118.731 |
| O2 |
C1 |
N3 |
122.585 |
|
O2 |
C1 |
N4 |
122.585 |
| N3 |
C1 |
N4 |
114.606 |
|
H5 |
N3 |
H6 |
115.020 |
| H7 |
N4 |
H8 |
115.020 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at QCISD(T)/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -224.759816 |
| Energy at 298.15K | |
| HF Energy | -224.026575 |
| Nuclear repulsion energy | 122.820066 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3759 |
3676 |
|
|
|
|
| 2 |
A1 |
3617 |
3537 |
|
|
|
|
| 3 |
A1 |
1752 |
1713 |
|
|
|
|
| 4 |
A1 |
1622 |
1586 |
|
|
|
|
| 5 |
A1 |
1133 |
1108 |
|
|
|
|
| 6 |
A1 |
962 |
941 |
|
|
|
|
| 7 |
A1 |
469 |
459 |
|
|
|
|
| 8 |
A2 |
361 |
353 |
|
|
|
|
| 9 |
A2 |
421i |
412i |
|
|
|
|
| 10 |
B1 |
755 |
738 |
|
|
|
|
| 11 |
B1 |
561 |
549 |
|
|
|
|
| 12 |
B1 |
348i |
340i |
|
|
|
|
| 13 |
B2 |
3757 |
3674 |
|
|
|
|
| 14 |
B2 |
3610 |
3530 |
|
|
|
|
| 15 |
B2 |
1624 |
1588 |
|
|
|
|
| 16 |
B2 |
1417 |
1386 |
|
|
|
|
| 17 |
B2 |
986 |
964 |
|
|
|
|
| 18 |
B2 |
552 |
539 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13083.3 cm
-1
Scaled (by 0.9778) Zero Point Vibrational Energy (zpe) 12792.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.142 |
| O2 |
0.000 |
0.000 |
1.376 |
| N3 |
0.000 |
1.167 |
-0.603 |
| N4 |
0.000 |
-1.167 |
-0.603 |
| H5 |
0.000 |
2.038 |
-0.092 |
| H6 |
0.000 |
1.182 |
-1.613 |
| H7 |
0.000 |
-2.038 |
-0.092 |
| H8 |
0.000 |
-1.182 |
-1.613 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2336 | 1.3843 | 1.3843 | 2.0519 | 2.1163 | 2.0519 | 2.1163 |
O2 | 1.2336 | | 2.2971 | 2.2971 | 2.5120 | 3.2141 | 2.5120 | 3.2141 | N3 | 1.3843 | 2.2971 | | 2.3333 | 1.0104 | 1.0101 | 3.2455 | 2.5570 | N4 | 1.3843 | 2.2971 | 2.3333 | | 3.2455 | 2.5570 | 1.0104 | 1.0101 | H5 | 2.0519 | 2.5120 | 1.0104 | 3.2455 | | 1.7451 | 4.0768 | 3.5618 | H6 | 2.1163 | 3.2141 | 1.0101 | 2.5570 | 1.7451 | | 3.5618 | 2.3648 | H7 | 2.0519 | 2.5120 | 3.2455 | 1.0104 | 4.0768 | 3.5618 | | 1.7451 | H8 | 2.1163 | 3.2141 | 2.5570 | 1.0101 | 3.5618 | 2.3648 | 1.7451 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.068 |
|
C1 |
N3 |
H6 |
123.461 |
| C1 |
N4 |
H7 |
117.068 |
|
C1 |
N4 |
H8 |
123.461 |
| O2 |
C1 |
N3 |
122.568 |
|
O2 |
C1 |
N4 |
122.568 |
| N3 |
C1 |
N4 |
114.865 |
|
H5 |
N3 |
H6 |
119.472 |
| H7 |
N4 |
H8 |
119.472 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
