Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -225.108771 |
| Energy at 298.15K | |
| HF Energy | -224.878628 |
| Nuclear repulsion energy | 123.389246 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3776 |
3634 |
58.02 |
|
|
|
| 2 |
A1 |
3634 |
3497 |
7.00 |
|
|
|
| 3 |
A1 |
1758 |
1692 |
489.72 |
|
|
|
| 4 |
A1 |
1625 |
1564 |
1.11 |
|
|
|
| 5 |
A1 |
1139 |
1096 |
1.65 |
|
|
|
| 6 |
A1 |
970 |
934 |
10.92 |
|
|
|
| 7 |
A1 |
475 |
457 |
2.90 |
|
|
|
| 8 |
A2 |
377 |
363 |
0.00 |
|
|
|
| 9 |
A2 |
353i |
340i |
0.00 |
|
|
|
| 10 |
B1 |
765 |
736 |
3.10 |
|
|
|
| 11 |
B1 |
569 |
547 |
9.55 |
|
|
|
| 12 |
B1 |
251i |
242i |
425.51 |
|
|
|
| 13 |
B2 |
3774 |
3631 |
44.58 |
|
|
|
| 14 |
B2 |
3626 |
3490 |
73.75 |
|
|
|
| 15 |
B2 |
1626 |
1565 |
263.72 |
|
|
|
| 16 |
B2 |
1422 |
1369 |
217.50 |
|
|
|
| 17 |
B2 |
989 |
951 |
16.06 |
|
|
|
| 18 |
B2 |
559 |
538 |
13.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13239.3 cm
-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 12740.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.139 |
| O2 |
0.000 |
0.000 |
1.368 |
| N3 |
0.000 |
1.162 |
-0.599 |
| N4 |
0.000 |
-1.162 |
-0.599 |
| H5 |
0.000 |
2.030 |
-0.089 |
| H6 |
0.000 |
1.183 |
-1.605 |
| H7 |
0.000 |
-2.030 |
-0.089 |
| H8 |
0.000 |
-1.183 |
-1.605 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2287 | 1.3769 | 1.3769 | 2.0423 | 2.1080 | 2.0423 | 2.1080 |
O2 | 1.2287 | | 2.2848 | 2.2848 | 2.4982 | 3.2000 | 2.4982 | 3.2000 | N3 | 1.3769 | 2.2848 | | 2.3242 | 1.0065 | 1.0062 | 3.2322 | 2.5521 | N4 | 1.3769 | 2.2848 | 2.3242 | | 3.2322 | 2.5521 | 1.0065 | 1.0062 | H5 | 2.0423 | 2.4982 | 1.0065 | 3.2322 | | 1.7365 | 4.0590 | 3.5528 | H6 | 2.1080 | 3.2000 | 1.0062 | 2.5521 | 1.7365 | | 3.5528 | 2.3667 | H7 | 2.0423 | 2.4982 | 3.2322 | 1.0065 | 4.0590 | 3.5528 | | 1.7365 | H8 | 2.1080 | 3.2000 | 2.5521 | 1.0062 | 3.5528 | 2.3667 | 1.7365 | |
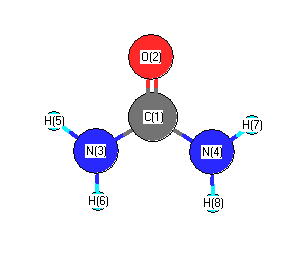 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.090 |
|
C1 |
N3 |
H6 |
123.648 |
| C1 |
N4 |
H7 |
117.090 |
|
C1 |
N4 |
H8 |
123.648 |
| O2 |
C1 |
N3 |
122.436 |
|
O2 |
C1 |
N4 |
122.436 |
| N3 |
C1 |
N4 |
115.128 |
|
H5 |
N3 |
H6 |
119.262 |
| H7 |
N4 |
H8 |
119.262 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
