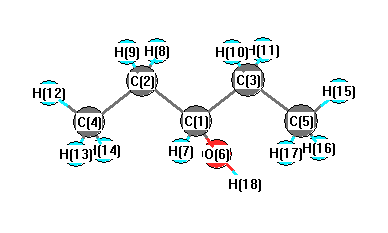
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3886 |
3724 |
22.88 |
|
|
|
| 2 |
A |
3156 |
3024 |
24.73 |
|
|
|
| 3 |
A |
3138 |
3007 |
30.50 |
|
|
|
| 4 |
A |
3132 |
3002 |
50.05 |
|
|
|
| 5 |
A |
3125 |
2995 |
52.45 |
|
|
|
| 6 |
A |
3104 |
2974 |
14.04 |
|
|
|
| 7 |
A |
3096 |
2967 |
0.74 |
|
|
|
| 8 |
A |
3059 |
2932 |
48.80 |
|
|
|
| 9 |
A |
3056 |
2929 |
33.65 |
|
|
|
| 10 |
A |
3052 |
2924 |
4.86 |
|
|
|
| 11 |
A |
3047 |
2920 |
28.44 |
|
|
|
| 12 |
A |
2979 |
2854 |
46.21 |
|
|
|
| 13 |
A |
1489 |
1427 |
7.95 |
|
|
|
| 14 |
A |
1486 |
1424 |
6.85 |
|
|
|
| 15 |
A |
1477 |
1416 |
7.97 |
|
|
|
| 16 |
A |
1474 |
1413 |
7.05 |
|
|
|
| 17 |
A |
1464 |
1403 |
3.32 |
|
|
|
| 18 |
A |
1454 |
1393 |
0.65 |
|
|
|
| 19 |
A |
1435 |
1375 |
9.35 |
|
|
|
| 20 |
A |
1400 |
1341 |
1.90 |
|
|
|
| 21 |
A |
1390 |
1332 |
5.28 |
|
|
|
| 22 |
A |
1386 |
1328 |
11.23 |
|
|
|
| 23 |
A |
1355 |
1299 |
0.27 |
|
|
|
| 24 |
A |
1319 |
1264 |
15.09 |
|
|
|
| 25 |
A |
1299 |
1245 |
1.12 |
|
|
|
| 26 |
A |
1274 |
1221 |
10.81 |
|
|
|
| 27 |
A |
1250 |
1198 |
25.32 |
|
|
|
| 28 |
A |
1179 |
1129 |
26.92 |
|
|
|
| 29 |
A |
1155 |
1107 |
4.49 |
|
|
|
| 30 |
A |
1101 |
1055 |
3.83 |
|
|
|
| 31 |
A |
1086 |
1041 |
21.16 |
|
|
|
| 32 |
A |
1039 |
996 |
2.40 |
|
|
|
| 33 |
A |
1028 |
985 |
2.23 |
|
|
|
| 34 |
A |
983 |
942 |
50.42 |
|
|
|
| 35 |
A |
943 |
903 |
8.64 |
|
|
|
| 36 |
A |
863 |
827 |
3.81 |
|
|
|
| 37 |
A |
778 |
745 |
1.58 |
|
|
|
| 38 |
A |
763 |
731 |
0.25 |
|
|
|
| 39 |
A |
495 |
474 |
3.27 |
|
|
|
| 40 |
A |
474 |
454 |
6.33 |
|
|
|
| 41 |
A |
397 |
380 |
1.04 |
|
|
|
| 42 |
A |
306 |
293 |
8.00 |
|
|
|
| 43 |
A |
267 |
256 |
83.16 |
|
|
|
| 44 |
A |
237 |
227 |
21.66 |
|
|
|
| 45 |
A |
210 |
201 |
0.73 |
|
|
|
| 46 |
A |
188 |
181 |
0.13 |
|
|
|
| 47 |
A |
103 |
99 |
1.62 |
|
|
|
| 48 |
A |
93 |
89 |
0.50 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36231.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 34720.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.726 |
|
|
|
| 2 |
C |
0.621 |
|
|
|
| 3 |
C |
0.510 |
|
|
|
| 4 |
C |
0.404 |
|
|
|
| 5 |
C |
0.452 |
|
|
|
| 6 |
O |
-0.588 |
|
|
|
| 7 |
H |
-0.450 |
|
|
|
| 8 |
H |
-0.266 |
|
|
|
| 9 |
H |
-0.270 |
|
|
|
| 10 |
H |
-0.272 |
|
|
|
| 11 |
H |
-0.255 |
|
|
|
| 12 |
H |
-0.111 |
|
|
|
| 13 |
H |
-0.124 |
|
|
|
| 14 |
H |
-0.093 |
|
|
|
| 15 |
H |
-0.138 |
|
|
|
| 16 |
H |
-0.142 |
|
|
|
| 17 |
H |
-0.114 |
|
|
|
| 18 |
H |
0.109 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.177 |
-0.701 |
0.822 |
1.598 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.449 |
2.165 |
0.194 |
| y |
2.165 |
-39.826 |
1.217 |
| z |
0.194 |
1.217 |
-40.446 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.687 |
2.165 |
0.194 |
| y |
2.165 |
0.122 |
1.217 |
| z |
0.194 |
1.217 |
-0.809 |
|
| Polar |
|---|
| 3z2-r2 | -1.618 |
|---|
| x2-y2 | 0.377 |
|---|
| xy | 2.165 |
|---|
| xz | 0.194 |
|---|
| yz | 1.217 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.028 |
0.052 |
0.015 |
| y |
0.052 |
9.763 |
0.131 |
| z |
0.015 |
0.131 |
9.243 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |