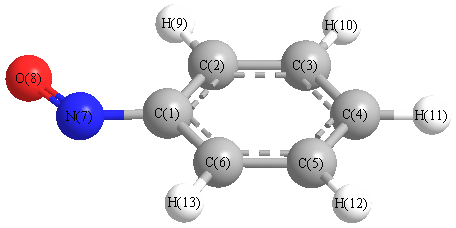
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3238 |
3103 |
7.24 |
|
|
|
| 2 |
A' |
3234 |
3099 |
6.97 |
|
|
|
| 3 |
A' |
3224 |
3090 |
5.67 |
|
|
|
| 4 |
A' |
3217 |
3083 |
4.43 |
|
|
|
| 5 |
A' |
3207 |
3073 |
0.38 |
|
|
|
| 6 |
A' |
1687 |
1617 |
47.03 |
|
|
|
| 7 |
A' |
1664 |
1595 |
2.56 |
|
|
|
| 8 |
A' |
1623 |
1555 |
186.28 |
|
|
|
| 9 |
A' |
1499 |
1436 |
7.73 |
|
|
|
| 10 |
A' |
1486 |
1424 |
25.42 |
|
|
|
| 11 |
A' |
1401 |
1343 |
16.07 |
|
|
|
| 12 |
A' |
1322 |
1267 |
6.51 |
|
|
|
| 13 |
A' |
1203 |
1153 |
48.39 |
|
|
|
| 14 |
A' |
1178 |
1129 |
1.58 |
|
|
|
| 15 |
A' |
1144 |
1096 |
119.71 |
|
|
|
| 16 |
A' |
1097 |
1051 |
5.52 |
|
|
|
| 17 |
A' |
1044 |
1000 |
6.50 |
|
|
|
| 18 |
A' |
1014 |
972 |
0.41 |
|
|
|
| 19 |
A' |
843 |
808 |
33.59 |
|
|
|
| 20 |
A' |
683 |
654 |
9.60 |
|
|
|
| 21 |
A' |
618 |
593 |
0.09 |
|
|
|
| 22 |
A' |
449 |
430 |
0.87 |
|
|
|
| 23 |
A' |
256 |
245 |
1.97 |
|
|
|
| 24 |
A" |
1011 |
969 |
0.02 |
|
|
|
| 25 |
A" |
999 |
958 |
0.00 |
|
|
|
| 26 |
A" |
959 |
919 |
4.35 |
|
|
|
| 27 |
A" |
861 |
825 |
0.01 |
|
|
|
| 28 |
A" |
776 |
743 |
58.52 |
|
|
|
| 29 |
A" |
693 |
664 |
27.58 |
|
|
|
| 30 |
A" |
471 |
451 |
2.66 |
|
|
|
| 31 |
A" |
418 |
400 |
0.00 |
|
|
|
| 32 |
A" |
248 |
237 |
0.27 |
|
|
|
| 33 |
A" |
111 |
107 |
0.11 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21438.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 20544.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.614 |
|
|
|
| 2 |
C |
0.752 |
|
|
|
| 3 |
C |
0.583 |
|
|
|
| 4 |
C |
0.471 |
|
|
|
| 5 |
C |
0.634 |
|
|
|
| 6 |
C |
0.503 |
|
|
|
| 7 |
N |
-0.158 |
|
|
|
| 8 |
O |
-0.552 |
|
|
|
| 9 |
H |
-0.596 |
|
|
|
| 10 |
H |
-0.564 |
|
|
|
| 11 |
H |
-0.559 |
|
|
|
| 12 |
H |
-0.546 |
|
|
|
| 13 |
H |
-0.581 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.143 |
-3.650 |
0.000 |
3.825 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.656 |
2.397 |
0.000 |
| y |
2.397 |
-48.829 |
0.000 |
| z |
0.000 |
0.000 |
-47.991 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.754 |
2.397 |
0.000 |
| y |
2.397 |
-4.006 |
0.000 |
| z |
0.000 |
0.000 |
-2.748 |
|
| Polar |
|---|
| 3z2-r2 | -5.497 |
|---|
| x2-y2 | 7.173 |
|---|
| xy | 2.397 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.717 |
-1.722 |
0.000 |
| y |
-1.722 |
16.734 |
0.000 |
| z |
0.000 |
0.000 |
7.063 |
<r2> (average value of r
2) Å
2
| <r2> |
249.148 |
| (<r2>)1/2 |
15.784 |