All results from a given calculation for C6H10 (1,5-Hexadiene)
using model chemistry: MP4/aug-cc-pVDZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2 |
1Ag |
Energy calculated at MP4/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -233.969199 |
| Energy at 298.15K | -233.978828 |
| HF Energy | -233.009037 |
| Nuclear repulsion energy | 210.411593 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Geometric Data calculated at MP4/aug-cc-pVDZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.429 |
0.646 |
-0.317 |
| C2 |
0.429 |
-0.646 |
-0.317 |
| C3 |
0.429 |
1.892 |
-0.354 |
| C4 |
-0.429 |
-1.892 |
-0.354 |
| C5 |
0.476 |
2.821 |
0.632 |
| C6 |
-0.476 |
-2.821 |
0.632 |
| H7 |
-1.099 |
0.628 |
-1.201 |
| H8 |
-1.073 |
0.658 |
0.583 |
| H9 |
1.099 |
-0.628 |
-1.201 |
| H10 |
1.073 |
-0.658 |
0.583 |
| H11 |
1.066 |
2.024 |
-1.242 |
| H12 |
-1.066 |
-2.024 |
-1.242 |
| H13 |
1.129 |
3.699 |
0.560 |
| H14 |
-0.143 |
2.724 |
1.534 |
| H15 |
-1.129 |
-3.699 |
0.560 |
| H16 |
0.143 |
-2.724 |
1.534 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.5509 | 1.5131 | 2.5382 | 2.5400 | 3.5950 | 1.1093 | 1.1071 | 2.1770 | 2.1833 | 2.2337 | 2.8965 | 3.5378 | 2.7973 | 4.4875 | 3.8869 |
C2 | 1.5509 | | 2.5382 | 1.5131 | 3.5950 | 2.5400 | 2.1770 | 2.1833 | 1.1093 | 1.1071 | 2.8965 | 2.2337 | 4.4875 | 3.8869 | 3.5378 | 2.7973 | C3 | 1.5131 | 2.5382 | | 3.8798 | 1.3554 | 4.8993 | 2.1566 | 2.1579 | 2.7417 | 2.7917 | 1.1015 | 4.2846 | 2.1424 | 2.1401 | 5.8752 | 4.9949 | C4 | 2.5382 | 1.5131 | 3.8798 | | 4.8993 | 1.3554 | 2.7417 | 2.7917 | 2.1566 | 2.1579 | 4.2846 | 1.1015 | 5.8752 | 4.9949 | 2.1424 | 2.1401 | C5 | 2.5400 | 3.5950 | 1.3554 | 4.8993 | | 5.7221 | 3.2637 | 2.6615 | 3.9552 | 3.5301 | 2.1205 | 5.4190 | 1.0959 | 1.0980 | 6.7151 | 5.6278 | C6 | 3.5950 | 2.5400 | 4.8993 | 1.3554 | 5.7221 | | 3.9552 | 3.5301 | 3.2637 | 2.6615 | 5.4190 | 2.1205 | 6.7151 | 5.6278 | 1.0959 | 1.0980 | H7 | 1.1093 | 2.1770 | 2.1566 | 2.7417 | 3.2637 | 3.9552 | | 1.7843 | 2.5321 | 3.0912 | 2.5767 | 2.6523 | 4.1828 | 3.5756 | 4.6716 | 4.5005 | H8 | 1.1071 | 2.1833 | 2.1579 | 2.7917 | 2.6615 | 3.5301 | 1.7843 | | 3.0912 | 2.5176 | 3.1264 | 3.2439 | 3.7546 | 2.4573 | 4.3572 | 3.7173 | H9 | 2.1770 | 1.1093 | 2.7417 | 2.1566 | 3.9552 | 3.2637 | 2.5321 | 3.0912 | | 1.7843 | 2.6523 | 2.5767 | 4.6716 | 4.5005 | 4.1828 | 3.5756 | H10 | 2.1833 | 1.1071 | 2.7917 | 2.1579 | 3.5301 | 2.6615 | 3.0912 | 2.5176 | 1.7843 | | 3.2439 | 3.1264 | 4.3572 | 3.7173 | 3.7546 | 2.4573 | H11 | 2.2337 | 2.8965 | 1.1015 | 4.2846 | 2.1205 | 5.4190 | 2.5767 | 3.1264 | 2.6523 | 3.2439 | | 4.5748 | 2.4616 | 3.1075 | 6.3887 | 5.5766 | H12 | 2.8965 | 2.2337 | 4.2846 | 1.1015 | 5.4190 | 2.1205 | 2.6523 | 3.2439 | 2.5767 | 3.1264 | 4.5748 | | 6.3887 | 5.5766 | 2.4616 | 3.1075 | H13 | 3.5378 | 4.4875 | 2.1424 | 5.8752 | 1.0959 | 6.7151 | 4.1828 | 3.7546 | 4.6716 | 4.3572 | 2.4616 | 6.3887 | | 1.8745 | 7.7345 | 6.5706 | H14 | 2.7973 | 3.8869 | 2.1401 | 4.9949 | 1.0980 | 5.6278 | 3.5756 | 2.4573 | 4.5005 | 3.7173 | 3.1075 | 5.5766 | 1.8745 | | 6.5706 | 5.4553 | H15 | 4.4875 | 3.5378 | 5.8752 | 2.1424 | 6.7151 | 1.0959 | 4.6716 | 4.3572 | 4.1828 | 3.7546 | 6.3887 | 2.4616 | 7.7345 | 6.5706 | | 1.8745 | H16 | 3.8869 | 2.7973 | 4.9949 | 2.1401 | 5.6278 | 1.0980 | 4.5005 | 3.7173 | 3.5756 | 2.4573 | 5.5766 | 3.1075 | 6.5706 | 5.4553 | 1.8745 | |
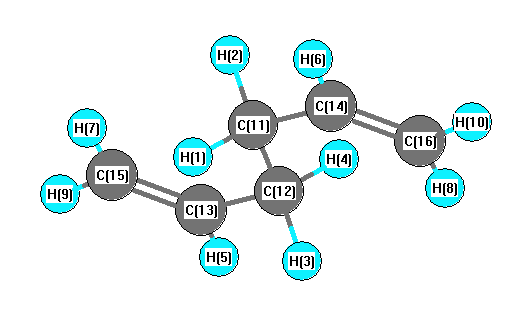 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
111.863 |
|
C1 |
C2 |
H9 |
108.708 |
| C1 |
C2 |
H10 |
109.320 |
|
C1 |
C3 |
C5 |
124.533 |
| C1 |
C3 |
H11 |
116.485 |
|
C2 |
C1 |
C3 |
111.863 |
| C2 |
C1 |
H7 |
108.708 |
|
C2 |
C1 |
H8 |
109.320 |
| C2 |
C4 |
C6 |
124.533 |
|
C2 |
C4 |
H12 |
116.485 |
| C3 |
C1 |
H7 |
109.683 |
|
C3 |
C1 |
H8 |
109.921 |
| C3 |
C5 |
H13 |
121.495 |
|
C3 |
C5 |
H14 |
121.109 |
| C4 |
C2 |
H9 |
109.683 |
|
C4 |
C2 |
H10 |
109.921 |
| C4 |
C6 |
H15 |
121.495 |
|
C4 |
C6 |
H16 |
121.109 |
| C5 |
C3 |
H11 |
118.970 |
|
C6 |
C4 |
H12 |
118.970 |
| H7 |
C1 |
H8 |
107.226 |
|
H9 |
C2 |
H10 |
107.226 |
| H13 |
C5 |
H14 |
117.396 |
|
H15 |
C6 |
H16 |
117.396 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
