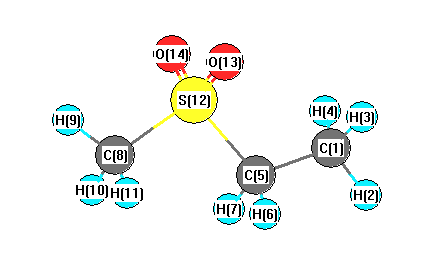
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3212 |
3078 |
0.80 |
|
|
|
| 2 |
A |
3207 |
3073 |
0.78 |
|
|
|
| 3 |
A |
3176 |
3044 |
6.74 |
|
|
|
| 4 |
A |
3160 |
3028 |
2.49 |
|
|
|
| 5 |
A |
3148 |
3017 |
11.94 |
|
|
|
| 6 |
A |
3096 |
2967 |
3.87 |
|
|
|
| 7 |
A |
3085 |
2956 |
1.94 |
|
|
|
| 8 |
A |
3072 |
2944 |
15.43 |
|
|
|
| 9 |
A |
1479 |
1418 |
6.48 |
|
|
|
| 10 |
A |
1476 |
1414 |
8.69 |
|
|
|
| 11 |
A |
1436 |
1376 |
8.70 |
|
|
|
| 12 |
A |
1428 |
1368 |
2.87 |
|
|
|
| 13 |
A |
1423 |
1364 |
4.81 |
|
|
|
| 14 |
A |
1393 |
1335 |
2.61 |
|
|
|
| 15 |
A |
1310 |
1255 |
20.84 |
|
|
|
| 16 |
A |
1289 |
1235 |
57.04 |
|
|
|
| 17 |
A |
1265 |
1213 |
131.32 |
|
|
|
| 18 |
A |
1243 |
1191 |
35.88 |
|
|
|
| 19 |
A |
1102 |
1056 |
131.86 |
|
|
|
| 20 |
A |
1087 |
1042 |
1.96 |
|
|
|
| 21 |
A |
1045 |
1001 |
14.20 |
|
|
|
| 22 |
A |
986 |
945 |
3.17 |
|
|
|
| 23 |
A |
949 |
909 |
31.84 |
|
|
|
| 24 |
A |
936 |
897 |
1.11 |
|
|
|
| 25 |
A |
785 |
752 |
42.54 |
|
|
|
| 26 |
A |
718 |
688 |
16.89 |
|
|
|
| 27 |
A |
642 |
615 |
9.42 |
|
|
|
| 28 |
A |
470 |
450 |
18.44 |
|
|
|
| 29 |
A |
426 |
408 |
31.43 |
|
|
|
| 30 |
A |
384 |
368 |
7.93 |
|
|
|
| 31 |
A |
302 |
289 |
0.95 |
|
|
|
| 32 |
A |
272 |
261 |
1.63 |
|
|
|
| 33 |
A |
217 |
208 |
2.30 |
|
|
|
| 34 |
A |
201 |
193 |
0.65 |
|
|
|
| 35 |
A |
176 |
169 |
0.18 |
|
|
|
| 36 |
A |
81 |
78 |
2.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24838.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 23802.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.471 |
|
|
|
| 2 |
H |
-0.072 |
|
|
|
| 3 |
H |
-0.120 |
|
|
|
| 4 |
H |
-0.149 |
|
|
|
| 5 |
C |
0.171 |
|
|
|
| 6 |
H |
-0.106 |
|
|
|
| 7 |
H |
-0.156 |
|
|
|
| 8 |
C |
0.161 |
|
|
|
| 9 |
H |
-0.046 |
|
|
|
| 10 |
H |
-0.034 |
|
|
|
| 11 |
H |
-0.026 |
|
|
|
| 12 |
S |
1.796 |
|
|
|
| 13 |
O |
-0.954 |
|
|
|
| 14 |
O |
-0.938 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.392 |
2.941 |
-2.299 |
5.044 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.490 |
-5.489 |
-1.323 |
| y |
-5.489 |
-41.938 |
-0.762 |
| z |
-1.323 |
-0.762 |
-48.775 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.867 |
-5.489 |
-1.323 |
| y |
-5.489 |
4.694 |
-0.762 |
| z |
-1.323 |
-0.762 |
-5.561 |
|
| Polar |
|---|
| 3z2-r2 | -11.123 |
|---|
| x2-y2 | -2.552 |
|---|
| xy | -5.489 |
|---|
| xz | -1.323 |
|---|
| yz | -0.762 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.726 |
-0.135 |
0.100 |
| y |
-0.135 |
9.687 |
0.044 |
| z |
0.100 |
0.044 |
9.182 |
<r2> (average value of r
2) Å
2
| <r2> |
193.283 |
| (<r2>)1/2 |
13.903 |