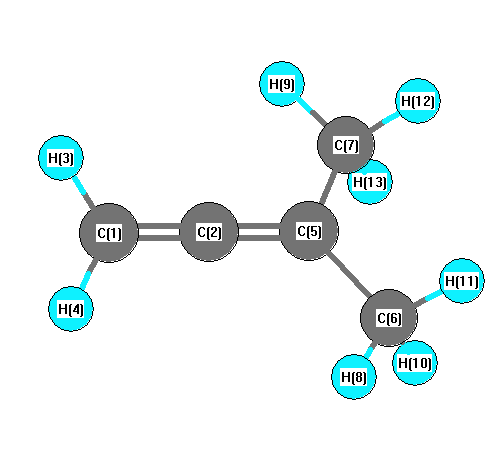
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3167 |
3035 |
15.90 |
|
|
|
| 2 |
A1 |
3143 |
3012 |
9.38 |
|
|
|
| 3 |
A1 |
3043 |
2916 |
60.53 |
|
|
|
| 4 |
A1 |
2085 |
1998 |
35.20 |
|
|
|
| 5 |
A1 |
1505 |
1442 |
2.63 |
|
|
|
| 6 |
A1 |
1456 |
1396 |
7.91 |
|
|
|
| 7 |
A1 |
1397 |
1338 |
0.56 |
|
|
|
| 8 |
A1 |
1318 |
1263 |
19.87 |
|
|
|
| 9 |
A1 |
1027 |
984 |
6.47 |
|
|
|
| 10 |
A1 |
753 |
722 |
0.08 |
|
|
|
| 11 |
A1 |
343 |
329 |
0.18 |
|
|
|
| 12 |
A2 |
3104 |
2975 |
0.00 |
|
|
|
| 13 |
A2 |
1452 |
1392 |
0.00 |
|
|
|
| 14 |
A2 |
988 |
947 |
0.00 |
|
|
|
| 15 |
A2 |
611 |
586 |
0.00 |
|
|
|
| 16 |
A2 |
166 |
159 |
0.00 |
|
|
|
| 17 |
B1 |
3227 |
3092 |
3.45 |
|
|
|
| 18 |
B1 |
3105 |
2975 |
39.30 |
|
|
|
| 19 |
B1 |
1471 |
1409 |
16.07 |
|
|
|
| 20 |
B1 |
1083 |
1038 |
0.03 |
|
|
|
| 21 |
B1 |
1010 |
967 |
1.51 |
|
|
|
| 22 |
B1 |
459 |
440 |
0.43 |
|
|
|
| 23 |
B1 |
251 |
241 |
5.64 |
|
|
|
| 24 |
B1 |
184 |
177 |
2.14 |
|
|
|
| 25 |
B2 |
3166 |
3034 |
14.58 |
|
|
|
| 26 |
B2 |
3039 |
2913 |
18.80 |
|
|
|
| 27 |
B2 |
1460 |
1400 |
6.81 |
|
|
|
| 28 |
B2 |
1383 |
1325 |
7.31 |
|
|
|
| 29 |
B2 |
1238 |
1186 |
4.59 |
|
|
|
| 30 |
B2 |
967 |
927 |
0.17 |
|
|
|
| 31 |
B2 |
880 |
844 |
49.22 |
|
|
|
| 32 |
B2 |
602 |
577 |
13.50 |
|
|
|
| 33 |
B2 |
183 |
176 |
0.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24631.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 23604.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.657 |
|
|
|
| 2 |
C |
0.051 |
|
|
|
| 3 |
H |
-0.353 |
|
|
|
| 4 |
H |
-0.353 |
|
|
|
| 5 |
C |
0.025 |
|
|
|
| 6 |
C |
0.247 |
|
|
|
| 7 |
C |
0.247 |
|
|
|
| 8 |
H |
-0.211 |
|
|
|
| 9 |
H |
-0.211 |
|
|
|
| 10 |
H |
-0.024 |
|
|
|
| 11 |
H |
-0.024 |
|
|
|
| 12 |
H |
-0.024 |
|
|
|
| 13 |
H |
-0.024 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.645 |
0.645 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.995 |
0.000 |
0.000 |
| y |
0.000 |
-32.546 |
0.000 |
| z |
0.000 |
0.000 |
-29.659 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.893 |
0.000 |
0.000 |
| y |
0.000 |
-1.719 |
0.000 |
| z |
0.000 |
0.000 |
2.612 |
|
| Polar |
|---|
| 3z2-r2 | 5.223 |
|---|
| x2-y2 | 0.551 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.216 |
0.000 |
0.000 |
| y |
0.000 |
8.568 |
0.000 |
| z |
0.000 |
0.000 |
14.380 |
<r2> (average value of r
2) Å
2
| <r2> |
142.923 |
| (<r2>)1/2 |
11.955 |