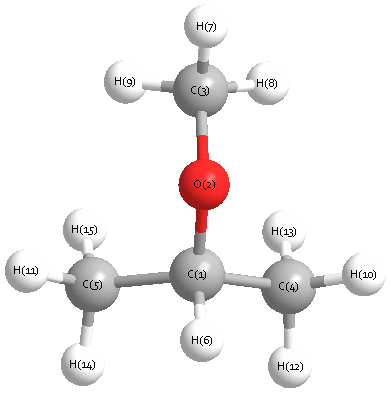
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3153 |
3022 |
38.61 |
|
|
|
| 2 |
A |
3151 |
3020 |
16.18 |
|
|
|
| 3 |
A |
3150 |
3019 |
9.96 |
|
|
|
| 4 |
A |
3143 |
3012 |
73.83 |
|
|
|
| 5 |
A |
3136 |
3005 |
0.17 |
|
|
|
| 6 |
A |
3080 |
2952 |
62.18 |
|
|
|
| 7 |
A |
3080 |
2951 |
14.51 |
|
|
|
| 8 |
A |
3062 |
2934 |
12.88 |
|
|
|
| 9 |
A |
3056 |
2929 |
9.97 |
|
|
|
| 10 |
A |
3017 |
2891 |
53.28 |
|
|
|
| 11 |
A |
1492 |
1430 |
5.29 |
|
|
|
| 12 |
A |
1487 |
1425 |
9.66 |
|
|
|
| 13 |
A |
1480 |
1418 |
3.02 |
|
|
|
| 14 |
A |
1475 |
1413 |
7.53 |
|
|
|
| 15 |
A |
1462 |
1401 |
0.02 |
|
|
|
| 16 |
A |
1461 |
1400 |
1.84 |
|
|
|
| 17 |
A |
1446 |
1385 |
0.53 |
|
|
|
| 18 |
A |
1402 |
1344 |
21.86 |
|
|
|
| 19 |
A |
1387 |
1329 |
26.05 |
|
|
|
| 20 |
A |
1381 |
1323 |
1.67 |
|
|
|
| 21 |
A |
1348 |
1292 |
0.36 |
|
|
|
| 22 |
A |
1226 |
1175 |
45.42 |
|
|
|
| 23 |
A |
1200 |
1150 |
110.49 |
|
|
|
| 24 |
A |
1172 |
1123 |
4.38 |
|
|
|
| 25 |
A |
1162 |
1114 |
2.36 |
|
|
|
| 26 |
A |
1151 |
1103 |
13.74 |
|
|
|
| 27 |
A |
1074 |
1029 |
24.74 |
|
|
|
| 28 |
A |
933 |
894 |
0.01 |
|
|
|
| 29 |
A |
931 |
892 |
14.33 |
|
|
|
| 30 |
A |
911 |
873 |
0.29 |
|
|
|
| 31 |
A |
797 |
764 |
9.35 |
|
|
|
| 32 |
A |
553 |
530 |
1.96 |
|
|
|
| 33 |
A |
420 |
402 |
8.48 |
|
|
|
| 34 |
A |
368 |
353 |
1.26 |
|
|
|
| 35 |
A |
308 |
295 |
0.71 |
|
|
|
| 36 |
A |
246 |
236 |
1.76 |
|
|
|
| 37 |
A |
244 |
234 |
0.03 |
|
|
|
| 38 |
A |
168 |
161 |
1.79 |
|
|
|
| 39 |
A |
24 |
23 |
2.40 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29866.9 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 28621.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.867 |
|
|
|
| 2 |
H |
-0.126 |
|
|
|
| 3 |
H |
-0.282 |
|
|
|
| 4 |
H |
-0.127 |
|
|
|
| 5 |
O |
-0.695 |
|
|
|
| 6 |
C |
0.766 |
|
|
|
| 7 |
H |
-0.166 |
|
|
|
| 8 |
H |
-0.114 |
|
|
|
| 9 |
H |
-0.118 |
|
|
|
| 10 |
C |
0.766 |
|
|
|
| 11 |
H |
-0.168 |
|
|
|
| 12 |
H |
-0.117 |
|
|
|
| 13 |
H |
-0.114 |
|
|
|
| 14 |
C |
0.082 |
|
|
|
| 15 |
H |
-0.453 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.685 |
0.003 |
1.124 |
1.316 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.733 |
0.001 |
2.207 |
| y |
0.001 |
-33.442 |
-0.004 |
| z |
2.207 |
-0.004 |
-34.413 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.194 |
0.001 |
2.207 |
| y |
0.001 |
-0.369 |
-0.004 |
| z |
2.207 |
-0.004 |
-1.826 |
|
| Polar |
|---|
| 3z2-r2 | -3.651 |
|---|
| x2-y2 | 1.709 |
|---|
| xy | 0.001 |
|---|
| xz | 2.207 |
|---|
| yz | -0.004 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.171 |
0.000 |
0.183 |
| y |
0.000 |
8.503 |
-0.000 |
| z |
0.183 |
-0.000 |
7.704 |
<r2> (average value of r
2) Å
2
| <r2> |
133.820 |
| (<r2>)1/2 |
11.568 |