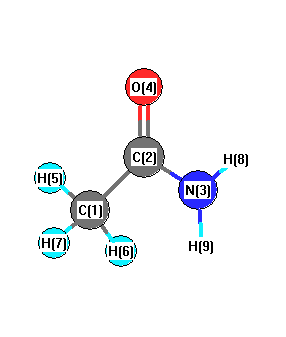
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3774 |
3617 |
40.14 |
|
|
|
| 2 |
A |
3628 |
3477 |
35.46 |
|
|
|
| 3 |
A |
3188 |
3055 |
6.17 |
|
|
|
| 4 |
A |
3153 |
3022 |
11.51 |
|
|
|
| 5 |
A |
3072 |
2943 |
6.91 |
|
|
|
| 6 |
A |
1791 |
1716 |
383.66 |
|
|
|
| 7 |
A |
1623 |
1556 |
112.63 |
|
|
|
| 8 |
A |
1469 |
1407 |
15.80 |
|
|
|
| 9 |
A |
1447 |
1387 |
9.74 |
|
|
|
| 10 |
A |
1395 |
1337 |
120.95 |
|
|
|
| 11 |
A |
1340 |
1284 |
43.50 |
|
|
|
| 12 |
A |
1116 |
1070 |
0.44 |
|
|
|
| 13 |
A |
1040 |
996 |
3.82 |
|
|
|
| 14 |
A |
979 |
938 |
7.78 |
|
|
|
| 15 |
A |
864 |
828 |
1.31 |
|
|
|
| 16 |
A |
662 |
634 |
8.07 |
|
|
|
| 17 |
A |
548 |
525 |
14.35 |
|
|
|
| 18 |
A |
522 |
500 |
6.74 |
|
|
|
| 19 |
A |
423 |
405 |
4.39 |
|
|
|
| 20 |
A |
208 |
199 |
187.28 |
|
|
|
| 21 |
A |
41 |
39 |
2.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16141.1 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 15468.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.361 |
|
|
|
| 2 |
C |
0.602 |
|
|
|
| 3 |
N |
-0.037 |
|
|
|
| 4 |
O |
-0.629 |
|
|
|
| 5 |
H |
-0.054 |
|
|
|
| 6 |
H |
-0.147 |
|
|
|
| 7 |
H |
0.005 |
|
|
|
| 8 |
H |
-0.057 |
|
|
|
| 9 |
H |
-0.043 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.237 |
-3.931 |
0.078 |
3.939 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.554 |
-2.377 |
0.105 |
| y |
-2.377 |
-26.395 |
-0.074 |
| z |
0.105 |
-0.074 |
-25.153 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.220 |
-2.377 |
0.105 |
| y |
-2.377 |
-3.541 |
-0.074 |
| z |
0.105 |
-0.074 |
-1.678 |
|
| Polar |
|---|
| 3z2-r2 | -3.357 |
|---|
| x2-y2 | 5.840 |
|---|
| xy | -2.377 |
|---|
| xz | 0.105 |
|---|
| yz | -0.074 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.375 |
-0.156 |
-0.032 |
| y |
-0.156 |
6.562 |
-0.007 |
| z |
-0.032 |
-0.007 |
4.449 |
<r2> (average value of r
2) Å
2
| <r2> |
75.144 |
| (<r2>)1/2 |
8.669 |