Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -190.626253 |
| Energy at 298.15K | |
| HF Energy | -190.626253 |
| Nuclear repulsion energy | 88.215295 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3104 |
2975 |
30.87 |
228.04 |
0.10 |
0.19 |
| 2 |
A1 |
2350 |
2252 |
1010.93 |
48.62 |
0.52 |
0.68 |
| 3 |
A1 |
1814 |
1738 |
42.34 |
2.47 |
0.40 |
0.57 |
| 4 |
A1 |
1468 |
1407 |
2.15 |
11.30 |
0.56 |
0.72 |
| 5 |
A1 |
938 |
899 |
6.51 |
52.11 |
0.16 |
0.28 |
| 6 |
B1 |
1030 |
987 |
24.53 |
1.70 |
0.75 |
0.86 |
| 7 |
B1 |
631 |
605 |
18.57 |
0.19 |
0.75 |
0.86 |
| 8 |
B1 |
205 |
197 |
0.62 |
2.00 |
0.75 |
0.86 |
| 9 |
B2 |
3184 |
3051 |
5.97 |
140.21 |
0.75 |
0.86 |
| 10 |
B2 |
1051 |
1007 |
2.14 |
0.00 |
0.75 |
0.86 |
| 11 |
B2 |
444 |
426 |
6.76 |
2.88 |
0.75 |
0.86 |
| 12 |
B2 |
173i |
165i |
15.87 |
0.13 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8022.8 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 7688.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.877 |
| C2 |
0.000 |
0.000 |
-0.558 |
| C3 |
0.000 |
0.000 |
0.723 |
| O4 |
0.000 |
0.000 |
1.899 |
| H5 |
0.000 |
0.929 |
-2.458 |
| H6 |
0.000 |
-0.929 |
-2.458 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3198 | 2.6006 | 3.7760 | 1.0950 | 1.0950 |
C2 | 1.3198 | | 1.2807 | 2.4562 | 2.1149 | 2.1149 | C3 | 2.6006 | 1.2807 | | 1.1755 | 3.3137 | 3.3137 | O4 | 3.7760 | 2.4562 | 1.1755 | | 4.4542 | 4.4542 | H5 | 1.0950 | 2.1149 | 3.3137 | 4.4542 | | 1.8570 | H6 | 1.0950 | 2.1149 | 3.3137 | 4.4542 | 1.8570 | |
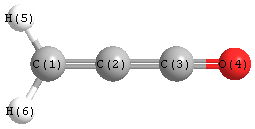 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.007 |
| C2 |
C1 |
H6 |
122.007 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.986 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.323 |
|
|
|
| 2 |
C |
0.339 |
|
|
|
| 3 |
C |
0.303 |
|
|
|
| 4 |
O |
-0.425 |
|
|
|
| 5 |
H |
-0.270 |
|
|
|
| 6 |
H |
-0.270 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.852 |
2.852 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.787 |
0.000 |
0.000 |
| y |
0.000 |
-22.561 |
0.000 |
| z |
0.000 |
0.000 |
-21.921 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.547 |
0.000 |
0.000 |
| y |
0.000 |
-0.207 |
0.000 |
| z |
0.000 |
0.000 |
0.753 |
|
| Polar |
|---|
| 3z2-r2 | 1.507 |
|---|
| x2-y2 | -0.227 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.691 |
0.000 |
0.000 |
| y |
0.000 |
4.180 |
0.000 |
| z |
0.000 |
0.000 |
11.292 |
<r2> (average value of r
2) Å
2
| <r2> |
82.799 |
| (<r2>)1/2 |
9.099 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -190.627683 |
| Energy at 298.15K | |
| HF Energy | -190.627683 |
| Nuclear repulsion energy | 88.495283 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3203 |
3069 |
5.48 |
116.03 |
0.72 |
0.83 |
| 2 |
A' |
3115 |
2985 |
28.84 |
178.94 |
0.11 |
0.19 |
| 3 |
A' |
2246 |
2152 |
1025.39 |
54.67 |
0.53 |
0.70 |
| 4 |
A' |
1781 |
1706 |
10.08 |
6.58 |
0.17 |
0.29 |
| 5 |
A' |
1471 |
1410 |
1.18 |
11.71 |
0.60 |
0.75 |
| 6 |
A' |
1064 |
1020 |
21.85 |
4.61 |
0.02 |
0.04 |
| 7 |
A' |
962 |
922 |
3.21 |
42.09 |
0.11 |
0.19 |
| 8 |
A' |
507 |
486 |
12.60 |
4.53 |
0.75 |
0.86 |
| 9 |
A' |
168 |
161 |
23.52 |
5.84 |
0.72 |
0.84 |
| 10 |
A" |
1037 |
994 |
24.43 |
1.92 |
0.75 |
0.86 |
| 11 |
A" |
703 |
674 |
12.12 |
0.34 |
0.75 |
0.86 |
| 12 |
A" |
271 |
260 |
1.64 |
2.52 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8263.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 7919.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.019 |
-1.510 |
0.000 |
| C2 |
0.000 |
-0.658 |
0.000 |
| C3 |
-0.347 |
0.601 |
0.000 |
| O4 |
-0.869 |
1.647 |
0.000 |
| H5 |
2.064 |
-1.184 |
0.000 |
| H6 |
0.854 |
-2.590 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3288 | 2.5146 | 3.6783 | 1.0949 | 1.0930 |
C2 | 1.3288 | | 1.3054 | 2.4626 | 2.1304 | 2.1133 | C3 | 2.5146 | 1.3054 | | 1.1687 | 3.0000 | 3.4100 | O4 | 3.6783 | 2.4626 | 1.1687 | | 4.0762 | 4.5742 | H5 | 1.0949 | 2.1304 | 3.0000 | 4.0762 | | 1.8558 | H6 | 1.0930 | 2.1133 | 3.4100 | 4.5742 | 1.8558 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
145.332 |
|
C2 |
C1 |
H5 |
122.749 |
| C2 |
C1 |
H6 |
121.222 |
|
C2 |
C3 |
O4 |
168.911 |
| H5 |
C1 |
H6 |
116.030 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.294 |
|
|
|
| 2 |
C |
0.496 |
|
|
|
| 3 |
C |
0.237 |
|
|
|
| 4 |
O |
-0.378 |
|
|
|
| 5 |
H |
-0.345 |
|
|
|
| 6 |
H |
-0.303 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.920 |
-1.636 |
0.000 |
2.522 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.948 |
-0.913 |
0.000 |
| y |
-0.913 |
-22.906 |
0.000 |
| z |
0.000 |
0.000 |
-22.956 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.983 |
-0.913 |
0.000 |
| y |
-0.913 |
-0.453 |
0.000 |
| z |
0.000 |
0.000 |
-0.529 |
|
| Polar |
|---|
| 3z2-r2 | -1.059 |
|---|
| x2-y2 | 0.957 |
|---|
| xy | -0.913 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.095 |
-2.672 |
0.000 |
| y |
-2.672 |
9.239 |
0.000 |
| z |
0.000 |
0.000 |
3.792 |
<r2> (average value of r
2) Å
2
| <r2> |
80.352 |
| (<r2>)1/2 |
8.964 |
